Jump to
S1C2
Vibrational Frequencies calculated at QCISD/6-31G**
Geometric Data calculated at QCISD/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -230.535676 |
| Energy at 298.15K | -230.541732 |
| HF Energy | -229.770118 |
| Nuclear repulsion energy | 161.197310 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3348 |
3151 |
13.11 |
|
|
|
| 2 |
A |
3347 |
3151 |
4.33 |
|
|
|
| 3 |
A |
3282 |
3090 |
12.70 |
|
|
|
| 4 |
A |
3254 |
3064 |
6.32 |
|
|
|
| 5 |
A |
3250 |
3059 |
1.47 |
|
|
|
| 6 |
A |
3235 |
3046 |
6.74 |
|
|
|
| 7 |
A |
1757 |
1654 |
88.27 |
|
|
|
| 8 |
A |
1742 |
1640 |
116.79 |
|
|
|
| 9 |
A |
1487 |
1400 |
2.18 |
|
|
|
| 10 |
A |
1469 |
1383 |
28.45 |
|
|
|
| 11 |
A |
1392 |
1311 |
33.30 |
|
|
|
| 12 |
A |
1359 |
1280 |
2.88 |
|
|
|
| 13 |
A |
1273 |
1199 |
250.62 |
|
|
|
| 14 |
A |
1159 |
1091 |
12.12 |
|
|
|
| 15 |
A |
1032 |
972 |
41.82 |
|
|
|
| 16 |
A |
1007 |
948 |
7.50 |
|
|
|
| 17 |
A |
994 |
936 |
35.15 |
|
|
|
| 18 |
A |
898 |
846 |
15.93 |
|
|
|
| 19 |
A |
874 |
822 |
41.64 |
|
|
|
| 20 |
A |
856 |
806 |
44.00 |
|
|
|
| 21 |
A |
743 |
699 |
3.13 |
|
|
|
| 22 |
A |
722 |
680 |
0.68 |
|
|
|
| 23 |
A |
591 |
556 |
2.77 |
|
|
|
| 24 |
A |
458 |
431 |
3.18 |
|
|
|
| 25 |
A |
293 |
276 |
1.75 |
|
|
|
| 26 |
A |
193 |
181 |
8.58 |
|
|
|
| 27 |
A |
93 |
88 |
0.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20054.5 cm
-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 18879.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.042 |
-0.822 |
0.051 |
| C2 |
1.291 |
-0.489 |
-0.008 |
| C3 |
-0.927 |
0.200 |
0.336 |
| C4 |
1.810 |
0.735 |
-0.150 |
| C5 |
-2.139 |
0.234 |
-0.218 |
| H6 |
1.899 |
-1.380 |
0.062 |
| H7 |
-0.578 |
0.912 |
1.075 |
| H8 |
2.883 |
0.847 |
-0.174 |
| H9 |
1.207 |
1.623 |
-0.263 |
| H10 |
-2.846 |
0.993 |
0.079 |
| H11 |
-2.439 |
-0.496 |
-0.955 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3757 | 1.3818 | 2.4282 | 2.3630 | 2.0196 | 2.0844 | 3.3755 | 2.7635 | 3.3404 | 2.6195 |
C2 | 1.3757 | | 2.3481 | 1.3367 | 3.5114 | 1.0812 | 2.5750 | 2.0846 | 2.1285 | 4.3953 | 3.8480 | C3 | 1.3818 | 2.3481 | | 2.8310 | 1.3326 | 3.2493 | 1.0842 | 3.8983 | 2.6338 | 2.0922 | 2.1061 | C4 | 2.4282 | 1.3367 | 2.8310 | | 3.9811 | 2.1276 | 2.6899 | 1.0791 | 1.0794 | 4.6686 | 4.4963 | C5 | 2.3630 | 3.5114 | 1.3326 | 3.9811 | | 4.3573 | 2.1372 | 5.0593 | 3.6226 | 1.0793 | 1.0800 | H6 | 2.0196 | 1.0812 | 3.2493 | 2.1276 | 4.3573 | | 3.5237 | 2.4462 | 3.0988 | 5.3053 | 4.5420 | H7 | 2.0844 | 2.5750 | 1.0842 | 2.6899 | 2.1372 | 3.5237 | | 3.6804 | 2.3412 | 2.4781 | 3.0930 | H8 | 3.3755 | 2.0846 | 3.8983 | 1.0791 | 5.0593 | 2.4462 | 3.6804 | | 1.8494 | 5.7363 | 5.5439 | H9 | 2.7635 | 2.1285 | 2.6338 | 1.0794 | 3.6226 | 3.0988 | 2.3412 | 1.8494 | | 4.1152 | 4.2730 | H10 | 3.3404 | 4.3953 | 2.0922 | 4.6686 | 1.0793 | 5.3053 | 2.4781 | 5.7363 | 4.1152 | | 1.8586 | H11 | 2.6195 | 3.8480 | 2.1061 | 4.4963 | 1.0800 | 4.5420 | 3.0930 | 5.5439 | 4.2730 | 1.8586 | |
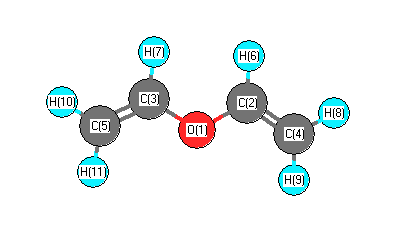 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
127.078 |
|
O1 |
C2 |
H6 |
109.998 |
| O1 |
C3 |
C5 |
121.033 |
|
O1 |
C3 |
H7 |
114.863 |
| C2 |
O1 |
C3 |
116.758 |
|
C2 |
C4 |
H8 |
118.905 |
| C2 |
C4 |
H9 |
123.164 |
|
C3 |
C5 |
H10 |
119.951 |
| C3 |
C5 |
H11 |
121.251 |
|
C4 |
C2 |
H6 |
122.918 |
| C5 |
C3 |
H7 |
123.998 |
|
H8 |
C4 |
H9 |
117.918 |
| H10 |
C5 |
H11 |
118.793 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
