Jump to
S1C2
Vibrational Frequencies calculated at PBE1PBE/6-31G**
Geometric Data calculated at PBE1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -230.918605 |
| Energy at 298.15K | -230.924694 |
| HF Energy | -230.918605 |
| Nuclear repulsion energy | 161.785393 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3312 |
3156 |
4.22 |
|
|
|
| 2 |
A |
3309 |
3154 |
6.63 |
|
|
|
| 3 |
A |
3246 |
3094 |
10.67 |
|
|
|
| 4 |
A |
3220 |
3069 |
1.62 |
|
|
|
| 5 |
A |
3218 |
3067 |
3.08 |
|
|
|
| 6 |
A |
3195 |
3045 |
10.17 |
|
|
|
| 7 |
A |
1756 |
1674 |
95.76 |
|
|
|
| 8 |
A |
1735 |
1653 |
184.39 |
|
|
|
| 9 |
A |
1459 |
1390 |
1.63 |
|
|
|
| 10 |
A |
1441 |
1373 |
30.75 |
|
|
|
| 11 |
A |
1384 |
1319 |
67.98 |
|
|
|
| 12 |
A |
1343 |
1280 |
5.08 |
|
|
|
| 13 |
A |
1280 |
1220 |
261.90 |
|
|
|
| 14 |
A |
1161 |
1106 |
16.41 |
|
|
|
| 15 |
A |
1030 |
982 |
28.25 |
|
|
|
| 16 |
A |
1013 |
965 |
5.88 |
|
|
|
| 17 |
A |
998 |
951 |
32.31 |
|
|
|
| 18 |
A |
899 |
857 |
19.09 |
|
|
|
| 19 |
A |
878 |
837 |
48.82 |
|
|
|
| 20 |
A |
859 |
819 |
61.56 |
|
|
|
| 21 |
A |
743 |
708 |
4.45 |
|
|
|
| 22 |
A |
727 |
693 |
1.58 |
|
|
|
| 23 |
A |
595 |
567 |
2.41 |
|
|
|
| 24 |
A |
458 |
436 |
2.46 |
|
|
|
| 25 |
A |
289 |
275 |
1.67 |
|
|
|
| 26 |
A |
212 |
202 |
9.21 |
|
|
|
| 27 |
A |
96 |
92 |
0.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19926.4 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 18991.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.045 |
-0.793 |
0.046 |
| C2 |
1.281 |
-0.493 |
0.005 |
| C3 |
-0.923 |
0.221 |
0.301 |
| C4 |
1.834 |
0.708 |
-0.149 |
| C5 |
-2.155 |
0.217 |
-0.194 |
| H6 |
1.867 |
-1.402 |
0.106 |
| H7 |
-0.545 |
0.986 |
0.978 |
| H8 |
2.914 |
0.795 |
-0.150 |
| H9 |
1.254 |
1.613 |
-0.299 |
| H10 |
-2.855 |
0.994 |
0.091 |
| H11 |
-2.490 |
-0.560 |
-0.873 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3601 | 1.3655 | 2.4131 | 2.3518 | 2.0076 | 2.0696 | 3.3640 | 2.7555 | 3.3300 | 2.6222 |
C2 | 1.3601 | | 2.3359 | 1.3317 | 3.5145 | 1.0858 | 2.5434 | 2.0858 | 2.1278 | 4.3956 | 3.8723 | C3 | 1.3655 | 2.3359 | | 2.8358 | 1.3275 | 3.2339 | 1.0894 | 3.9061 | 2.6523 | 2.0906 | 2.1075 | C4 | 2.4131 | 1.3317 | 2.8358 | | 4.0195 | 2.1257 | 2.6469 | 1.0836 | 1.0846 | 4.7031 | 4.5639 | C5 | 2.3518 | 3.5145 | 1.3275 | 4.0195 | | 4.3462 | 2.1346 | 5.1022 | 3.6849 | 1.0836 | 1.0849 | H6 | 2.0076 | 1.0858 | 3.2339 | 2.1257 | 4.3462 | | 3.5045 | 2.4468 | 3.1024 | 5.2947 | 4.5441 | H7 | 2.0696 | 2.5434 | 1.0894 | 2.6469 | 2.1346 | 3.5045 | | 3.6433 | 2.2930 | 2.4735 | 3.0980 | H8 | 3.3640 | 2.0858 | 3.9061 | 1.0836 | 5.1022 | 2.4468 | 3.6433 | | 1.8566 | 5.7769 | 5.6180 | H9 | 2.7555 | 2.1278 | 2.6523 | 1.0846 | 3.6849 | 3.1024 | 2.2930 | 1.8566 | | 4.1727 | 4.3663 | H10 | 3.3300 | 4.3956 | 2.0906 | 4.7031 | 1.0836 | 5.2947 | 2.4735 | 5.7769 | 4.1727 | | 1.8651 | H11 | 2.6222 | 3.8723 | 2.1075 | 4.5639 | 1.0849 | 4.5441 | 3.0980 | 5.6180 | 4.3663 | 1.8651 | |
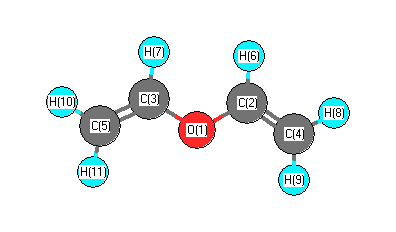 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
127.384 |
|
O1 |
C2 |
H6 |
109.822 |
| O1 |
C3 |
C5 |
121.681 |
|
O1 |
C3 |
H7 |
114.466 |
| C2 |
O1 |
C3 |
117.965 |
|
C2 |
C4 |
H8 |
119.082 |
| C2 |
C4 |
H9 |
123.096 |
|
C3 |
C5 |
H10 |
119.902 |
| C3 |
C5 |
H11 |
121.436 |
|
C4 |
C2 |
H6 |
122.789 |
| C5 |
C3 |
H7 |
123.767 |
|
H8 |
C4 |
H9 |
117.806 |
| H10 |
C5 |
H11 |
118.656 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.443 |
|
|
|
| 2 |
C |
0.126 |
|
|
|
| 3 |
C |
0.129 |
|
|
|
| 4 |
C |
-0.457 |
|
|
|
| 5 |
C |
-0.441 |
|
|
|
| 6 |
H |
0.182 |
|
|
|
| 7 |
H |
0.178 |
|
|
|
| 8 |
H |
0.180 |
|
|
|
| 9 |
H |
0.182 |
|
|
|
| 10 |
H |
0.178 |
|
|
|
| 11 |
H |
0.186 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.358 |
0.670 |
0.283 |
0.810 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.582 |
-1.677 |
0.567 |
| y |
-1.677 |
-28.961 |
0.970 |
| z |
0.567 |
0.970 |
-31.900 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.849 |
-1.677 |
0.567 |
| y |
-1.677 |
0.280 |
0.970 |
| z |
0.567 |
0.970 |
-4.129 |
|
| Polar |
|---|
| 3z2-r2 | -8.258 |
|---|
| x2-y2 | 2.379 |
|---|
| xy | -1.677 |
|---|
| xz | 0.567 |
|---|
| yz | 0.970 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.212 |
0.315 |
0.624 |
| y |
0.315 |
6.187 |
0.275 |
| z |
0.624 |
0.275 |
3.558 |
<r2> (average value of r
2) Å
2
| <r2> |
129.742 |
| (<r2>)1/2 |
11.390 |
