Jump to
S1C2
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
Geometric Data calculated at HSEh1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -230.947853 |
| Energy at 298.15K | -230.953936 |
| HF Energy | -230.947853 |
| Nuclear repulsion energy | 161.812333 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3309 |
3158 |
3.79 |
|
|
|
| 2 |
A |
3306 |
3155 |
6.23 |
|
|
|
| 3 |
A |
3238 |
3090 |
10.20 |
|
|
|
| 4 |
A |
3212 |
3065 |
1.30 |
|
|
|
| 5 |
A |
3210 |
3063 |
3.44 |
|
|
|
| 6 |
A |
3187 |
3041 |
10.23 |
|
|
|
| 7 |
A |
1751 |
1671 |
96.14 |
|
|
|
| 8 |
A |
1729 |
1650 |
183.89 |
|
|
|
| 9 |
A |
1446 |
1380 |
1.38 |
|
|
|
| 10 |
A |
1429 |
1363 |
30.36 |
|
|
|
| 11 |
A |
1376 |
1313 |
70.39 |
|
|
|
| 12 |
A |
1334 |
1273 |
4.95 |
|
|
|
| 13 |
A |
1273 |
1215 |
260.79 |
|
|
|
| 14 |
A |
1154 |
1102 |
16.47 |
|
|
|
| 15 |
A |
1025 |
978 |
29.08 |
|
|
|
| 16 |
A |
1011 |
965 |
4.11 |
|
|
|
| 17 |
A |
996 |
951 |
31.19 |
|
|
|
| 18 |
A |
895 |
854 |
22.17 |
|
|
|
| 19 |
A |
879 |
839 |
42.60 |
|
|
|
| 20 |
A |
861 |
821 |
58.12 |
|
|
|
| 21 |
A |
742 |
708 |
4.28 |
|
|
|
| 22 |
A |
727 |
694 |
1.46 |
|
|
|
| 23 |
A |
594 |
567 |
2.43 |
|
|
|
| 24 |
A |
457 |
436 |
2.43 |
|
|
|
| 25 |
A |
288 |
275 |
1.62 |
|
|
|
| 26 |
A |
213 |
203 |
9.17 |
|
|
|
| 27 |
A |
96 |
92 |
0.29 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19868.3 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 18960.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.046 |
-0.791 |
0.046 |
| C2 |
1.282 |
-0.494 |
0.007 |
| C3 |
-0.923 |
0.225 |
0.296 |
| C4 |
1.835 |
0.707 |
-0.148 |
| C5 |
-2.157 |
0.214 |
-0.192 |
| H6 |
1.868 |
-1.402 |
0.109 |
| H7 |
-0.543 |
0.998 |
0.962 |
| H8 |
2.914 |
0.792 |
-0.148 |
| H9 |
1.255 |
1.609 |
-0.300 |
| H10 |
-2.854 |
0.994 |
0.090 |
| H11 |
-2.492 |
-0.571 |
-0.860 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3606 | 1.3659 | 2.4122 | 2.3509 | 2.0095 | 2.0707 | 3.3623 | 2.7520 | 3.3282 | 2.6184 |
C2 | 1.3606 | | 2.3370 | 1.3312 | 3.5164 | 1.0853 | 2.5430 | 2.0840 | 2.1258 | 4.3961 | 3.8729 | C3 | 1.3659 | 2.3370 | | 2.8349 | 1.3269 | 3.2359 | 1.0889 | 3.9043 | 2.6483 | 2.0888 | 2.1052 | C4 | 2.4122 | 1.3312 | 2.8349 | | 4.0224 | 2.1242 | 2.6404 | 1.0827 | 1.0838 | 4.7040 | 4.5676 | C5 | 2.3509 | 3.5164 | 1.3269 | 4.0224 | | 4.3477 | 2.1335 | 5.1042 | 3.6873 | 1.0827 | 1.0840 | H6 | 2.0095 | 1.0853 | 3.2359 | 2.1242 | 4.3477 | | 3.5068 | 2.4437 | 3.0999 | 5.2950 | 4.5432 | H7 | 2.0707 | 2.5430 | 1.0889 | 2.6404 | 2.1335 | 3.5068 | | 3.6366 | 2.2797 | 2.4710 | 3.0954 | H8 | 3.3623 | 2.0840 | 3.9043 | 1.0827 | 5.1042 | 2.4437 | 3.6366 | | 1.8564 | 5.7769 | 5.6210 | H9 | 2.7520 | 2.1258 | 2.6483 | 1.0838 | 3.6873 | 3.0999 | 2.2797 | 1.8564 | | 4.1729 | 4.3710 | H10 | 3.3282 | 4.3961 | 2.0888 | 4.7040 | 1.0827 | 5.2950 | 2.4710 | 5.7769 | 4.1729 | | 1.8651 | H11 | 2.6184 | 3.8729 | 2.1052 | 4.5676 | 1.0840 | 4.5432 | 3.0954 | 5.6210 | 4.3710 | 1.8651 | |
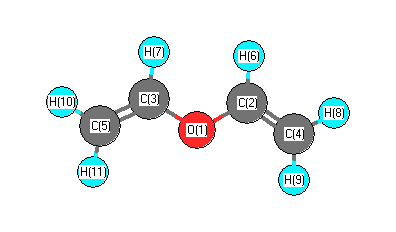 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
127.300 |
|
O1 |
C2 |
H6 |
109.967 |
| O1 |
C3 |
C5 |
121.613 |
|
O1 |
C3 |
H7 |
114.559 |
| C2 |
O1 |
C3 |
117.991 |
|
C2 |
C4 |
H8 |
119.028 |
| C2 |
C4 |
H9 |
123.017 |
|
C3 |
C5 |
H10 |
119.845 |
| C3 |
C5 |
H11 |
121.336 |
|
C4 |
C2 |
H6 |
122.728 |
| C5 |
C3 |
H7 |
123.745 |
|
H8 |
C4 |
H9 |
117.939 |
| H10 |
C5 |
H11 |
118.813 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.451 |
|
|
|
| 2 |
C |
0.163 |
|
|
|
| 3 |
C |
0.166 |
|
|
|
| 4 |
C |
-0.353 |
|
|
|
| 5 |
C |
-0.340 |
|
|
|
| 6 |
H |
0.137 |
|
|
|
| 7 |
H |
0.134 |
|
|
|
| 8 |
H |
0.135 |
|
|
|
| 9 |
H |
0.135 |
|
|
|
| 10 |
H |
0.134 |
|
|
|
| 11 |
H |
0.141 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.361 |
0.665 |
0.277 |
0.806 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.631 |
-1.653 |
0.552 |
| y |
-1.653 |
-28.962 |
0.941 |
| z |
0.552 |
0.941 |
-31.865 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.783 |
-1.653 |
0.552 |
| y |
-1.653 |
0.286 |
0.941 |
| z |
0.552 |
0.941 |
-4.069 |
|
| Polar |
|---|
| 3z2-r2 | -8.138 |
|---|
| x2-y2 | 2.331 |
|---|
| xy | -1.653 |
|---|
| xz | 0.552 |
|---|
| yz | 0.941 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.340 |
0.324 |
0.618 |
| y |
0.324 |
6.271 |
0.274 |
| z |
0.618 |
0.274 |
3.608 |
<r2> (average value of r
2) Å
2
| <r2> |
129.755 |
| (<r2>)1/2 |
11.391 |
