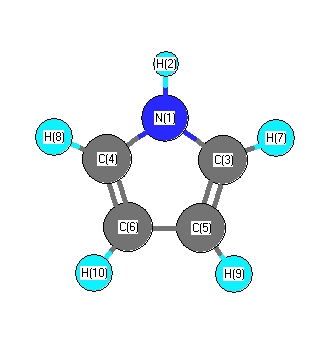
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3717 |
3543 |
60.95 |
|
|
|
| 2 |
A1 |
3312 |
3157 |
0.07 |
|
|
|
| 3 |
A1 |
3290 |
3136 |
4.64 |
|
|
|
| 4 |
A1 |
1538 |
1466 |
9.43 |
|
|
|
| 5 |
A1 |
1463 |
1395 |
4.07 |
|
|
|
| 6 |
A1 |
1202 |
1145 |
3.46 |
|
|
|
| 7 |
A1 |
1115 |
1063 |
7.84 |
|
|
|
| 8 |
A1 |
1056 |
1006 |
26.61 |
|
|
|
| 9 |
A1 |
902 |
859 |
0.16 |
|
|
|
| 10 |
A2 |
879 |
838 |
0.00 |
|
|
|
| 11 |
A2 |
682 |
650 |
0.00 |
|
|
|
| 12 |
A2 |
634 |
604 |
0.00 |
|
|
|
| 13 |
B1 |
827 |
788 |
4.52 |
|
|
|
| 14 |
B1 |
737 |
702 |
133.65 |
|
|
|
| 15 |
B1 |
649 |
619 |
0.63 |
|
|
|
| 16 |
B1 |
476 |
453 |
83.24 |
|
|
|
| 17 |
B2 |
3306 |
3151 |
5.35 |
|
|
|
| 18 |
B2 |
3279 |
3125 |
2.62 |
|
|
|
| 19 |
B2 |
1615 |
1539 |
4.98 |
|
|
|
| 20 |
B2 |
1505 |
1435 |
6.91 |
|
|
|
| 21 |
B2 |
1324 |
1262 |
2.77 |
|
|
|
| 22 |
B2 |
1184 |
1129 |
3.95 |
|
|
|
| 23 |
B2 |
1088 |
1037 |
24.44 |
|
|
|
| 24 |
B2 |
876 |
835 |
1.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18328.3 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 17468.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.588 |
|
|
|
| 2 |
H |
0.356 |
|
|
|
| 3 |
C |
0.004 |
|
|
|
| 4 |
C |
0.004 |
|
|
|
| 5 |
C |
-0.224 |
|
|
|
| 6 |
C |
-0.224 |
|
|
|
| 7 |
H |
0.176 |
|
|
|
| 8 |
H |
0.176 |
|
|
|
| 9 |
H |
0.161 |
|
|
|
| 10 |
H |
0.161 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.949 |
1.949 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.087 |
0.000 |
0.000 |
| y |
0.000 |
-26.033 |
0.000 |
| z |
0.000 |
0.000 |
-22.843 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.649 |
0.000 |
0.000 |
| y |
0.000 |
1.932 |
0.000 |
| z |
0.000 |
0.000 |
6.717 |
|
| Polar |
|---|
| 3z2-r2 | 13.434 |
|---|
| x2-y2 | -7.054 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.681 |
0.000 |
0.000 |
| y |
0.000 |
7.727 |
0.000 |
| z |
0.000 |
0.000 |
7.593 |
<r2> (average value of r
2) Å
2
| <r2> |
84.858 |
| (<r2>)1/2 |
9.212 |