Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Geometric Data calculated at MP2=FULL/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -251.122799 |
| Energy at 298.15K | -251.137265 |
| Nuclear repulsion energy | 259.679844 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3567 |
3333 |
0.43 |
|
|
|
| 2 |
A' |
3183 |
2975 |
78.30 |
|
|
|
| 3 |
A' |
3176 |
2968 |
25.70 |
|
|
|
| 4 |
A' |
3168 |
2960 |
24.56 |
|
|
|
| 5 |
A' |
3114 |
2910 |
38.30 |
|
|
|
| 6 |
A' |
3112 |
2907 |
22.54 |
|
|
|
| 7 |
A' |
3109 |
2905 |
25.86 |
|
|
|
| 8 |
A' |
1558 |
1456 |
1.91 |
|
|
|
| 9 |
A' |
1544 |
1442 |
9.98 |
|
|
|
| 10 |
A' |
1539 |
1438 |
1.35 |
|
|
|
| 11 |
A' |
1438 |
1344 |
5.74 |
|
|
|
| 12 |
A' |
1427 |
1333 |
0.59 |
|
|
|
| 13 |
A' |
1376 |
1286 |
0.21 |
|
|
|
| 14 |
A' |
1321 |
1234 |
10.01 |
|
|
|
| 15 |
A' |
1231 |
1150 |
4.05 |
|
|
|
| 16 |
A' |
1084 |
1013 |
0.50 |
|
|
|
| 17 |
A' |
1050 |
981 |
9.38 |
|
|
|
| 18 |
A' |
958 |
895 |
16.55 |
|
|
|
| 19 |
A' |
892 |
833 |
38.11 |
|
|
|
| 20 |
A' |
850 |
794 |
0.90 |
|
|
|
| 21 |
A' |
808 |
755 |
91.43 |
|
|
|
| 22 |
A' |
564 |
527 |
2.54 |
|
|
|
| 23 |
A' |
442 |
413 |
0.95 |
|
|
|
| 24 |
A' |
408 |
382 |
5.48 |
|
|
|
| 25 |
A' |
250 |
234 |
4.69 |
|
|
|
| 26 |
A" |
3181 |
2972 |
16.13 |
|
|
|
| 27 |
A" |
3168 |
2961 |
56.83 |
|
|
|
| 28 |
A" |
3111 |
2907 |
16.28 |
|
|
|
| 29 |
A" |
3107 |
2903 |
37.32 |
|
|
|
| 30 |
A" |
1538 |
1437 |
3.74 |
|
|
|
| 31 |
A" |
1538 |
1437 |
0.43 |
|
|
|
| 32 |
A" |
1511 |
1411 |
10.07 |
|
|
|
| 33 |
A" |
1426 |
1333 |
0.51 |
|
|
|
| 34 |
A" |
1414 |
1321 |
2.65 |
|
|
|
| 35 |
A" |
1386 |
1295 |
1.07 |
|
|
|
| 36 |
A" |
1337 |
1249 |
0.62 |
|
|
|
| 37 |
A" |
1253 |
1171 |
11.86 |
|
|
|
| 38 |
A" |
1176 |
1099 |
15.31 |
|
|
|
| 39 |
A" |
1147 |
1072 |
1.65 |
|
|
|
| 40 |
A" |
1101 |
1028 |
3.61 |
|
|
|
| 41 |
A" |
976 |
912 |
4.81 |
|
|
|
| 42 |
A" |
910 |
850 |
0.01 |
|
|
|
| 43 |
A" |
831 |
777 |
0.08 |
|
|
|
| 44 |
A" |
459 |
429 |
0.95 |
|
|
|
| 45 |
A" |
250 |
233 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35992.4 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 33631.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.649 |
1.313 |
0.000 |
| H2 |
0.600 |
2.403 |
0.000 |
| H3 |
1.709 |
1.044 |
0.000 |
| C4 |
-0.008 |
0.743 |
1.254 |
| C5 |
-0.008 |
0.743 |
-1.254 |
| C6 |
-0.008 |
-0.785 |
-1.207 |
| C7 |
-0.008 |
-0.785 |
1.207 |
| N8 |
-0.617 |
-1.344 |
0.000 |
| H9 |
-1.605 |
-1.102 |
0.000 |
| H10 |
0.502 |
1.093 |
2.153 |
| H11 |
0.502 |
1.093 |
-2.153 |
| H12 |
-1.042 |
1.096 |
1.315 |
| H13 |
-1.042 |
1.096 |
-1.315 |
| H14 |
1.024 |
-1.145 |
-1.244 |
| H15 |
1.024 |
-1.145 |
1.244 |
| H16 |
-0.522 |
-1.200 |
-2.075 |
| H17 |
-0.522 |
-1.200 |
2.075 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0907 | 1.0937 | 1.5265 | 1.5265 | 2.5080 | 2.5080 | 2.9431 | 3.3035 | 2.1696 | 2.1696 | 2.1527 | 2.1527 | 2.7804 | 2.7804 | 3.4629 | 3.4629 |
H2 | 1.0907 | | 1.7537 | 2.1677 | 2.1677 | 3.4623 | 3.4623 | 3.9393 | 4.1408 | 2.5223 | 2.5223 | 2.4764 | 2.4764 | 3.7834 | 3.7834 | 4.3063 | 4.3063 | H3 | 1.0937 | 1.7537 | | 2.1476 | 2.1476 | 2.7840 | 2.7840 | 3.3337 | 3.9482 | 2.4689 | 2.4689 | 3.0491 | 3.0491 | 2.6096 | 2.6096 | 3.7840 | 3.7840 | C4 | 1.5265 | 2.1677 | 2.1476 | | 2.5084 | 2.8968 | 1.5279 | 2.5093 | 2.7433 | 1.0918 | 3.4634 | 1.0937 | 2.7918 | 3.2967 | 2.1511 | 3.8884 | 2.1706 | C5 | 1.5265 | 2.1677 | 2.1476 | 2.5084 | | 1.5279 | 2.8968 | 2.5093 | 2.7433 | 3.4634 | 1.0918 | 2.7918 | 1.0937 | 2.1511 | 3.2967 | 2.1706 | 3.8884 | C6 | 2.5080 | 3.4623 | 2.7840 | 2.8968 | 1.5279 | | 2.4146 | 1.4633 | 2.0269 | 3.8835 | 2.1638 | 3.3115 | 2.1482 | 1.0935 | 2.6840 | 1.0904 | 3.3479 | C7 | 2.5080 | 3.4623 | 2.7840 | 1.5279 | 2.8968 | 2.4146 | | 1.4633 | 2.0269 | 2.1638 | 3.8835 | 2.1482 | 3.3115 | 2.6840 | 1.0935 | 3.3479 | 1.0904 | N8 | 2.9431 | 3.9393 | 3.3337 | 2.5093 | 2.5093 | 1.4633 | 1.4633 | | 1.0167 | 3.4393 | 3.4393 | 2.8035 | 2.8035 | 2.0689 | 2.0689 | 2.0818 | 2.0818 | H9 | 3.3035 | 4.1408 | 3.9482 | 2.7433 | 2.7433 | 2.0269 | 2.0269 | 1.0167 | | 3.7279 | 3.7279 | 2.6224 | 2.6224 | 2.9085 | 2.9085 | 2.3424 | 2.3424 | H10 | 2.1696 | 2.5223 | 2.4689 | 1.0918 | 3.4634 | 3.8835 | 2.1638 | 3.4393 | 3.7279 | | 4.3068 | 1.7567 | 3.7966 | 4.1017 | 2.4714 | 4.9179 | 2.5129 | H11 | 2.1696 | 2.5223 | 2.4689 | 3.4634 | 1.0918 | 2.1638 | 3.8835 | 3.4393 | 3.7279 | 4.3068 | | 3.7966 | 1.7567 | 2.4714 | 4.1017 | 2.5129 | 4.9179 | H12 | 2.1527 | 2.4764 | 3.0491 | 1.0937 | 2.7918 | 3.3115 | 2.1482 | 2.8035 | 2.6224 | 1.7567 | 3.7966 | | 2.6304 | 3.9793 | 3.0479 | 4.1270 | 2.4733 | H13 | 2.1527 | 2.4764 | 3.0491 | 2.7918 | 1.0937 | 2.1482 | 3.3115 | 2.8035 | 2.6224 | 3.7966 | 1.7567 | 2.6304 | | 3.0479 | 3.9793 | 2.4733 | 4.1270 | H14 | 2.7804 | 3.7834 | 2.6096 | 3.2967 | 2.1511 | 1.0935 | 2.6840 | 2.0689 | 2.9085 | 4.1017 | 2.4714 | 3.9793 | 3.0479 | | 2.4882 | 1.7555 | 3.6615 | H15 | 2.7804 | 3.7834 | 2.6096 | 2.1511 | 3.2967 | 2.6840 | 1.0935 | 2.0689 | 2.9085 | 2.4714 | 4.1017 | 3.0479 | 3.9793 | 2.4882 | | 3.6615 | 1.7555 | H16 | 3.4629 | 4.3063 | 3.7840 | 3.8884 | 2.1706 | 1.0904 | 3.3479 | 2.0818 | 2.3424 | 4.9179 | 2.5129 | 4.1270 | 2.4733 | 1.7555 | 3.6615 | | 4.1494 | H17 | 3.4629 | 4.3063 | 3.7840 | 2.1706 | 3.8884 | 3.3479 | 1.0904 | 2.0818 | 2.3424 | 2.5129 | 4.9179 | 2.4733 | 4.1270 | 3.6615 | 1.7555 | 4.1494 | |
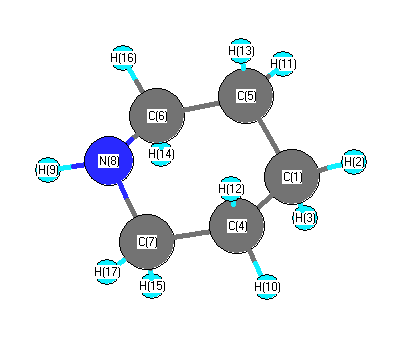 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.392 |
|
C1 |
C4 |
H10 |
110.824 |
| C1 |
C4 |
H12 |
109.373 |
|
C1 |
C5 |
C6 |
110.392 |
| C1 |
C5 |
H11 |
110.824 |
|
C1 |
C5 |
H13 |
109.373 |
| H2 |
C1 |
H3 |
106.801 |
|
H2 |
C1 |
C4 |
110.743 |
| H2 |
C1 |
C5 |
110.743 |
|
H3 |
C1 |
C4 |
108.977 |
| H3 |
C1 |
C5 |
108.977 |
|
C4 |
C1 |
C5 |
110.497 |
| C4 |
C7 |
N8 |
114.026 |
|
C4 |
C7 |
H15 |
109.164 |
| C4 |
C7 |
H17 |
110.885 |
|
C5 |
C6 |
N8 |
114.026 |
| C5 |
C6 |
H14 |
109.164 |
|
C5 |
C6 |
H16 |
110.885 |
| C6 |
C5 |
H11 |
110.258 |
|
C6 |
C5 |
H13 |
108.927 |
| C6 |
N8 |
C7 |
111.189 |
|
C6 |
N8 |
H9 |
108.281 |
| C7 |
C4 |
H10 |
110.258 |
|
C7 |
C4 |
H12 |
108.927 |
| C7 |
N8 |
H9 |
108.281 |
|
N8 |
C6 |
H14 |
107.141 |
| N8 |
C6 |
H16 |
108.331 |
|
N8 |
C7 |
H15 |
107.141 |
| N8 |
C7 |
H17 |
108.331 |
|
H10 |
C4 |
H12 |
106.982 |
| H11 |
C5 |
H13 |
106.982 |
|
H14 |
C6 |
H16 |
106.998 |
| H15 |
C7 |
H17 |
106.998 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
