return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6OS (Vinyl sulfoxide)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-628.119449
Energy at 298.15K-628.125752
Nuclear repulsion energy290.400047
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3357 3137 0.34      
2 A' 3278 3063 4.71      
3 A' 3243 3031 1.91      
4 A' 1693 1582 2.32      
5 A' 1450 1355 10.77      
6 A' 1310 1224 0.49      
7 A' 1130 1056 82.73      
8 A' 1033 965 9.92      
9 A' 1019 952 39.86      
10 A' 957 894 20.68      
11 A' 726 678 67.91      
12 A' 663 619 5.56      
13 A' 511 477 1.18      
14 A' 316 295 2.37      
15 A' 216 202 2.80      
16 A' 92 86 0.42      
17 A" 3357 3136 0.71      
18 A" 3274 3059 2.99      
19 A" 3243 3030 2.45      
20 A" 1685 1574 3.34      
21 A" 1446 1351 3.17      
22 A" 1292 1207 7.91      
23 A" 1023 956 5.09      
24 A" 1007 941 34.57      
25 A" 955 893 9.33      
26 A" 682 637 16.27      
27 A" 576 539 3.95      
28 A" 488 456 7.62      
29 A" 259 242 8.26      
30 A" 170 159 0.55      

Unscaled Zero Point Vibrational Energy (zpe) 20224.2 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 18897.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.16478 0.07580 0.06695

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.592 -0.511 0.000
O2 1.293 0.827 0.000
C3 -0.618 -0.488 1.316
C4 -0.618 -0.488 -1.316
C5 -0.618 0.527 2.178
C6 -0.618 0.527 -2.178
H7 -1.266 -1.353 1.379
H8 -1.266 -1.353 -1.379
H9 -1.307 0.559 3.008
H10 -1.307 0.559 -3.008
H11 0.081 1.341 2.044
H12 0.081 1.341 -2.044

Atom - Atom Distances (Å)
  S1 O2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12
S11.51071.78741.78742.69912.69912.46232.46233.71453.71452.80532.8053
O21.51072.66692.66692.91292.91293.63353.63353.98473.98472.43152.4315
C31.78742.66692.63211.33223.63901.08232.90392.10594.50192.08923.8894
C41.78742.66692.63213.63901.33222.90391.08234.50192.10593.88942.0892
C52.69912.91291.33223.63904.35682.14294.07561.07915.23171.08084.3569
C62.69912.91293.63901.33224.35684.07562.14295.23171.07914.35691.0808
H72.46233.63351.08232.90392.14294.07562.75872.51154.78553.08404.5598
H82.46233.63352.90391.08234.07562.14292.75874.78552.51154.55983.0840
H93.71453.98472.10594.50191.07915.23172.51154.78556.01501.86165.2973
H103.71453.98474.50192.10595.23171.07914.78552.51156.01505.29731.8616
H112.80532.43152.08923.88941.08084.35693.08404.55981.86165.29734.0890
H122.80532.43153.88942.08924.35691.08084.55983.08405.29731.86164.0890

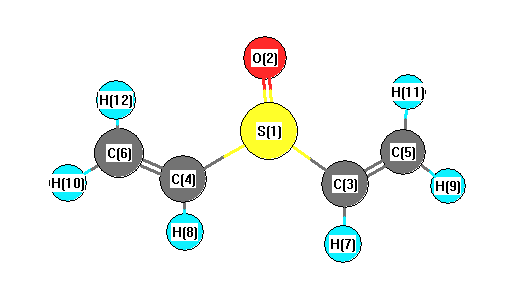
picture of Vinyl sulfoxide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C3 C5 119.088 S1 C3 H7 116.010
S1 C4 C6 119.088 S1 C4 H8 116.010
O2 S1 C3 107.628 O2 S1 C4 107.628
C3 S1 C4 94.831 C3 C5 H9 121.341
C3 C5 H11 119.587 C4 C6 H10 121.341
C4 C6 H12 119.587 C5 C3 H7 124.798
C6 C4 H8 124.798 H9 C5 H11 119.071
H10 C6 H12 119.071
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability