return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C9H20 (Nonane)

using model chemistry: PBE1PBE/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at PBE1PBE/6-31G**
 hartrees
Energy at 0K-354.575440
Energy at 298.15K-354.597095
HF Energy-354.575440
Nuclear repulsion energy445.217149
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3148 3000 42.80      
2 A1 3066 2922 35.13      
3 A1 3057 2914 170.81      
4 A1 3050 2907 20.60      
5 A1 3040 2897 1.32      
6 A1 3039 2896 1.00      
7 A1 1540 1468 11.16      
8 A1 1533 1461 0.45      
9 A1 1520 1449 0.05      
10 A1 1514 1443 0.00      
11 A1 1510 1439 0.00      
12 A1 1438 1371 3.64      
13 A1 1433 1366 0.03      
14 A1 1385 1320 0.01      
15 A1 1296 1235 0.01      
16 A1 1175 1120 0.74      
17 A1 1103 1051 0.58      
18 A1 1070 1019 0.32      
19 A1 1044 995 0.32      
20 A1 916 873 1.40      
21 A1 498 475 0.04      
22 A1 291 277 0.03      
23 A1 244 233 0.00      
24 A1 61 59 0.00      
25 A2 3141 2994 0.00      
26 A2 3103 2957 0.00      
27 A2 3084 2939 0.00      
28 A2 3067 2924 0.00      
29 A2 1526 1455 0.00      
30 A2 1358 1295 0.00      
31 A2 1341 1278 0.00      
32 A2 1322 1260 0.00      
33 A2 1243 1184 0.00      
34 A2 1054 1005 0.00      
35 A2 916 873 0.00      
36 A2 785 748 0.00      
37 A2 739 704 0.00      
38 A2 249 237 0.00      
39 A2 167 159 0.00      
40 A2 104 99 0.00      
41 A2 70 67 0.00      
42 B1 3141 2994 110.26      
43 B1 3110 2964 162.71      
44 B1 3094 2949 1.54      
45 B1 3074 2930 0.16      
46 B1 3065 2921 0.04      
47 B1 1526 1455 15.34      
48 B1 1355 1291 0.12      
49 B1 1346 1283 0.84      
50 B1 1283 1223 0.00      
51 B1 1219 1161 0.09      
52 B1 991 944 0.62      
53 B1 844 804 1.15      
54 B1 751 715 1.87      
55 B1 738 704 7.97      
56 B1 248 237 0.00      
57 B1 162 154 0.00      
58 B1 122 116 0.00      
59 B1 45 43 0.00      
60 B2 3147 3000 31.09      
61 B2 3065 2922 45.92      
62 B2 3056 2912 0.06      
63 B2 3043 2900 0.02      
64 B2 3038 2895 0.01      
65 B2 1538 1466 0.96      
66 B2 1527 1455 1.03      
67 B2 1515 1444 0.71      
68 B2 1510 1439 0.03      
69 B2 1439 1372 1.97      
70 B2 1435 1368 1.16      
71 B2 1417 1350 0.68      
72 B2 1342 1279 1.50      
73 B2 1250 1191 1.55      
74 B2 1125 1073 6.25      
75 B2 1100 1048 0.06      
76 B2 1093 1042 0.10      
77 B2 1029 981 0.04      
78 B2 917 874 3.38      
79 B2 460 438 0.11      
80 B2 398 379 0.03      
81 B2 159 152 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 60978.3 cm-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 58118.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at PBE1PBE/6-31G**
ABC
0.31248 0.01159 0.01139

See section I.F.4 to change rotational constant units
Geometric Data calculated at PBE1PBE/6-31G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.358
C2 0.000 1.276 -0.478
C3 0.000 -1.276 -0.478
C4 0.000 2.551 0.359
C5 0.000 -2.551 0.359
C6 0.000 3.828 -0.477
C7 0.000 -3.828 -0.477
C8 0.000 5.095 0.369
C9 0.000 -5.095 0.369
H10 0.878 0.000 1.021
H11 -0.878 0.000 1.021
H12 0.878 1.276 -1.141
H13 -0.878 1.276 -1.141
H14 -0.878 -1.276 -1.141
H15 0.878 -1.276 -1.141
H16 0.878 2.552 1.022
H17 -0.878 2.552 1.022
H18 -0.878 -2.552 1.022
H19 0.878 -2.552 1.022
H20 -0.878 3.826 -1.138
H21 0.878 3.826 -1.138
H22 0.878 -3.826 -1.138
H23 -0.878 -3.826 -1.138
H24 0.000 5.996 -0.254
H25 -0.884 5.137 1.017
H26 0.884 5.137 1.017
H27 0.000 -5.996 -0.254
H28 0.884 -5.137 1.017
H29 -0.884 -5.137 1.017

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 C9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20 H21 H22 H23 H24 H25 H26 H27 H28 H29
C11.52551.52552.55102.55103.91763.91765.09485.09481.10011.10012.15552.15552.15552.15552.77882.77882.77882.77884.20144.20144.20144.20146.02705.25435.25436.02705.25435.2543
C21.52552.55131.52543.91712.55195.10323.91206.42662.15552.15551.10011.10012.77852.77852.15602.15604.20344.20342.77712.77715.21895.21894.72554.23434.23437.27496.64416.6441
C31.52552.55133.91711.52545.10322.55196.42663.91202.15552.15552.77852.77851.10011.10014.20344.20342.15602.15605.21895.21892.77712.77717.27496.64416.64414.72554.23434.2343
C42.55101.52543.91715.10201.52566.43302.54387.64582.77802.77802.15542.15544.20284.20281.10021.10025.21995.21992.15362.15366.60926.60923.49882.81142.81148.56877.76707.7670
C52.55103.91711.52545.10206.43301.52567.64582.54382.77802.77804.20284.20282.15542.15545.21995.21991.10021.10026.60926.60922.15362.15368.56877.76707.76703.49882.81142.8114
C63.91762.55195.10321.52566.43307.65511.52378.96244.20294.20292.77922.77925.22075.22072.15502.15506.61136.61131.09911.09917.73247.73242.17972.17442.17449.82599.13149.1314
C73.91765.10322.55196.43301.52567.65518.96241.52374.20294.20295.22075.22072.77922.77926.61136.61132.15502.15507.73247.73241.09911.09919.82599.13149.13142.17972.17442.1744
C85.09483.91206.42662.54387.64581.52378.962410.18955.21085.21084.19964.19966.60576.60572.76842.76847.72427.72422.15702.15709.09019.09011.09551.09661.096611.108110.290610.2906
C95.09486.42663.91207.64582.54388.96241.523710.18955.21085.21086.60576.60574.19964.19967.72427.72422.76842.76849.09019.09012.15702.157011.108110.290610.29061.09551.09661.0966
H101.10012.15552.15552.77802.77804.20294.20295.21085.21081.75612.51033.06363.06362.51032.55163.09753.09752.55164.73154.39374.39374.73156.19245.43125.13746.19245.13745.4312
H111.10012.15552.15552.77802.77804.20294.20295.21085.21081.75613.06362.51032.51033.06363.09752.55162.55163.09754.39374.73154.73154.39376.19245.13745.43126.19245.43125.1374
H122.15551.10012.77852.15544.20282.77925.22074.19966.60572.51033.06361.75623.09752.55152.51083.06404.73384.39603.09652.55065.10215.39574.88234.76164.42357.37796.76646.9921
H132.15551.10012.77852.15544.20282.77925.22074.19966.60573.06362.51031.75622.55153.09753.06402.51084.39604.73382.55063.09655.39575.10214.88234.42354.76167.37796.99216.7664
H142.15552.77851.10014.20282.15545.22072.77926.60574.19963.06362.51033.09752.55151.75624.73384.39602.51083.06405.10215.39573.09652.55067.37796.76646.99214.88234.76164.4235
H152.15552.77851.10014.20282.15545.22072.77926.60574.19962.51033.06362.55153.09751.75624.39604.73383.06402.51085.39575.10212.55063.09657.37796.99216.76644.88234.42354.7616
H162.77882.15604.20341.10025.21992.15506.61132.76847.72422.55163.09752.51083.06404.73384.39601.75615.39695.10323.06152.50816.73386.95893.77623.12912.58588.68657.68907.8883
H172.77882.15604.20341.10025.21992.15506.61132.76847.72423.09752.55163.06402.51084.39604.73381.75615.10325.39692.50813.06156.95896.73383.77622.58583.12918.68657.88837.6890
H182.77884.20342.15605.21991.10026.61132.15507.72422.76843.09752.55164.73384.39602.51083.06405.39695.10321.75616.73386.95893.06152.50818.68657.68907.88833.77623.12912.5858
H192.77884.20342.15605.21991.10026.61132.15507.72422.76842.55163.09754.39604.73383.06402.51085.10325.39691.75616.95896.73382.50813.06158.68657.88837.68903.77622.58583.1291
H204.20142.77715.21892.15366.60921.09917.73242.15709.09014.73154.39373.09652.55065.10215.39573.06152.50816.73386.95891.75527.85147.65272.50192.52273.07699.90099.38609.2192
H214.20142.77715.21892.15366.60921.09917.73242.15709.09014.39374.73152.55063.09655.39575.10212.50813.06156.95896.73381.75527.65277.85142.50193.07692.52279.90099.21929.3860
H224.20145.21892.77716.60922.15367.73241.09919.09012.15704.39374.73155.10215.39573.09652.55066.73386.95893.06152.50817.85147.65271.75529.90099.38609.21922.50192.52273.0769
H234.20145.21892.77716.60922.15367.73241.09919.09012.15704.73154.39375.39575.10212.55063.09656.95896.73382.50813.06157.65277.85141.75529.90099.21929.38602.50193.07692.5227
H246.02704.72557.27493.49888.56872.17979.82591.095511.10816.19246.19244.88234.88237.37797.37793.77623.77628.68658.68652.50192.50199.90099.90091.76991.769911.991611.240311.2403
H255.25434.23436.64412.81147.76702.17449.13141.096610.29065.43125.13744.76164.42356.76646.99213.12912.58587.68907.88832.52273.07699.38609.21921.76991.768211.240310.425810.2747
H265.25434.23436.64412.81147.76702.17449.13141.096610.29065.13745.43124.42354.76166.99216.76642.58583.12917.88837.68903.07692.52279.21929.38601.76991.768211.240310.274710.4258
H276.02707.27494.72558.56873.49889.82592.179711.10811.09556.19246.19247.37797.37794.88234.88238.68658.68653.77623.77629.90099.90092.50192.501911.991611.240311.24031.76991.7699
H285.25436.64414.23437.76702.81149.13142.174410.29061.09665.13745.43126.76646.99214.76164.42357.68907.88833.12912.58589.38609.21922.52273.076911.240310.425810.27471.76991.7682
H295.25436.64414.23437.76702.81149.13142.174410.29061.09665.43125.13746.99216.76644.42354.76167.88837.68902.58583.12919.21929.38603.07692.522711.240310.274710.42581.76991.7682

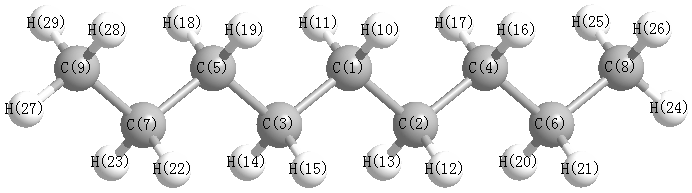
picture of Nonane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 113.474 C1 C2 H12 109.293
C1 C2 H13 109.293 C1 C3 C5 113.474
C1 C3 H14 109.293 C1 C3 H15 109.293
C2 C1 C3 113.490 C2 C1 H10 109.290
C2 C1 H11 109.290 C2 C4 C6 113.527
C2 C4 H16 109.332 C2 C4 H17 109.332
C3 C1 H10 109.290 C3 C1 H11 109.290
C3 C5 C7 113.527 C3 C5 H18 109.332
C3 C5 H19 109.332 C4 C2 H12 109.293
C4 C2 H13 109.293 C4 C6 C8 113.065
C4 C6 H20 109.192 C4 C6 H21 109.192
C5 C3 H14 109.293 C5 C3 H15 109.293
C5 C7 C9 113.065 C5 C7 H22 109.192
C5 C7 H23 109.192 C6 C4 H16 109.236
C6 C4 H17 109.236 C6 C8 H24 111.604
C6 C8 H25 111.110 C6 C8 H26 111.110
C7 C5 H14 96.552 C7 C5 H15 96.552
C7 C9 H27 111.604 C7 C9 H28 111.110
C7 C9 H29 111.110 C8 C6 H20 109.586
C8 C6 H21 109.586 C9 C7 H22 109.586
C9 C7 H23 109.586 H10 C1 H11 105.910
H12 C2 H13 105.918 H14 C3 H15 105.918
H16 C4 H17 105.896 H18 C5 H19 105.896
H20 C6 H21 105.971 H22 C7 H23 105.971
H24 C8 H25 107.686 H24 C8 H26 107.686
H25 C8 H26 107.454 H27 C9 H28 107.686
H27 C9 H29 107.686 H28 C9 H29 107.454
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G** Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.317      
2 C -0.316      
3 C -0.316      
4 C -0.310      
5 C -0.310      
6 C -0.316      
7 C -0.316      
8 C -0.520      
9 C -0.520      
10 H 0.158      
11 H 0.158      
12 H 0.158      
13 H 0.158      
14 H 0.158      
15 H 0.158      
16 H 0.158      
17 H 0.158      
18 H 0.158      
19 H 0.158      
20 H 0.162      
21 H 0.162      
22 H 0.162      
23 H 0.162      
24 H 0.170      
25 H 0.168      
26 H 0.168      
27 H 0.170      
28 H 0.168      
29 H 0.168      


Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section VII.A.3)
  x y z Total
  0.000 0.000 -0.054 0.054
CHELPG        
AIM        
ESP        


Electric Quadrupole moment
Quadrupole components in D Å
Primitive
 xyz
x -59.890 0.000 0.000
y 0.000 -62.352 0.000
z 0.000 0.000 -61.526
Traceless
 xyz
x 2.049 0.000 0.000
y 0.000 -1.644 0.000
z 0.000 0.000 -0.405
Polar
3z2-r2-0.811
x2-y22.462
xy0.000
xz0.000
yz0.000


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 12.562 0.000 0.000
y 0.000 18.175 0.000
z 0.000 0.000 12.917


<r2> (average value of r2) Å2
<r2> 936.001
(<r2>)1/2 30.594