return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6S (Thiophene, 2,3-dihydro-)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-553.168268
Energy at 298.15K-553.175268
HF Energy-552.462307
Nuclear repulsion energy217.675776
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3321 3104 7.24      
2 A 3293 3077 0.74      
3 A 3238 3026 8.90      
4 A 3188 2978 20.34      
5 A 3156 2949 16.70      
6 A 3106 2902 30.36      
7 A 1656 1547 8.34      
8 A 1544 1443 0.56      
9 A 1526 1426 1.82      
10 A 1385 1294 0.52      
11 A 1372 1282 4.17      
12 A 1326 1239 15.80      
13 A 1241 1159 1.76      
14 A 1185 1108 0.28      
15 A 1151 1075 1.43      
16 A 1056 986 4.75      
17 A 1038 970 2.30      
18 A 957 894 2.11      
19 A 927 866 1.16      
20 A 894 835 1.30      
21 A 822 768 20.51      
22 A 720 673 6.13      
23 A 701 655 27.30      
24 A 665 622 19.37      
25 A 510 476 0.65      
26 A 412 385 2.52      
27 A 190 178 1.22      

Unscaled Zero Point Vibrational Energy (zpe) 20289.8 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 18958.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.23142 0.15686 0.09914

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.065 -1.243 0.234
H2 -0.079 -1.451 1.303
H3 0.139 -2.164 -0.306
C4 -1.371 -0.576 -0.196
H5 -2.215 -1.000 0.348
H6 -1.551 -0.738 -1.263
C7 0.106 1.272 0.086
H8 0.471 2.282 0.187
C9 -1.180 0.894 0.066
H10 -2.003 1.589 0.136
S11 1.269 -0.037 -0.097

Atom - Atom Distances (Å)
  C1 H2 H3 C4 H5 H6 C7 H8 C9 H10 S11
C11.08901.08651.52892.16722.16942.52553.56602.41693.43381.8281
H21.08901.77322.16372.38323.04302.98793.93442.87083.78222.4027
H31.08651.77322.19432.70592.40963.45864.48573.35114.34392.4176
C41.52892.16372.19431.08951.09452.38223.42141.50502.27942.6962
H52.16722.38322.70591.08951.76183.25814.24382.17652.60613.6416
H62.16943.04302.40961.09451.76182.93323.91262.13672.75243.1308
C72.52552.98793.45862.38223.25812.93321.07861.34052.13331.7609
H83.56603.93444.48573.42144.24383.91261.07862.16032.56982.4692
C92.41692.87083.35111.50502.17652.13671.34052.16031.07962.6253
H103.43383.78224.34392.27942.60612.75242.13332.56981.07963.6615
S111.82812.40272.41762.69623.64163.13081.76092.46922.62533.6615

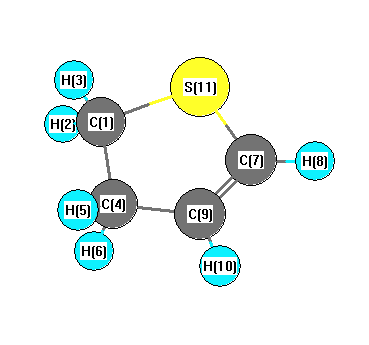
picture of Thiophene, 2,3-dihydro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H5 110.598 C1 C4 H6 110.474
C1 C4 C9 105.617 C1 S11 C7 89.424
H2 C1 H3 109.189 H2 C1 C4 110.351
H2 C1 S11 108.224 H3 C1 C4 112.974
H3 C1 S11 109.444 C4 C1 S11 106.526
C4 C9 C7 113.562 C4 C9 H10 122.908
H5 C4 H6 107.548 H5 C4 C9 113.071
H6 C4 C9 109.553 C7 C9 H10 123.291
H8 C7 C9 126.164 H8 C7 S11 118.842
C9 C7 S11 114.987
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability