return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6O (Cyclobutanone)

using model chemistry: QCISD/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/6-31G**
 hartrees
Energy at 0K-230.572217
Energy at 298.15K-230.579062
HF Energy-229.805882
Nuclear repulsion energy167.244930
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3215 3027 33.27      
2 A' 3193 3006 3.42      
3 A' 3152 2968 22.07      
4 A' 3135 2951 8.82      
5 A' 1894 1783 224.74      
6 A' 1556 1465 0.05      
7 A' 1510 1422 1.48      
8 A' 1289 1214 2.75      
9 A' 1252 1178 0.29      
10 A' 1129 1062 0.37      
11 A' 998 940 1.03      
12 A' 882 831 0.74      
13 A' 754 710 2.32      
14 A' 686 646 2.10      
15 A' 404 380 1.76      
16 A' 79 74 3.38      
17 A" 3199 3011 4.40      
18 A" 3132 2948 15.96      
19 A" 1498 1411 10.89      
20 A" 1313 1236 8.87      
21 A" 1267 1193 3.21      
22 A" 1224 1152 1.46      
23 A" 1129 1063 74.68      
24 A" 980 922 0.25      
25 A" 955 899 0.18      
26 A" 678 638 0.01      
27 A" 463 436 4.72      

Unscaled Zero Point Vibrational Energy (zpe) 20482.3 cm-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 19282.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G**
ABC
0.35887 0.15988 0.11896

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 -0.158 1.874 0.000
C2 0.000 0.674 0.000
C3 0.101 -0.386 1.108
C4 0.101 -0.386 -1.108
C5 -0.069 -1.466 0.000
H6 -1.064 -1.909 0.000
H7 0.671 -2.264 0.000
H8 1.088 -0.385 1.573
H9 1.088 -0.385 -1.573
H10 -0.657 -0.331 1.889
H11 -0.657 -0.331 -1.889

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11
O11.20982.53002.53003.34083.88924.21993.02183.02182.94592.9459
C21.20981.53681.53682.14132.79333.01392.18672.18672.23842.2384
C32.53001.53682.21531.55602.21362.25351.09182.85671.08973.0913
C42.53001.53682.21531.55602.21362.25352.85671.09183.09131.0897
C53.34082.14131.55601.55601.08921.08812.23152.23152.28062.2806
H63.88922.79332.21362.21361.08921.77083.07013.07012.49422.4942
H74.21993.01392.25352.25351.08811.77082.48532.48533.01113.0111
H83.02182.18671.09182.85672.23153.07012.48533.14561.77483.8772
H93.02182.18672.85671.09182.23153.07012.48533.14563.87721.7748
H102.94592.23841.08973.09132.28062.49423.01111.77483.87723.7776
H112.94592.23843.09131.08972.28062.49423.01113.87721.77483.7776

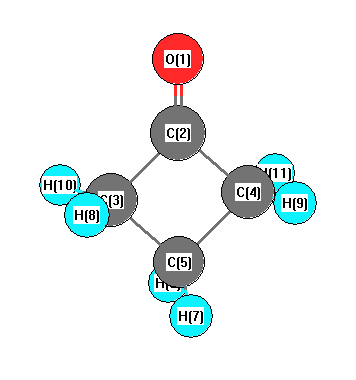
picture of Cyclobutanone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C3 133.846 O1 C2 C4 133.846
C2 C3 C5 87.629 C2 C3 H8 111.458
C2 C3 H10 115.869 C2 C4 C5 87.629
C2 C4 H9 111.458 C2 C4 H11 115.869
C3 C2 C4 92.234 C3 C5 C4 90.772
C3 C5 H6 112.418 C3 C5 H7 115.784
C4 C5 H6 112.418 C4 C5 H7 115.784
C5 C3 H8 113.712 C5 C3 H10 118.014
C5 C4 H9 113.712 C5 C4 H11 118.014
H6 C5 H7 108.843 H8 C3 H10 108.888
H9 C4 H11 108.888
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability