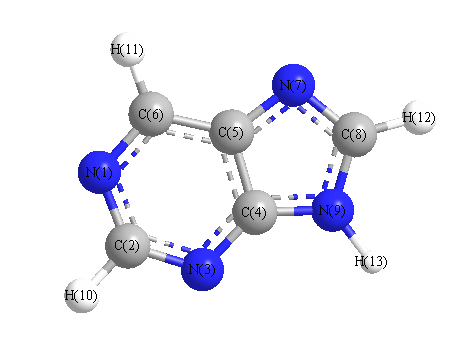
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3707 |
3537 |
98.64 |
|
|
|
| 2 |
A' |
3278 |
3128 |
0.59 |
|
|
|
| 3 |
A' |
3212 |
3065 |
16.24 |
|
|
|
| 4 |
A' |
3203 |
3057 |
15.43 |
|
|
|
| 5 |
A' |
1695 |
1618 |
65.81 |
|
|
|
| 6 |
A' |
1653 |
1578 |
80.64 |
|
|
|
| 7 |
A' |
1563 |
1492 |
29.14 |
|
|
|
| 8 |
A' |
1514 |
1445 |
4.09 |
|
|
|
| 9 |
A' |
1460 |
1393 |
67.79 |
|
|
|
| 10 |
A' |
1440 |
1374 |
20.35 |
|
|
|
| 11 |
A' |
1393 |
1329 |
63.79 |
|
|
|
| 12 |
A' |
1342 |
1281 |
18.62 |
|
|
|
| 13 |
A' |
1334 |
1273 |
11.10 |
|
|
|
| 14 |
A' |
1302 |
1243 |
29.92 |
|
|
|
| 15 |
A' |
1222 |
1166 |
7.40 |
|
|
|
| 16 |
A' |
1155 |
1102 |
6.08 |
|
|
|
| 17 |
A' |
1102 |
1052 |
13.28 |
|
|
|
| 18 |
A' |
952 |
909 |
1.47 |
|
|
|
| 19 |
A' |
920 |
878 |
12.19 |
|
|
|
| 20 |
A' |
818 |
781 |
14.82 |
|
|
|
| 21 |
A' |
664 |
633 |
0.46 |
|
|
|
| 22 |
A' |
573 |
547 |
4.08 |
|
|
|
| 23 |
A' |
448 |
428 |
12.85 |
|
|
|
| 24 |
A" |
1000 |
954 |
0.46 |
|
|
|
| 25 |
A" |
947 |
903 |
9.16 |
|
|
|
| 26 |
A" |
889 |
848 |
5.57 |
|
|
|
| 27 |
A" |
815 |
778 |
9.03 |
|
|
|
| 28 |
A" |
671 |
641 |
6.38 |
|
|
|
| 29 |
A" |
627 |
598 |
26.19 |
|
|
|
| 30 |
A" |
546 |
521 |
109.67 |
|
|
|
| 31 |
A" |
420 |
401 |
2.64 |
|
|
|
| 32 |
A" |
249 |
238 |
0.37 |
|
|
|
| 33 |
A" |
233 |
222 |
4.35 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21172.5 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 20204.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.447 |
|
|
|
| 2 |
C |
0.207 |
|
|
|
| 3 |
N |
-0.494 |
|
|
|
| 4 |
C |
0.531 |
|
|
|
| 5 |
C |
0.173 |
|
|
|
| 6 |
C |
0.098 |
|
|
|
| 7 |
N |
-0.504 |
|
|
|
| 8 |
C |
0.277 |
|
|
|
| 9 |
N |
-0.608 |
|
|
|
| 10 |
H |
0.144 |
|
|
|
| 11 |
H |
0.155 |
|
|
|
| 12 |
H |
0.164 |
|
|
|
| 13 |
H |
0.304 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.216 |
2.984 |
0.000 |
3.717 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.584 |
6.167 |
0.000 |
| y |
6.167 |
-46.402 |
0.000 |
| z |
0.000 |
0.000 |
-51.197 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.784 |
6.167 |
0.000 |
| y |
6.167 |
3.989 |
0.000 |
| z |
0.000 |
0.000 |
-3.205 |
|
| Polar |
|---|
| 3z2-r2 | -6.409 |
|---|
| x2-y2 | -3.182 |
|---|
| xy | 6.167 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.624 |
1.343 |
0.000 |
| y |
1.343 |
11.348 |
0.000 |
| z |
0.000 |
0.000 |
4.252 |
<r2> (average value of r
2) Å
2
| <r2> |
252.498 |
| (<r2>)1/2 |
15.890 |