return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

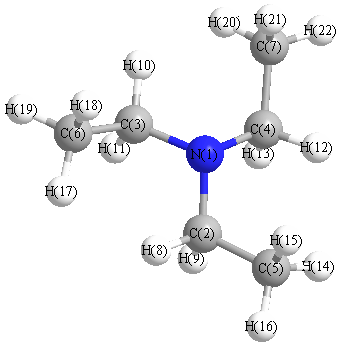
All results from a given calculation for C6H15N (triethylamine)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C3 1A
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-291.486209
Energy at 298.15K-291.503611
HF Energy-290.394706
Nuclear repulsion energy337.829938
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3244 3031 36.41      
2 A 3226 3015 5.68      
3 A 3181 2973 1.79      
4 A 3135 2929 5.62      
5 A 3034 2835 163.25      
6 A 1573 1470 7.12      
7 A 1555 1453 1.17      
8 A 1546 1445 0.72      
9 A 1472 1376 14.65      
10 A 1448 1353 0.49      
11 A 1355 1266 3.41      
12 A 1214 1135 9.41      
13 A 1127 1053 1.56      
14 A 1067 997 7.20      
15 A 839 784 0.13      
16 A 775 724 10.85      
17 A 454 424 3.39      
18 A 318 297 0.79      
19 A 203 190 0.63      
20 A 104 97 0.01      
21 E 3245 3032 21.26      
22 E 3228 3017 42.52      
23 E 3182 2973 28.15      
24 E 3135 2929 22.98      
25 E 3027 2828 12.80      
26 E 1570 1467 3.03      
26 E 1570 1467 3.03      
27 E 1561 1459 1.72      
27 E 1561 1459 1.72      
28 E 1546 1445 1.22      
28 E 1546 1445 1.22      
29 E 1470 1373 17.53      
30 E 1430 1337 9.55      
30 E 1430 1337 9.55      
31 E 1366 1277 19.47      
32 E 1278 1194 17.88      
33 E 1143 1068 8.94      
33 E 1143 1068 8.94      
34 E 1128 1054 18.55      
35 E 962 899 1.81      
35 E 962 899 1.81      
36 E 824 770 2.27      
36 E 824 770 2.27      
37 E 485 453 2.41      
37 E 485 453 2.41      
38 E 327 305 0.44      
38 E 327 305 0.44      
39 E 230 215 0.03      
39 E 230 215 0.03      
40 E 96 90 0.21      
40 E 96 90 0.21      

Unscaled Zero Point Vibrational Energy (zpe) 46667.7 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 43606.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.07776 0.07776 0.04465

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is C3

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.000 0.000 0.013
C2 0.334 1.351 -0.445
C3 1.003 -0.964 -0.445
C4 -1.336 -0.386 -0.445
C5 -0.293 2.423 0.430
C6 2.245 -0.958 0.430
C7 -1.952 -1.466 0.430
H8 1.416 1.462 -0.403
H9 0.048 1.494 -1.498
H10 0.559 -1.957 -0.403
H11 1.270 -0.788 -1.498
H12 -1.974 0.495 -0.403
H13 -1.318 -0.706 -1.498
H14 -1.380 2.377 0.415
H15 0.036 2.296 1.458
H16 0.000 3.415 0.087
H17 2.749 0.006 0.415
H18 1.970 -1.179 1.458
H19 2.957 -1.707 0.087
H20 -1.369 -2.383 0.415
H21 -2.006 -1.117 1.458
H22 -2.957 -1.707 0.087

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20 H21 H22
N11.46441.46441.46442.47652.47652.47652.07742.12582.07742.12582.07742.12582.77792.71303.41532.77792.71303.41532.77792.71303.4153
C21.46442.40962.40961.51943.12233.73101.08891.10143.31592.56202.46182.84032.17442.14512.15712.89463.56344.06404.19293.89634.5233
C31.46442.40962.40963.73101.51943.12232.46182.84031.08891.10143.31592.56204.19293.89634.52332.17442.14512.15712.89463.56344.0640
C41.46442.40962.40963.12233.73101.51943.31592.56202.46182.84031.08891.10142.89463.56344.06404.19293.89634.52332.17442.14512.1571
C52.47651.51943.73103.12234.22814.22812.13022.16754.53984.05952.69043.81601.08761.08701.08903.88524.37695.26734.92584.06494.9271
C62.47653.12231.51943.73104.22814.22812.69043.81602.13022.16754.53984.05954.92584.06494.92711.08761.08701.08903.88524.37695.2673
C72.47653.73103.12231.51944.22814.22814.53984.05952.69043.81602.13022.16753.88524.37695.26734.92584.06494.92711.08761.08701.0890
H82.07741.08892.46183.31592.13022.69044.53981.75333.52562.50773.52563.65743.05302.46162.46062.13653.27803.55824.81814.67155.4229
H92.12581.10142.84032.56202.16753.81604.05951.75333.65742.58962.50772.58962.54543.06312.49043.62924.42534.60734.55044.44704.6681
H102.07743.31591.08892.46184.53982.13022.69043.52563.65741.75333.52562.50774.81814.67155.42293.05302.46162.46062.13653.27813.5582
H112.12582.56201.10142.84034.05952.16753.81602.50772.58961.75333.65742.58964.55044.44704.66812.54543.06312.49043.62924.42534.6073
H122.07742.46183.31591.08892.69044.53982.13023.52562.50773.52563.65741.75332.13653.27803.55824.81814.67155.42293.05302.46162.4607
H132.12582.84032.56201.10143.81604.05952.16753.65742.58962.50772.58961.75333.62924.42534.60734.55044.44704.66812.54543.06312.4904
H142.77792.17444.19292.89461.08764.92583.88523.05302.54544.81814.55042.13653.62921.76021.75724.76064.99555.96644.76063.69954.3907
H152.71302.14513.89633.56341.08704.06494.37692.46163.06314.67154.44703.27804.42531.76021.77013.69953.97685.14164.99553.97685.1829
H163.41532.15714.52334.06401.08904.92715.26732.46062.49045.42294.66813.55824.60731.75721.77014.39075.18295.91415.96645.14165.9141
H172.77792.89462.17444.19293.88521.08764.92582.13653.62923.05302.54544.81814.55044.76063.69954.39071.76021.75724.76064.99565.9664
H182.71303.56342.14513.89634.37691.08704.06493.27804.42532.46163.06314.67154.44704.99553.97685.18291.76021.77013.69953.97685.1416
H193.41534.06402.15714.52335.26731.08904.92713.55824.60732.46062.49045.42294.66815.96645.14165.91411.75721.77014.39075.18295.9141
H202.77794.19292.89462.17444.92583.88521.08764.81814.55042.13653.62923.05302.54544.76064.99555.96644.76063.69954.39071.76021.7572
H212.71303.89633.56342.14514.06494.37691.08704.67154.44703.27814.42532.46163.06313.69953.97685.14164.99563.97685.18291.76021.7701
H223.41534.52334.06402.15714.92715.26731.08905.42294.66813.55824.60732.46072.49044.39075.18295.91415.96645.14165.91411.75721.7701

picture of triethylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C5 112.180 N1 C2 H8 107.995
N1 C2 H9 111.104 N1 C3 C6 112.180
N1 C3 H10 107.995 N1 C3 H11 111.104
N1 C4 C7 112.180 N1 C4 H12 107.995
N1 C4 H13 111.104 C2 N1 C3 110.716
C2 N1 C4 110.716 C2 C5 H14 111.975
C2 C5 H15 109.661 C2 C5 H16 110.491
C3 N1 C4 110.716 C3 C6 H17 111.976
C3 C6 H18 109.661 C3 C6 H19 110.491
C4 C7 H20 111.975 C4 C7 H21 109.661
C4 C7 H22 110.491 C5 C2 H8 108.378
C5 C2 H9 110.584 C6 C3 H10 108.378
C6 C3 H11 110.584 C7 C4 H12 108.378
C7 C4 H13 110.584 H8 C2 H9 106.355
H10 C3 H11 106.355 H12 C4 H13 106.355
H14 C5 H15 108.086 H14 C5 H16 107.663
H15 C5 H16 108.873 H17 C6 H18 108.086
H17 C6 H19 107.663 H18 C6 H19 108.873
H20 C7 H21 108.086 H20 C7 H22 107.663
H21 C7 H22 108.873
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability