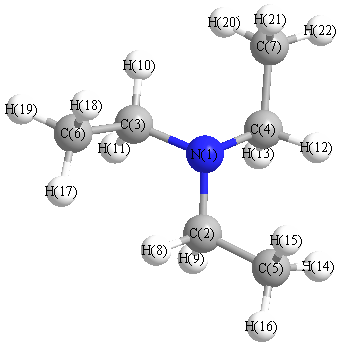
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3165 |
3016 |
49.42 |
|
|
|
| 2 |
A |
3148 |
3001 |
0.40 |
|
|
|
| 3 |
A |
3115 |
2969 |
0.25 |
|
|
|
| 4 |
A |
3075 |
2931 |
7.78 |
|
|
|
| 5 |
A |
2962 |
2823 |
190.73 |
|
|
|
| 6 |
A |
1542 |
1470 |
11.00 |
|
|
|
| 7 |
A |
1518 |
1447 |
1.92 |
|
|
|
| 8 |
A |
1512 |
1441 |
0.73 |
|
|
|
| 9 |
A |
1448 |
1380 |
15.59 |
|
|
|
| 10 |
A |
1422 |
1355 |
0.21 |
|
|
|
| 11 |
A |
1336 |
1274 |
3.17 |
|
|
|
| 12 |
A |
1187 |
1131 |
11.38 |
|
|
|
| 13 |
A |
1106 |
1054 |
2.57 |
|
|
|
| 14 |
A |
1056 |
1006 |
8.03 |
|
|
|
| 15 |
A |
821 |
783 |
0.01 |
|
|
|
| 16 |
A |
769 |
733 |
10.89 |
|
|
|
| 17 |
A |
439 |
418 |
3.34 |
|
|
|
| 18 |
A |
310 |
295 |
1.16 |
|
|
|
| 19 |
A |
191 |
182 |
0.71 |
|
|
|
| 20 |
A |
92 |
88 |
0.01 |
|
|
|
| 21 |
E |
3164 |
3016 |
17.08 |
|
|
|
| 21 |
E |
3164 |
3016 |
17.08 |
|
|
|
| 22 |
E |
3152 |
3004 |
61.96 |
|
|
|
| 22 |
E |
3152 |
3004 |
61.96 |
|
|
|
| 23 |
E |
3115 |
2969 |
22.23 |
|
|
|
| 23 |
E |
3115 |
2969 |
22.23 |
|
|
|
| 24 |
E |
3075 |
2930 |
25.60 |
|
|
|
| 24 |
E |
3075 |
2930 |
25.60 |
|
|
|
| 25 |
E |
2951 |
2812 |
17.26 |
|
|
|
| 25 |
E |
2951 |
2812 |
17.26 |
|
|
|
| 26 |
E |
1537 |
1465 |
5.35 |
|
|
|
| 26 |
E |
1537 |
1465 |
5.35 |
|
|
|
| 27 |
E |
1527 |
1456 |
1.86 |
|
|
|
| 27 |
E |
1527 |
1456 |
1.86 |
|
|
|
| 28 |
E |
1512 |
1441 |
1.68 |
|
|
|
| 28 |
E |
1512 |
1441 |
1.68 |
|
|
|
| 29 |
E |
1444 |
1376 |
26.04 |
|
|
|
| 29 |
E |
1444 |
1376 |
26.03 |
|
|
|
| 30 |
E |
1411 |
1345 |
3.89 |
|
|
|
| 30 |
E |
1411 |
1345 |
3.89 |
|
|
|
| 31 |
E |
1349 |
1285 |
26.15 |
|
|
|
| 31 |
E |
1349 |
1285 |
26.15 |
|
|
|
| 32 |
E |
1260 |
1201 |
22.93 |
|
|
|
| 32 |
E |
1260 |
1201 |
22.93 |
|
|
|
| 33 |
E |
1124 |
1071 |
19.56 |
|
|
|
| 33 |
E |
1124 |
1071 |
19.56 |
|
|
|
| 34 |
E |
1114 |
1062 |
9.18 |
|
|
|
| 34 |
E |
1114 |
1062 |
9.18 |
|
|
|
| 35 |
E |
949 |
905 |
0.91 |
|
|
|
| 35 |
E |
949 |
905 |
0.91 |
|
|
|
| 36 |
E |
808 |
770 |
3.55 |
|
|
|
| 36 |
E |
808 |
770 |
3.55 |
|
|
|
| 37 |
E |
479 |
457 |
2.65 |
|
|
|
| 37 |
E |
479 |
457 |
2.65 |
|
|
|
| 38 |
E |
315 |
300 |
0.35 |
|
|
|
| 38 |
E |
315 |
300 |
0.35 |
|
|
|
| 39 |
E |
219 |
209 |
0.03 |
|
|
|
| 39 |
E |
219 |
209 |
0.03 |
|
|
|
| 40 |
E |
87 |
83 |
0.22 |
|
|
|
| 40 |
E |
87 |
83 |
0.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 45697.0 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 43553.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.384 |
|
|
|
| 2 |
C |
-0.184 |
|
|
|
| 3 |
C |
-0.184 |
|
|
|
| 4 |
C |
-0.184 |
|
|
|
| 5 |
C |
-0.517 |
|
|
|
| 6 |
C |
-0.517 |
|
|
|
| 7 |
C |
-0.517 |
|
|
|
| 8 |
H |
0.168 |
|
|
|
| 9 |
H |
0.141 |
|
|
|
| 10 |
H |
0.168 |
|
|
|
| 11 |
H |
0.141 |
|
|
|
| 12 |
H |
0.168 |
|
|
|
| 13 |
H |
0.141 |
|
|
|
| 14 |
H |
0.171 |
|
|
|
| 15 |
H |
0.183 |
|
|
|
| 16 |
H |
0.167 |
|
|
|
| 17 |
H |
0.171 |
|
|
|
| 18 |
H |
0.183 |
|
|
|
| 19 |
H |
0.167 |
|
|
|
| 20 |
H |
0.171 |
|
|
|
| 21 |
H |
0.183 |
|
|
|
| 22 |
H |
0.167 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.551 |
0.551 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.768 |
0.000 |
0.000 |
| y |
0.000 |
-45.768 |
0.000 |
| z |
0.000 |
0.000 |
-48.151 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.191 |
0.000 |
0.000 |
| y |
0.000 |
1.191 |
0.000 |
| z |
0.000 |
0.000 |
-2.382 |
|
| Polar |
|---|
| 3z2-r2 | -4.765 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.367 |
-0.000 |
0.000 |
| y |
-0.000 |
11.367 |
0.000 |
| z |
0.000 |
0.000 |
9.551 |
<r2> (average value of r
2) Å
2
| <r2> |
286.189 |
| (<r2>)1/2 |
16.917 |