return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H15N (triethylamine)

using model chemistry: CCD/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C3 1A
Energy calculated at CCD/6-31G**
 hartrees
Energy at 0K-291.531650
Energy at 298.15K-291.549040
HF Energy-290.394799
Nuclear repulsion energy336.837595
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3216 3015 47.22      
2 A 3201 3001 6.96      
3 A 3168 2971 1.09      
4 A 3121 2926 6.33      
5 A 3034 2844 163.16      
6 A 1578 1479 5.74      
7 A 1556 1459 0.18      
8 A 1550 1454 1.59      
9 A 1485 1392 12.82      
10 A 1458 1367 0.28      
11 A 1360 1275 3.90      
12 A 1219 1143 9.45      
13 A 1129 1059 2.10      
14 A 1069 1002 5.83      
15 A 839 787 0.37      
16 A 778 729 9.96      
17 A 455 427 3.63      
18 A 314 295 0.78      
19 A 202 189 0.65      
20 A 104 98 0.01      
21 E 3217 3016 31.99      
22 E 3203 3003 50.60      
23 E 3167 2970 27.09      
24 E 3121 2926 25.55      
25 E 3025 2836 10.70      
26 E 1575 1476 2.29      
26 E 1575 1476 2.29      
27 E 1562 1465 0.68      
27 E 1562 1465 0.68      
28 E 1550 1453 1.83      
28 E 1550 1453 1.83      
29 E 1481 1389 19.54      
30 E 1444 1353 10.73      
31 E 1374 1288 19.15      
32 E 1287 1207 15.68      
33 E 1145 1073 11.39      
34 E 1132 1061 12.90      
35 E 963 903 1.76      
35 E 963 903 1.76      
36 E 824 773 1.89      
36 E 824 773 1.89      
37 E 486 455 2.05      
37 E 486 455 2.05      
38 E 327 306 0.45      
38 E 327 306 0.45      
39 E 226 212 0.02      
39 E 226 212 0.02      
40 E 94 88 0.20      
40 E 94 88 0.20      

Unscaled Zero Point Vibrational Energy (zpe) 46619.5 cm-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 43710.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/6-31G**
ABC
0.07720 0.07720 0.04444

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/6-31G**

Point Group is C3

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.000 0.000 0.000
C2 -0.329 1.357 0.451
C3 -1.011 -0.963 0.451
C4 1.340 -0.394 0.451
C5 0.291 2.429 -0.439
C6 -2.249 -0.962 -0.439
C7 1.958 -1.466 -0.439
H8 -1.413 1.471 0.419
H9 -0.033 1.510 1.501
H10 -0.567 -1.959 0.419
H11 -1.291 -0.784 1.501
H12 1.980 0.488 0.419
H13 1.324 -0.726 1.501
H14 1.380 2.383 -0.435
H15 -0.048 2.297 -1.465
H16 0.000 3.423 -0.097
H17 -2.753 0.004 -0.435
H18 -1.965 -1.190 -1.465
H19 -2.964 -1.711 -0.097
H20 1.373 -2.386 -0.435
H21 2.013 -1.107 -1.465
H22 2.964 -1.711 -0.097

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20 H21 H22
N11.46741.46741.46742.48512.48512.48512.08202.12932.08202.12932.08202.12932.78732.72493.42422.78732.72493.42422.78732.72493.4242
C21.46742.41882.41881.52443.13963.74071.09011.10183.32482.57132.46752.85962.18072.15242.16222.91433.58304.08184.20653.90234.5345
C31.46742.41882.41883.74071.52443.13962.46752.85961.09011.10183.32482.57134.20653.90234.53452.18072.15242.16222.91433.58304.0818
C41.46742.41882.41883.13963.74071.52443.32482.57132.46752.85961.09011.10182.91433.58304.08184.20653.90234.53452.18072.15242.1622
C52.48511.52443.74073.13964.23674.23672.13442.17064.55234.07242.71203.84511.08971.08911.09093.89214.38655.27784.93504.06424.9400
C62.48513.13961.52443.74074.23674.23672.71203.84512.13442.17064.55234.07244.93504.06424.94001.08971.08911.09093.89214.38655.2778
C72.48513.74073.13961.52444.23674.23674.55234.07242.71203.84512.13442.17063.89214.38655.27784.93504.06424.94001.08971.08911.0909
H82.08201.09012.46753.32482.13442.71204.55231.75403.53242.50363.53243.67283.05902.46842.46412.16303.30713.57784.83404.68305.4361
H92.12931.10182.85962.57132.17063.84514.07241.75403.67282.61602.50362.61602.55043.06882.49263.66284.45234.63934.57224.45334.6812
H102.08203.32481.09012.46754.55232.13442.71203.53243.67281.75403.53242.50364.83404.68305.43613.05902.46842.46412.16303.30713.5778
H112.12932.57131.10182.85964.07242.17063.84512.50362.61601.75403.67282.61604.57224.45334.68122.55043.06882.49263.66284.45234.6393
H122.08202.46753.32481.09012.71204.55232.13443.53242.50363.53243.67281.75402.16303.30713.57784.83404.68305.43613.05902.46842.4641
H132.12932.85962.57131.10183.84514.07242.17063.67282.61602.50362.61601.75403.66284.45234.63934.57224.45334.68122.55043.06882.4926
H142.78732.18074.20652.91431.08974.93503.89213.05902.55044.83404.57222.16303.66281.76301.76084.76865.00145.97874.76863.69284.4029
H152.72492.15243.90233.58301.08914.06424.38652.46843.06884.68304.45333.30714.45231.76301.77273.69283.97935.14225.00143.97935.1977
H163.42422.16224.53454.08181.09094.94005.27782.46412.49265.43614.68123.57784.63931.76081.77274.40295.19775.92865.97875.14225.9286
H172.78732.91432.18074.20653.89211.08974.93502.16303.66283.05902.55044.83404.57224.76863.69284.40291.76301.76084.76865.00145.9787
H182.72493.58302.15243.90234.38651.08914.06423.30714.45232.46843.06884.68304.45335.00143.97935.19771.76301.77273.69283.97935.1422
H193.42424.08182.16224.53455.27781.09094.94003.57784.63932.46412.49265.43614.68125.97875.14225.92861.76081.77274.40295.19775.9286
H202.78734.20652.91432.18074.93503.89211.08974.83404.57222.16303.66283.05902.55044.76865.00145.97874.76863.69284.40291.76301.7608
H212.72493.90233.58302.15244.06424.38651.08914.68304.45333.30714.45232.46843.06883.69283.97935.14225.00143.97935.19771.76301.7727
H223.42424.53454.08182.16224.94005.27781.09095.43614.68123.57784.63932.46412.49264.40295.19775.92865.97875.14225.92861.76081.7727

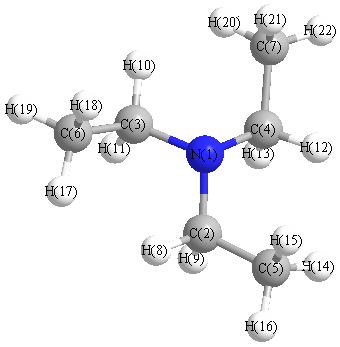
picture of triethylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C5 112.314 N1 C2 H8 108.079
N1 C2 H9 111.149 N1 C3 C6 112.314
N1 C3 H10 108.079 N1 C3 H11 111.149
N1 C4 C7 112.314 N1 C4 H12 108.079
N1 C4 H13 111.149 C2 N1 C3 111.008
C2 N1 C4 111.008 C2 C5 H14 111.993
C2 C5 H15 109.764 C2 C5 H16 110.433
C3 N1 C4 111.008 C3 C6 H17 111.993
C3 C6 H18 109.764 C3 C6 H19 110.433
C4 C7 H20 111.993 C4 C7 H21 109.764
C4 C7 H22 110.433 C5 C2 H8 108.294
C5 C2 H9 110.454 C6 C3 H10 108.294
C6 C3 H11 110.454 C7 C4 H12 108.294
C7 C4 H13 110.454 H8 C2 H9 106.297
H10 C3 H11 106.297 H12 C4 H13 106.296
H14 C5 H15 108.037 H14 C5 H16 107.703
H15 C5 H16 108.817 H17 C6 H18 108.037
H17 C6 H19 107.703 H18 C6 H19 108.817
H20 C7 H21 108.037 H20 C7 H22 107.703
H21 C7 H22 108.817
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability