return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C7H8 (Norbornadiene)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-270.654006
Energy at 298.15K-270.663429
Nuclear repulsion energy288.767920
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3321 3103 6.46      
2 A1 3208 2998 7.64      
3 A1 3144 2938 36.13      
4 A1 1651 1543 0.10      
5 A1 1545 1443 0.16      
6 A1 1295 1210 3.16      
7 A1 1154 1078 0.19      
8 A1 979 914 2.23      
9 A1 925 864 4.64      
10 A1 803 751 1.34      
11 A1 751 702 49.95      
12 A1 436 408 3.94      
13 A2 3294 3078 0.00      
14 A2 1339 1251 0.00      
15 A2 1303 1217 0.00      
16 A2 1166 1089 0.00      
17 A2 958 895 0.00      
18 A2 904 845 0.00      
19 A2 728 681 0.00      
20 A2 457 427 0.00      
21 B1 3319 3101 17.91      
22 B1 3221 3010 31.46      
23 B1 1613 1507 3.76      
24 B1 1272 1189 1.24      
25 B1 1109 1037 0.18      
26 B1 1061 991 0.19      
27 B1 951 888 1.26      
28 B1 684 639 31.21      
29 B1 515 481 5.38      
30 B2 3296 3080 7.61      
31 B2 3206 2996 48.57      
32 B2 1381 1291 18.65      
33 B2 1311 1225 0.02      
34 B2 1205 1126 0.58      
35 B2 994 929 0.00      
36 B2 923 862 0.03      
37 B2 905 845 3.69      
38 B2 835 780 5.92      
39 B2 557 521 1.36      

Unscaled Zero Point Vibrational Energy (zpe) 28858.5 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 26965.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.14404 0.12173 0.10691

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.343
C2 0.000 1.117 0.272
C3 0.000 -1.117 0.272
C4 1.234 0.671 -0.517
C5 1.234 -0.671 -0.517
C6 -1.234 0.671 -0.517
C7 -1.234 -0.671 -0.517
H8 -0.898 0.000 1.961
H9 0.898 0.000 1.961
H10 0.000 2.149 0.614
H11 0.000 -2.149 0.614
H12 1.920 1.336 -1.018
H13 1.920 -1.336 -1.018
H14 -1.920 1.336 -1.018
H15 -1.920 -1.336 -1.018

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15
C11.54831.54832.33092.33092.33092.33091.08951.08952.26942.26943.32413.32413.32413.3241
C21.54832.23481.53062.31171.53062.31172.21522.21521.08693.28432.32353.37192.32353.3719
C31.54832.23482.31171.53062.31171.53062.21522.21523.28431.08693.37192.32353.37192.3235
C42.33091.53062.31171.34302.46772.80953.33642.58852.23253.27961.07902.18003.26213.7722
C52.33092.31171.53061.34302.80952.46773.33642.58853.27962.23252.18001.07903.77223.2621
C62.33091.53062.31172.46772.80951.34302.58853.33642.23253.27963.26213.77221.07902.1800
C72.33092.31171.53062.80952.46771.34302.58853.33643.27962.23253.77223.26212.18001.0790
H81.08952.21522.21523.33643.33642.58852.58851.79562.69022.69024.31274.31273.42103.4210
H91.08952.21522.21522.58852.58853.33643.33641.79562.69022.69023.42103.42104.31274.3127
H102.26941.08693.28432.23253.27962.23253.27962.69022.69024.29792.64824.30072.64824.3007
H112.26943.28431.08693.27962.23253.27962.23252.69022.69024.29794.30072.64824.30072.6482
H123.32412.32353.37191.07902.18003.26213.77224.31273.42102.64824.30072.67193.84044.6784
H133.32413.37192.32352.18001.07903.77223.26214.31273.42104.30072.64822.67194.67843.8404
H143.32412.32353.37193.26213.77221.07902.18003.42104.31272.64824.30073.84044.67842.6719
H153.32413.37192.32353.77223.26212.18001.07903.42104.31274.30072.64824.67843.84042.6719

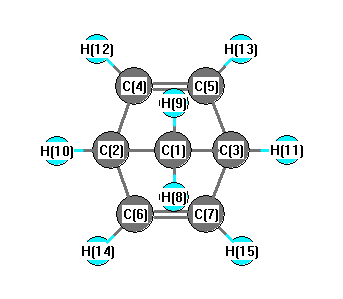
picture of Norbornadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 98.410 C1 C2 C6 98.410
C1 C2 H10 117.830 C1 C3 C5 98.410
C1 C3 C7 98.410 C1 C3 H11 117.830
C2 C1 C3 92.389 C2 C1 H8 113.086
C2 C1 H9 113.086 C2 C4 C5 106.938
C2 C4 H12 124.947 C2 C6 C7 106.938
C2 C6 H14 124.947 C3 C1 H8 113.086
C3 C1 H9 113.086 C3 C5 C4 106.938
C3 C5 H13 124.947 C3 C7 C6 106.938
C3 C7 H15 124.947 C4 C2 C6 107.440
C4 C2 H10 116.025 C4 C5 H13 128.011
C5 C3 C7 107.440 C5 C3 H11 116.025
C5 C4 H12 128.011 C6 C2 H10 116.025
C6 C7 H15 128.011 C7 C3 H11 116.025
C7 C6 H14 128.011 H8 C1 H9 110.991
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability