Jump to
S1C2
S1C3
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -208.610957 |
| Energy at 298.15K | -208.616976 |
| HF Energy | -207.967721 |
| Nuclear repulsion energy | 119.102917 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3708 |
3465 |
38.19 |
|
|
|
| 2 |
A' |
3242 |
3030 |
15.28 |
|
|
|
| 3 |
A' |
3127 |
2922 |
49.29 |
|
|
|
| 4 |
A' |
3050 |
2850 |
85.90 |
|
|
|
| 5 |
A' |
1837 |
1716 |
404.91 |
|
|
|
| 6 |
A' |
1592 |
1487 |
11.87 |
|
|
|
| 7 |
A' |
1520 |
1421 |
28.83 |
|
|
|
| 8 |
A' |
1519 |
1419 |
0.15 |
|
|
|
| 9 |
A' |
1441 |
1347 |
16.41 |
|
|
|
| 10 |
A' |
1334 |
1246 |
98.10 |
|
|
|
| 11 |
A' |
1203 |
1124 |
30.70 |
|
|
|
| 12 |
A' |
1040 |
972 |
41.96 |
|
|
|
| 13 |
A' |
617 |
577 |
13.80 |
|
|
|
| 14 |
A' |
349 |
326 |
7.86 |
|
|
|
| 15 |
A" |
3211 |
3000 |
27.88 |
|
|
|
| 16 |
A" |
1545 |
1443 |
4.62 |
|
|
|
| 17 |
A" |
1186 |
1108 |
0.97 |
|
|
|
| 18 |
A" |
1057 |
988 |
0.43 |
|
|
|
| 19 |
A" |
615 |
575 |
139.29 |
|
|
|
| 20 |
A" |
200 |
187 |
1.79 |
|
|
|
| 21 |
A" |
74 |
69 |
1.93 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16732.6 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 15635.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.276 |
-0.758 |
0.000 |
| O2 |
1.402 |
-1.241 |
0.000 |
| N3 |
0.000 |
0.573 |
0.000 |
| C4 |
-1.329 |
1.139 |
0.000 |
| H5 |
-0.640 |
-1.370 |
0.000 |
| H6 |
0.803 |
1.182 |
0.000 |
| H7 |
-2.050 |
0.326 |
0.000 |
| H8 |
-1.505 |
1.748 |
0.885 |
| H9 |
-1.505 |
1.748 |
-0.885 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2251 | 1.3600 | 2.4856 | 1.1017 | 2.0108 | 2.5661 | 3.1992 | 3.1992 |
O2 | 1.2251 | | 2.2932 | 3.6231 | 2.0464 | 2.4962 | 3.7908 | 4.2622 | 4.2622 | N3 | 1.3600 | 2.2932 | | 1.4448 | 2.0457 | 1.0080 | 2.0648 | 2.1040 | 2.1040 | C4 | 2.4856 | 3.6231 | 1.4448 | | 2.6015 | 2.1333 | 1.0867 | 1.0886 | 1.0886 | H5 | 1.1017 | 2.0464 | 2.0457 | 2.6015 | | 2.9318 | 2.2045 | 3.3538 | 3.3538 | H6 | 2.0108 | 2.4962 | 1.0080 | 2.1333 | 2.9318 | | 2.9791 | 2.5360 | 2.5360 | H7 | 2.5661 | 3.7908 | 2.0648 | 1.0867 | 2.2045 | 2.9791 | | 1.7617 | 1.7617 | H8 | 3.1992 | 4.2622 | 2.1040 | 1.0886 | 3.3538 | 2.5360 | 1.7617 | | 1.7709 | H9 | 3.1992 | 4.2622 | 2.1040 | 1.0886 | 3.3538 | 2.5360 | 1.7617 | 1.7709 | |
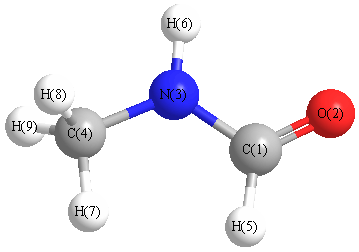 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
124.767 |
|
C1 |
N3 |
H6 |
115.433 |
| O2 |
C1 |
N3 |
124.935 |
|
O2 |
C1 |
H5 |
123.083 |
| N3 |
C1 |
H5 |
111.982 |
|
N3 |
C4 |
H7 |
108.474 |
| N3 |
C4 |
H8 |
111.529 |
|
N3 |
C4 |
H9 |
111.529 |
| C4 |
N3 |
H6 |
119.800 |
|
H7 |
C4 |
H8 |
108.168 |
| H7 |
C4 |
H9 |
108.168 |
|
H8 |
C4 |
H9 |
108.856 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -208.610957 |
| Energy at 298.15K | -208.616976 |
| HF Energy | -207.967721 |
| Nuclear repulsion energy | 119.102917 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -208.613095 |
| Energy at 298.15K | -208.619068 |
| HF Energy | -207.969393 |
| Nuclear repulsion energy | 121.412638 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3738 |
3493 |
38.66 |
|
|
|
| 2 |
A |
3272 |
3057 |
1.05 |
|
|
|
| 3 |
A |
3218 |
3007 |
28.37 |
|
|
|
| 4 |
A |
3135 |
2930 |
41.54 |
|
|
|
| 5 |
A |
3065 |
2864 |
111.42 |
|
|
|
| 6 |
A |
1823 |
1704 |
284.08 |
|
|
|
| 7 |
A |
1602 |
1497 |
67.33 |
|
|
|
| 8 |
A |
1565 |
1462 |
6.40 |
|
|
|
| 9 |
A |
1555 |
1453 |
65.75 |
|
|
|
| 10 |
A |
1489 |
1391 |
18.90 |
|
|
|
| 11 |
A |
1459 |
1363 |
3.38 |
|
|
|
| 12 |
A |
1258 |
1175 |
78.18 |
|
|
|
| 13 |
A |
1191 |
1113 |
1.49 |
|
|
|
| 14 |
A |
1186 |
1108 |
16.65 |
|
|
|
| 15 |
A |
1037 |
969 |
0.44 |
|
|
|
| 16 |
A |
999 |
933 |
19.41 |
|
|
|
| 17 |
A |
778 |
727 |
0.62 |
|
|
|
| 18 |
A |
534 |
499 |
47.43 |
|
|
|
| 19 |
A |
310 |
290 |
13.41 |
|
|
|
| 20 |
A |
251 |
235 |
89.86 |
|
|
|
| 21 |
A |
31 |
29 |
0.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16748.3 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 15649.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.858 |
0.443 |
0.005 |
| O2 |
1.374 |
-0.671 |
-0.000 |
| N3 |
-0.480 |
0.661 |
-0.022 |
| C4 |
-1.409 |
-0.450 |
0.005 |
| H5 |
1.440 |
1.376 |
0.019 |
| H6 |
-0.819 |
1.604 |
0.061 |
| H7 |
-2.406 |
-0.078 |
-0.208 |
| H8 |
-1.125 |
-1.175 |
-0.751 |
| H9 |
-1.409 |
-0.947 |
0.974 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2279 | 1.3560 | 2.4369 | 1.0995 | 2.0400 | 3.3119 | 2.6685 | 2.8300 |
O2 | 1.2279 | | 2.2831 | 2.7918 | 2.0479 | 3.1600 | 3.8313 | 2.6569 | 2.9608 | N3 | 1.3560 | 2.2831 | | 1.4489 | 2.0494 | 1.0047 | 2.0712 | 2.0785 | 2.1073 | C4 | 2.4369 | 2.7918 | 1.4489 | | 3.3843 | 2.1375 | 1.0847 | 1.0857 | 1.0885 | H5 | 1.0995 | 2.0479 | 2.0494 | 3.3843 | | 2.2711 | 4.1179 | 3.6985 | 3.7979 | H6 | 2.0400 | 3.1600 | 1.0047 | 2.1375 | 2.2711 | | 2.3278 | 2.9112 | 2.7725 | H7 | 3.3119 | 3.8313 | 2.0712 | 1.0847 | 4.1179 | 2.3278 | | 1.7718 | 1.7737 | H8 | 2.6685 | 2.6569 | 2.0785 | 1.0857 | 3.6985 | 2.9112 | 1.7718 | | 1.7623 | H9 | 2.8300 | 2.9608 | 2.1073 | 1.0885 | 3.7979 | 2.7725 | 1.7737 | 1.7623 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
120.600 |
|
C1 |
N3 |
H6 |
118.820 |
| O2 |
C1 |
N3 |
124.089 |
|
O2 |
C1 |
H5 |
123.174 |
| N3 |
C1 |
H5 |
112.733 |
|
N3 |
C4 |
H7 |
108.813 |
| N3 |
C4 |
H8 |
109.345 |
|
N3 |
C4 |
H9 |
111.507 |
| C4 |
N3 |
H6 |
120.101 |
|
H7 |
C4 |
H8 |
109.438 |
| H7 |
C4 |
H9 |
109.410 |
|
H8 |
C4 |
H9 |
108.305 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
