Jump to
S1C2
S1C3
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -208.985370 |
| Energy at 298.15K | -208.991451 |
| HF Energy | -208.985370 |
| Nuclear repulsion energy | 119.453189 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3659 |
3492 |
28.21 |
|
|
|
| 2 |
A' |
3156 |
3012 |
14.68 |
|
|
|
| 3 |
A' |
3053 |
2914 |
52.88 |
|
|
|
| 4 |
A' |
2959 |
2824 |
92.45 |
|
|
|
| 5 |
A' |
1861 |
1776 |
456.03 |
|
|
|
| 6 |
A' |
1547 |
1476 |
23.31 |
|
|
|
| 7 |
A' |
1502 |
1434 |
13.82 |
|
|
|
| 8 |
A' |
1476 |
1409 |
6.24 |
|
|
|
| 9 |
A' |
1407 |
1342 |
13.84 |
|
|
|
| 10 |
A' |
1318 |
1258 |
90.06 |
|
|
|
| 11 |
A' |
1186 |
1132 |
24.96 |
|
|
|
| 12 |
A' |
1026 |
979 |
44.96 |
|
|
|
| 13 |
A' |
621 |
592 |
16.01 |
|
|
|
| 14 |
A' |
346 |
330 |
8.52 |
|
|
|
| 15 |
A" |
3119 |
2976 |
29.50 |
|
|
|
| 16 |
A" |
1492 |
1424 |
6.37 |
|
|
|
| 17 |
A" |
1154 |
1101 |
0.20 |
|
|
|
| 18 |
A" |
1049 |
1001 |
0.11 |
|
|
|
| 19 |
A" |
628 |
599 |
135.07 |
|
|
|
| 20 |
A" |
209 |
199 |
1.28 |
|
|
|
| 21 |
A" |
107 |
102 |
0.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16436.8 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 15685.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.280 |
-0.757 |
0.000 |
| O2 |
1.397 |
-1.232 |
0.000 |
| N3 |
0.000 |
0.570 |
0.000 |
| C4 |
-1.325 |
1.132 |
0.000 |
| H5 |
-0.642 |
-1.374 |
0.000 |
| H6 |
0.802 |
1.183 |
0.000 |
| H7 |
-2.053 |
0.317 |
0.000 |
| H8 |
-1.508 |
1.746 |
0.889 |
| H9 |
-1.508 |
1.746 |
-0.889 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2136 | 1.3558 | 2.4792 | 1.1103 | 2.0086 | 2.5685 | 3.2016 | 3.2016 |
O2 | 1.2136 | | 2.2799 | 3.6056 | 2.0444 | 2.4870 | 3.7818 | 4.2538 | 4.2538 | N3 | 1.3558 | 2.2799 | | 1.4395 | 2.0474 | 1.0098 | 2.0685 | 2.1088 | 2.1088 | C4 | 2.4792 | 3.6056 | 1.4395 | | 2.5981 | 2.1279 | 1.0933 | 1.0956 | 1.0956 | H5 | 1.1103 | 2.0444 | 2.0474 | 2.5981 | | 2.9370 | 2.2023 | 3.3577 | 3.3577 | H6 | 2.0086 | 2.4870 | 1.0098 | 2.1279 | 2.9370 | | 2.9838 | 2.5385 | 2.5385 | H7 | 2.5685 | 3.7818 | 2.0685 | 1.0933 | 2.2023 | 2.9838 | | 1.7692 | 1.7692 | H8 | 3.2016 | 4.2538 | 2.1088 | 1.0956 | 3.3577 | 2.5385 | 1.7692 | | 1.7787 | H9 | 3.2016 | 4.2538 | 2.1088 | 1.0956 | 3.3577 | 2.5385 | 1.7692 | 1.7787 | |
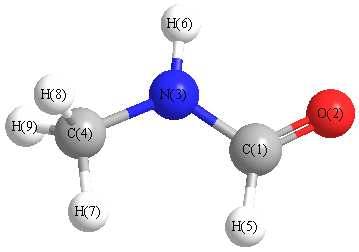 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
124.945 |
|
C1 |
N3 |
H6 |
115.453 |
| O2 |
C1 |
N3 |
124.985 |
|
O2 |
C1 |
H5 |
123.154 |
| N3 |
C1 |
H5 |
111.861 |
|
N3 |
C4 |
H7 |
108.740 |
| N3 |
C4 |
H8 |
111.856 |
|
N3 |
C4 |
H9 |
111.856 |
| C4 |
N3 |
H6 |
119.602 |
|
H7 |
C4 |
H8 |
107.852 |
| H7 |
C4 |
H9 |
107.852 |
|
H8 |
C4 |
H9 |
108.529 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.406 |
|
|
|
| 2 |
O |
-0.477 |
|
|
|
| 3 |
N |
-0.508 |
|
|
|
| 4 |
C |
-0.242 |
|
|
|
| 5 |
H |
0.080 |
|
|
|
| 6 |
H |
0.290 |
|
|
|
| 7 |
H |
0.152 |
|
|
|
| 8 |
H |
0.150 |
|
|
|
| 9 |
H |
0.150 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.914 |
2.768 |
0.000 |
4.019 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.865 |
3.911 |
0.000 |
| y |
3.911 |
-24.012 |
0.000 |
| z |
0.000 |
0.000 |
-24.451 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.634 |
3.911 |
0.000 |
| y |
3.911 |
1.146 |
0.000 |
| z |
0.000 |
0.000 |
0.488 |
|
| Polar |
|---|
| 3z2-r2 | 0.975 |
|---|
| x2-y2 | -1.853 |
|---|
| xy | 3.911 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.464 |
-1.053 |
0.000 |
| y |
-1.053 |
5.561 |
0.000 |
| z |
0.000 |
0.000 |
3.013 |
<r2> (average value of r
2) Å
2
| <r2> |
88.521 |
| (<r2>)1/2 |
9.409 |
Jump to
S1C1
S1C3
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -208.985370 |
| Energy at 298.15K | -208.991451 |
| HF Energy | -208.985370 |
| Nuclear repulsion energy | 119.453189 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -208.987131 |
| Energy at 298.15K | -208.993241 |
| HF Energy | -208.987131 |
| Nuclear repulsion energy | 121.731980 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3691 |
3522 |
27.54 |
|
|
|
| 2 |
A |
3190 |
3044 |
0.53 |
|
|
|
| 3 |
A |
3128 |
2985 |
29.59 |
|
|
|
| 4 |
A |
3061 |
2921 |
43.60 |
|
|
|
| 5 |
A |
2983 |
2846 |
117.21 |
|
|
|
| 6 |
A |
1846 |
1762 |
316.04 |
|
|
|
| 7 |
A |
1577 |
1505 |
101.80 |
|
|
|
| 8 |
A |
1511 |
1442 |
8.27 |
|
|
|
| 9 |
A |
1504 |
1436 |
34.69 |
|
|
|
| 10 |
A |
1445 |
1379 |
21.87 |
|
|
|
| 11 |
A |
1433 |
1368 |
0.95 |
|
|
|
| 12 |
A |
1244 |
1187 |
67.31 |
|
|
|
| 13 |
A |
1165 |
1112 |
22.66 |
|
|
|
| 14 |
A |
1158 |
1106 |
0.51 |
|
|
|
| 15 |
A |
1031 |
984 |
0.68 |
|
|
|
| 16 |
A |
993 |
948 |
20.39 |
|
|
|
| 17 |
A |
779 |
743 |
0.46 |
|
|
|
| 18 |
A |
545 |
520 |
50.72 |
|
|
|
| 19 |
A |
305 |
291 |
14.50 |
|
|
|
| 20 |
A |
279 |
267 |
79.44 |
|
|
|
| 21 |
A |
92 |
88 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16480.0 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 15726.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.860 |
0.436 |
0.004 |
| O2 |
1.366 |
-0.671 |
-0.000 |
| N3 |
-0.473 |
0.659 |
-0.020 |
| C4 |
-1.411 |
-0.442 |
0.005 |
| H5 |
1.452 |
1.370 |
0.018 |
| H6 |
-0.807 |
1.605 |
0.057 |
| H7 |
-2.411 |
-0.066 |
-0.218 |
| H8 |
-1.126 |
-1.180 |
-0.749 |
| H9 |
-1.421 |
-0.939 |
0.980 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2176 | 1.3510 | 2.4350 | 1.1058 | 2.0362 | 3.3162 | 2.6688 | 2.8371 |
O2 | 1.2176 | | 2.2693 | 2.7868 | 2.0430 | 3.1473 | 3.8312 | 2.6516 | 2.9672 | N3 | 1.3510 | 2.2693 | | 1.4471 | 2.0518 | 1.0065 | 2.0787 | 2.0828 | 2.1109 | C4 | 2.4350 | 2.7868 | 1.4471 | | 3.3884 | 2.1354 | 1.0909 | 1.0921 | 1.0950 | H5 | 1.1058 | 2.0430 | 2.0518 | 3.3884 | | 2.2709 | 4.1274 | 3.7060 | 3.8099 | H6 | 2.0362 | 3.1473 | 1.0065 | 2.1354 | 2.2709 | | 2.3325 | 2.9164 | 2.7759 | H7 | 3.3162 | 3.8312 | 2.0787 | 1.0909 | 4.1274 | 2.3325 | | 1.7809 | 1.7825 | H8 | 2.6688 | 2.6516 | 2.0828 | 1.0921 | 3.7060 | 2.9164 | 1.7809 | | 1.7703 | H9 | 2.8371 | 2.9672 | 2.1109 | 1.0950 | 3.8099 | 2.7759 | 1.7825 | 1.7703 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
120.935 |
|
C1 |
N3 |
H6 |
118.748 |
| O2 |
C1 |
N3 |
124.048 |
|
O2 |
C1 |
H5 |
123.058 |
| N3 |
C1 |
H5 |
112.891 |
|
N3 |
C4 |
H7 |
109.167 |
| N3 |
C4 |
H8 |
109.426 |
|
N3 |
C4 |
H9 |
111.524 |
| C4 |
N3 |
H6 |
119.910 |
|
H7 |
C4 |
H8 |
109.333 |
| H7 |
C4 |
H9 |
109.271 |
|
H8 |
C4 |
H9 |
108.088 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.398 |
|
|
|
| 2 |
O |
-0.475 |
|
|
|
| 3 |
N |
-0.519 |
|
|
|
| 4 |
C |
-0.248 |
|
|
|
| 5 |
H |
0.088 |
|
|
|
| 6 |
H |
0.283 |
|
|
|
| 7 |
H |
0.144 |
|
|
|
| 8 |
H |
0.186 |
|
|
|
| 9 |
H |
0.144 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.846 |
2.545 |
0.000 |
3.818 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.151 |
2.448 |
0.000 |
| y |
2.448 |
-21.702 |
0.000 |
| z |
0.000 |
0.000 |
-24.469 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.066 |
2.448 |
0.000 |
| y |
2.448 |
3.109 |
0.000 |
| z |
0.000 |
0.000 |
-1.043 |
|
| Polar |
|---|
| 3z2-r2 | -2.086 |
|---|
| x2-y2 | -3.450 |
|---|
| xy | 2.448 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.932 |
-0.093 |
0.000 |
| y |
-0.093 |
4.867 |
0.000 |
| z |
0.000 |
0.000 |
2.999 |
<r2> (average value of r
2) Å
2
| <r2> |
77.747 |
| (<r2>)1/2 |
8.817 |
