Jump to
S1C2
S1C3
Energy calculated at MP3=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -208.629026 |
| Energy at 298.15K | -208.635044 |
| HF Energy | -207.968931 |
| Nuclear repulsion energy | 119.492737 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3756 |
3756 |
35.66 |
|
|
|
| 2 |
A' |
3238 |
3238 |
18.87 |
|
|
|
| 3 |
A' |
3136 |
3136 |
49.50 |
|
|
|
| 4 |
A' |
3077 |
3077 |
75.31 |
|
|
|
| 5 |
A' |
1898 |
1898 |
497.73 |
|
|
|
| 6 |
A' |
1601 |
1601 |
15.88 |
|
|
|
| 7 |
A' |
1555 |
1555 |
24.20 |
|
|
|
| 8 |
A' |
1534 |
1534 |
9.52 |
|
|
|
| 9 |
A' |
1460 |
1460 |
13.59 |
|
|
|
| 10 |
A' |
1349 |
1349 |
126.47 |
|
|
|
| 11 |
A' |
1215 |
1215 |
30.53 |
|
|
|
| 12 |
A' |
1050 |
1050 |
44.21 |
|
|
|
| 13 |
A' |
629 |
629 |
15.49 |
|
|
|
| 14 |
A' |
354 |
354 |
8.80 |
|
|
|
| 15 |
A" |
3209 |
3209 |
31.86 |
|
|
|
| 16 |
A" |
1551 |
1551 |
4.22 |
|
|
|
| 17 |
A" |
1193 |
1193 |
1.40 |
|
|
|
| 18 |
A" |
1081 |
1081 |
0.00 |
|
|
|
| 19 |
A" |
607 |
607 |
147.86 |
|
|
|
| 20 |
A" |
196 |
196 |
1.44 |
|
|
|
| 21 |
A" |
66 |
66 |
2.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16878.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16878.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.283 |
-0.756 |
0.000 |
| O2 |
1.402 |
-1.227 |
0.000 |
| N3 |
0.000 |
0.571 |
0.000 |
| C4 |
-1.334 |
1.127 |
0.000 |
| H5 |
-0.624 |
-1.378 |
0.000 |
| H6 |
0.795 |
1.185 |
0.000 |
| H7 |
-2.051 |
0.311 |
0.000 |
| H8 |
-1.513 |
1.735 |
0.885 |
| H9 |
-1.513 |
1.735 |
-0.885 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2140 | 1.3569 | 2.4823 | 1.0994 | 2.0076 | 2.5662 | 3.1957 | 3.1957 |
O2 | 1.2140 | | 2.2797 | 3.6093 | 2.0317 | 2.4870 | 3.7798 | 4.2485 | 4.2485 | N3 | 1.3569 | 2.2797 | | 1.4455 | 2.0462 | 1.0044 | 2.0674 | 2.1040 | 2.1040 | C4 | 2.4823 | 3.6093 | 1.4455 | | 2.6037 | 2.1297 | 1.0863 | 1.0881 | 1.0881 | H5 | 1.0994 | 2.0317 | 2.0462 | 2.6037 | | 2.9294 | 2.2109 | 3.3557 | 3.3557 | H6 | 2.0076 | 2.4870 | 1.0044 | 2.1297 | 2.9294 | | 2.9769 | 2.5320 | 2.5320 | H7 | 2.5662 | 3.7798 | 2.0674 | 1.0863 | 2.2109 | 2.9769 | | 1.7604 | 1.7604 | H8 | 3.1957 | 4.2485 | 2.1040 | 1.0881 | 3.3557 | 2.5320 | 1.7604 | | 1.7697 | H9 | 3.1957 | 4.2485 | 2.1040 | 1.0881 | 3.3557 | 2.5320 | 1.7604 | 1.7697 | |
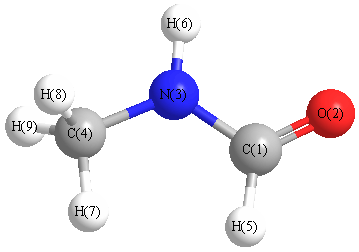 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
124.662 |
|
C1 |
N3 |
H6 |
115.666 |
| O2 |
C1 |
N3 |
124.828 |
|
O2 |
C1 |
H5 |
122.772 |
| N3 |
C1 |
H5 |
112.399 |
|
N3 |
C4 |
H7 |
108.655 |
| N3 |
C4 |
H8 |
111.508 |
|
N3 |
C4 |
H9 |
111.508 |
| C4 |
N3 |
H6 |
119.672 |
|
H7 |
C4 |
H8 |
108.114 |
| H7 |
C4 |
H9 |
108.114 |
|
H8 |
C4 |
H9 |
108.825 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP3=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -208.629026 |
| Energy at 298.15K | -208.635044 |
| HF Energy | -207.968931 |
| Nuclear repulsion energy | 119.492737 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3=FULL/6-31G**
Geometric Data calculated at MP3=FULL/6-31G**
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP3=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -208.631093 |
| Energy at 298.15K | -208.637059 |
| HF Energy | -207.970630 |
| Nuclear repulsion energy | 121.889507 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3787 |
3787 |
38.07 |
|
|
|
| 2 |
A |
3272 |
3272 |
6.28 |
|
|
|
| 3 |
A |
3226 |
3226 |
27.37 |
|
|
|
| 4 |
A |
3144 |
3144 |
37.22 |
|
|
|
| 5 |
A |
3094 |
3094 |
102.99 |
|
|
|
| 6 |
A |
1884 |
1884 |
335.89 |
|
|
|
| 7 |
A |
1614 |
1614 |
121.08 |
|
|
|
| 8 |
A |
1573 |
1573 |
19.65 |
|
|
|
| 9 |
A |
1558 |
1558 |
29.35 |
|
|
|
| 10 |
A |
1510 |
1510 |
13.96 |
|
|
|
| 11 |
A |
1483 |
1483 |
4.44 |
|
|
|
| 12 |
A |
1281 |
1281 |
73.74 |
|
|
|
| 13 |
A |
1207 |
1207 |
0.86 |
|
|
|
| 14 |
A |
1191 |
1191 |
11.11 |
|
|
|
| 15 |
A |
1066 |
1066 |
1.64 |
|
|
|
| 16 |
A |
1010 |
1010 |
17.62 |
|
|
|
| 17 |
A |
792 |
792 |
6.82 |
|
|
|
| 18 |
A |
501 |
501 |
42.84 |
|
|
|
| 19 |
A |
300 |
300 |
13.52 |
|
|
|
| 20 |
A |
246 |
246 |
106.51 |
|
|
|
| 21 |
A |
41 |
41 |
0.24 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16888.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16888.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3=FULL/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.861 |
0.433 |
0.008 |
| O2 |
1.381 |
-0.665 |
-0.001 |
| N3 |
-0.477 |
0.656 |
-0.036 |
| C4 |
-1.423 |
-0.441 |
0.008 |
| H5 |
1.442 |
1.364 |
0.031 |
| H6 |
-0.804 |
1.593 |
0.097 |
| H7 |
-2.346 |
-0.141 |
-0.477 |
| H8 |
-0.994 |
-1.282 |
-0.524 |
| H9 |
-1.641 |
-0.754 |
1.029 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2156 | 1.3566 | 2.4459 | 1.0978 | 2.0309 | 3.2943 | 2.5819 | 2.9513 |
O2 | 1.2156 | | 2.2798 | 2.8134 | 2.0307 | 3.1439 | 3.7943 | 2.5091 | 3.1938 | N3 | 1.3566 | 2.2798 | | 1.4496 | 2.0467 | 1.0015 | 2.0799 | 2.0640 | 2.1156 | C4 | 2.4459 | 2.8134 | 1.4496 | | 3.3870 | 2.1281 | 1.0853 | 1.0836 | 1.0891 | H5 | 1.0978 | 2.0307 | 2.0467 | 3.3870 | | 2.2590 | 4.1084 | 3.6395 | 3.8716 | H6 | 2.0309 | 3.1439 | 1.0015 | 2.1281 | 2.2590 | | 2.3905 | 2.9470 | 2.6605 | H7 | 3.2943 | 3.7943 | 2.0799 | 1.0853 | 4.1084 | 2.3905 | | 1.7698 | 1.7721 | H8 | 2.5819 | 2.5091 | 2.0640 | 1.0836 | 3.6395 | 2.9470 | 1.7698 | | 1.7627 | H9 | 2.9513 | 3.1938 | 2.1156 | 1.0891 | 3.8716 | 2.6605 | 1.7721 | 1.7627 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
121.255 |
|
C1 |
N3 |
H6 |
118.129 |
| O2 |
C1 |
N3 |
124.740 |
|
O2 |
C1 |
H5 |
122.673 |
| N3 |
C1 |
H5 |
112.575 |
|
N3 |
C4 |
H7 |
109.430 |
| N3 |
C4 |
H8 |
108.266 |
|
N3 |
C4 |
H9 |
112.114 |
| C4 |
N3 |
H6 |
119.382 |
|
H7 |
C4 |
H8 |
109.371 |
| H7 |
C4 |
H9 |
109.169 |
|
H8 |
C4 |
H9 |
108.449 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
