Jump to
S1C2
S1C3
Energy calculated at TPSSh/6-31G**
| | hartrees |
|---|
| Energy at 0K | -209.211930 |
| Energy at 298.15K | -209.217963 |
| HF Energy | -209.211930 |
| Nuclear repulsion energy | 118.834336 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3604 |
3455 |
19.55 |
|
|
|
| 2 |
A' |
3131 |
3002 |
20.15 |
|
|
|
| 3 |
A' |
3034 |
2909 |
58.70 |
|
|
|
| 4 |
A' |
2941 |
2820 |
102.23 |
|
|
|
| 5 |
A' |
1812 |
1737 |
421.15 |
|
|
|
| 6 |
A' |
1552 |
1488 |
13.44 |
|
|
|
| 7 |
A' |
1489 |
1427 |
6.58 |
|
|
|
| 8 |
A' |
1480 |
1419 |
12.10 |
|
|
|
| 9 |
A' |
1402 |
1344 |
14.12 |
|
|
|
| 10 |
A' |
1300 |
1246 |
94.28 |
|
|
|
| 11 |
A' |
1164 |
1116 |
36.12 |
|
|
|
| 12 |
A' |
1009 |
967 |
39.19 |
|
|
|
| 13 |
A' |
604 |
579 |
15.06 |
|
|
|
| 14 |
A' |
333 |
319 |
8.42 |
|
|
|
| 15 |
A" |
3093 |
2965 |
37.97 |
|
|
|
| 16 |
A" |
1508 |
1446 |
4.41 |
|
|
|
| 17 |
A" |
1152 |
1105 |
0.50 |
|
|
|
| 18 |
A" |
1033 |
990 |
0.67 |
|
|
|
| 19 |
A" |
618 |
592 |
127.90 |
|
|
|
| 20 |
A" |
207 |
198 |
1.26 |
|
|
|
| 21 |
A" |
104 |
99 |
1.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16284.9 cm
-1
Scaled (by 0.9587) Zero Point Vibrational Energy (zpe) 15612.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.281 |
-0.762 |
0.000 |
| O2 |
1.405 |
-1.239 |
0.000 |
| N3 |
0.000 |
0.572 |
0.000 |
| C4 |
-1.335 |
1.141 |
0.000 |
| H5 |
-0.642 |
-1.377 |
0.000 |
| H6 |
0.804 |
1.188 |
0.000 |
| H7 |
-2.059 |
0.321 |
0.000 |
| H8 |
-1.511 |
1.754 |
0.891 |
| H9 |
-1.511 |
1.754 |
-0.891 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2218 | 1.3631 | 2.4961 | 1.1087 | 2.0191 | 2.5782 | 3.2144 | 3.2144 |
O2 | 1.2218 | | 2.2924 | 3.6296 | 2.0517 | 2.5009 | 3.7996 | 4.2727 | 4.2727 | N3 | 1.3631 | 2.2924 | | 1.4511 | 2.0521 | 1.0127 | 2.0740 | 2.1149 | 2.1149 | C4 | 2.4961 | 3.6296 | 1.4511 | | 2.6118 | 2.1391 | 1.0932 | 1.0956 | 1.0956 | H5 | 1.1087 | 2.0517 | 2.0521 | 2.6118 | | 2.9446 | 2.2124 | 3.3696 | 3.3696 | H6 | 2.0191 | 2.5009 | 1.0127 | 2.1391 | 2.9446 | | 2.9907 | 2.5435 | 2.5435 | H7 | 2.5782 | 3.7996 | 2.0740 | 1.0932 | 2.2124 | 2.9907 | | 1.7736 | 1.7736 | H8 | 3.2144 | 4.2727 | 2.1149 | 1.0956 | 3.3696 | 2.5435 | 1.7736 | | 1.7815 | H9 | 3.2144 | 4.2727 | 2.1149 | 1.0956 | 3.3696 | 2.5435 | 1.7736 | 1.7815 | |
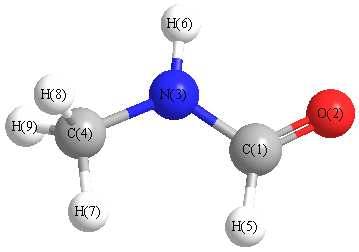 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
124.961 |
|
C1 |
N3 |
H6 |
115.603 |
| O2 |
C1 |
N3 |
124.870 |
|
O2 |
C1 |
H5 |
123.302 |
| N3 |
C1 |
H5 |
111.828 |
|
N3 |
C4 |
H7 |
108.386 |
| N3 |
C4 |
H8 |
111.530 |
|
N3 |
C4 |
H9 |
111.530 |
| C4 |
N3 |
H6 |
119.435 |
|
H7 |
C4 |
H8 |
108.253 |
| H7 |
C4 |
H9 |
108.253 |
|
H8 |
C4 |
H9 |
108.783 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.406 |
|
|
|
| 2 |
O |
-0.479 |
|
|
|
| 3 |
N |
-0.503 |
|
|
|
| 4 |
C |
-0.216 |
|
|
|
| 5 |
H |
0.078 |
|
|
|
| 6 |
H |
0.286 |
|
|
|
| 7 |
H |
0.144 |
|
|
|
| 8 |
H |
0.142 |
|
|
|
| 9 |
H |
0.142 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.908 |
2.780 |
0.000 |
4.023 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.573 |
-1.092 |
0.000 |
| y |
-1.092 |
5.688 |
0.000 |
| z |
0.000 |
0.000 |
3.032 |
<r2> (average value of r
2) Å
2
| <r2> |
89.310 |
| (<r2>)1/2 |
9.450 |
Jump to
S1C1
S1C3
Energy calculated at TPSSh/6-31G**
| | hartrees |
|---|
| Energy at 0K | -209.211930 |
| Energy at 298.15K | -209.217963 |
| HF Energy | -209.211930 |
| Nuclear repulsion energy | 118.834336 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-31G**
Geometric Data calculated at TPSSh/6-31G**
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at TPSSh/6-31G**
| | hartrees |
|---|
| Energy at 0K | -209.213685 |
| Energy at 298.15K | -209.219749 |
| HF Energy | -209.213685 |
| Nuclear repulsion energy | 121.127645 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3628 |
3479 |
16.99 |
|
|
|
| 2 |
A |
3169 |
3038 |
1.77 |
|
|
|
| 3 |
A |
3105 |
2977 |
38.35 |
|
|
|
| 4 |
A |
3045 |
2919 |
49.47 |
|
|
|
| 5 |
A |
2963 |
2841 |
128.78 |
|
|
|
| 6 |
A |
1799 |
1724 |
291.48 |
|
|
|
| 7 |
A |
1565 |
1501 |
62.08 |
|
|
|
| 8 |
A |
1527 |
1464 |
5.92 |
|
|
|
| 9 |
A |
1511 |
1449 |
56.74 |
|
|
|
| 10 |
A |
1449 |
1389 |
20.62 |
|
|
|
| 11 |
A |
1429 |
1370 |
3.48 |
|
|
|
| 12 |
A |
1218 |
1167 |
85.46 |
|
|
|
| 13 |
A |
1159 |
1111 |
17.90 |
|
|
|
| 14 |
A |
1159 |
1111 |
1.06 |
|
|
|
| 15 |
A |
1013 |
971 |
0.27 |
|
|
|
| 16 |
A |
966 |
926 |
15.52 |
|
|
|
| 17 |
A |
763 |
732 |
0.31 |
|
|
|
| 18 |
A |
540 |
518 |
47.06 |
|
|
|
| 19 |
A |
294 |
282 |
13.99 |
|
|
|
| 20 |
A |
265 |
254 |
77.45 |
|
|
|
| 21 |
A |
95 |
92 |
0.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16330.9 cm
-1
Scaled (by 0.9587) Zero Point Vibrational Energy (zpe) 15656.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.865 |
0.440 |
0.005 |
| O2 |
1.377 |
-0.674 |
-0.001 |
| N3 |
-0.475 |
0.661 |
-0.020 |
| C4 |
-1.422 |
-0.445 |
0.005 |
| H5 |
1.454 |
1.376 |
0.017 |
| H6 |
-0.812 |
1.611 |
0.056 |
| H7 |
-2.420 |
-0.065 |
-0.220 |
| H8 |
-1.136 |
-1.183 |
-0.748 |
| H9 |
-1.434 |
-0.940 |
0.982 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2255 | 1.3590 | 2.4528 | 1.1056 | 2.0461 | 3.3314 | 2.6844 | 2.8542 |
O2 | 1.2255 | | 2.2833 | 2.8082 | 2.0512 | 3.1643 | 3.8514 | 2.6704 | 2.9897 | N3 | 1.3590 | 2.2833 | | 1.4568 | 2.0573 | 1.0100 | 2.0855 | 2.0901 | 2.1187 | C4 | 2.4528 | 2.8082 | 1.4568 | | 3.4040 | 2.1452 | 1.0912 | 1.0921 | 1.0953 | H5 | 1.1056 | 2.0512 | 2.0573 | 3.4040 | | 2.2781 | 4.1397 | 3.7201 | 3.8255 | H6 | 2.0461 | 3.1643 | 1.0100 | 2.1452 | 2.2781 | | 2.3387 | 2.9251 | 2.7840 | H7 | 3.3314 | 3.8514 | 2.0855 | 1.0912 | 4.1397 | 2.3387 | | 1.7825 | 1.7837 | H8 | 2.6844 | 2.6704 | 2.0901 | 1.0921 | 3.7201 | 2.9251 | 1.7825 | | 1.7721 | H9 | 2.8542 | 2.9897 | 2.1187 | 1.0953 | 3.8255 | 2.7840 | 1.7837 | 1.7721 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
121.131 |
|
C1 |
N3 |
H6 |
118.729 |
| O2 |
C1 |
N3 |
124.037 |
|
O2 |
C1 |
H5 |
123.183 |
| N3 |
C1 |
H5 |
112.774 |
|
N3 |
C4 |
H7 |
109.029 |
| N3 |
C4 |
H8 |
109.333 |
|
N3 |
C4 |
H9 |
111.449 |
| C4 |
N3 |
H6 |
119.737 |
|
H7 |
C4 |
H8 |
109.460 |
| H7 |
C4 |
H9 |
109.330 |
|
H8 |
C4 |
H9 |
108.215 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.397 |
|
|
|
| 2 |
O |
-0.475 |
|
|
|
| 3 |
N |
-0.514 |
|
|
|
| 4 |
C |
-0.221 |
|
|
|
| 5 |
H |
0.086 |
|
|
|
| 6 |
H |
0.277 |
|
|
|
| 7 |
H |
0.136 |
|
|
|
| 8 |
H |
0.178 |
|
|
|
| 9 |
H |
0.136 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.860 |
2.501 |
0.000 |
3.799 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.072 |
-0.094 |
0.000 |
| y |
-0.094 |
4.965 |
0.000 |
| z |
0.000 |
0.000 |
3.017 |
<r2> (average value of r
2) Å
2
| <r2> |
78.363 |
| (<r2>)1/2 |
8.852 |
