return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CHOCH2CH2CH3 (Butanal)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-231.766956
Energy at 298.15K-231.775642
HF Energy-230.997301
Nuclear repulsion energy172.344125
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3227 3016 27.01      
2 A' 3151 2944 15.63      
3 A' 3127 2922 21.57      
4 A' 3102 2899 16.77      
5 A' 3001 2804 138.72      
6 A' 1793 1675 85.05      
7 A' 1568 1465 3.98      
8 A' 1548 1446 2.63      
9 A' 1512 1413 10.70      
10 A' 1472 1375 3.06      
11 A' 1461 1365 2.69      
12 A' 1444 1350 16.64      
13 A' 1349 1261 4.65      
14 A' 1173 1096 10.16      
15 A' 1098 1026 0.15      
16 A' 994 929 5.69      
17 A' 891 832 7.16      
18 A' 708 662 13.49      
19 A' 355 332 0.94      
20 A' 203 190 8.03      
21 A" 3224 3012 45.80      
22 A" 3200 2990 2.09      
23 A" 3149 2942 8.05      
24 A" 1564 1461 5.90      
25 A" 1363 1274 0.23      
26 A" 1294 1209 0.04      
27 A" 1188 1110 0.00      
28 A" 972 908 0.02      
29 A" 812 759 1.02      
30 A" 692 647 2.72      
31 A" 261 244 0.03      
32 A" 172 161 1.42      
33 A" 85 79 1.69      

Unscaled Zero Point Vibrational Energy (zpe) 25576.3 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 23898.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.49898 0.08623 0.07660

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.268 -0.349 0.000
H2 2.794 -1.302 0.000
H3 2.588 0.208 0.880
H4 2.588 0.208 -0.880
C5 0.760 -0.553 0.000
H6 0.458 -1.136 -0.870
H7 0.458 -1.136 0.870
C8 0.000 0.762 0.000
H9 0.264 1.372 0.870
H10 0.264 1.372 -0.870
C11 -1.495 0.597 0.000
H12 -2.075 1.538 0.000
O13 -2.067 -0.483 0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 C8 H9 H10 C11 H12 O13
C11.08791.08911.08911.52202.15682.15682.52592.78142.78143.88024.73574.3378
H21.08791.75951.75952.16692.49782.49783.47333.78223.78224.68995.63644.9295
H31.08911.75951.75942.16633.06612.51792.78862.59883.13274.19414.92794.7876
H41.08911.75951.75942.16632.51793.06612.78863.13272.59884.19414.92794.7876
C51.52202.16692.16632.16631.08951.08951.51912.17022.17022.53133.52302.8284
H62.15682.49783.06612.51791.08951.73972.13723.05822.51542.75173.78442.7496
H72.15682.49782.51793.06611.08951.73972.13722.51543.05822.75173.78442.7496
C82.52593.47332.78862.78861.51912.13722.13721.09471.09471.50382.21552.4136
H92.78143.78222.59883.13272.17023.05822.51541.09471.73922.10982.50133.1043
H102.78143.78223.13272.59882.17022.51543.05821.09471.73922.10982.50133.1043
C113.88024.68994.19414.19412.53132.75172.75171.50382.10982.10981.10551.2227
H124.73575.63644.92794.92793.52303.78443.78442.21552.50132.50131.10552.0212
O134.33784.92954.78764.78762.82842.74962.74962.41363.10433.10431.22272.0212

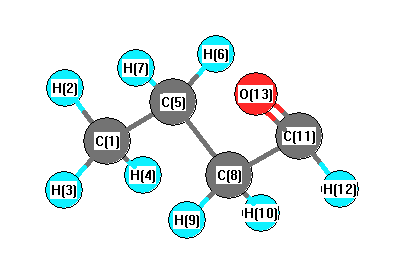
picture of Butanal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 110.265 C1 C5 H7 110.265
C1 C5 C8 112.318 H2 C1 H3 107.842
H2 C1 H4 107.842 H2 C1 C5 111.166
H3 C1 H4 107.749 H3 C1 C5 111.043
H4 C1 C5 111.043 C5 C8 H9 111.215
C5 C8 H10 111.215 C5 C8 C11 113.720
H6 C5 H7 105.958 H6 C5 C8 108.910
H7 C5 C8 108.910 C8 C11 H12 115.370
C8 C11 O13 124.231 H9 C8 H10 105.189
H9 C8 C11 107.525 H10 C8 C11 107.525
H12 C11 O13 120.399
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability