return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H9N (Pyrrolidine)

using model chemistry: CCD/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCD/6-31G**
 hartrees
Energy at 0K-211.951671
Energy at 298.15K-211.963264
HF Energy-211.160347
Nuclear repulsion energy190.490768
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3612 3386 0.82      
2 A' 3207 3007 60.95      
3 A' 3170 2972 10.25      
4 A' 3151 2954 41.75      
5 A' 3037 2848 117.01      
6 A' 1594 1495 0.68      
7 A' 1569 1471 2.25      
8 A' 1452 1361 3.66      
9 A' 1371 1286 1.06      
10 A' 1292 1211 0.76      
11 A' 1270 1191 8.84      
12 A' 1108 1039 0.59      
13 A' 1037 973 6.16      
14 A' 978 917 3.39      
15 A' 937 879 9.17      
16 A' 921 863 59.65      
17 A' 794 744 21.78      
18 A' 595 558 39.57      
19 A' 305 286 4.46      
20 A" 3189 2990 1.28      
21 A" 3165 2968 66.35      
22 A" 3142 2946 17.60      
23 A" 3033 2844 36.61      
24 A" 1577 1479 3.03      
25 A" 1547 1450 0.43      
26 A" 1484 1391 4.46      
27 A" 1374 1289 12.31      
28 A" 1351 1266 14.30      
29 A" 1289 1208 6.72      
30 A" 1241 1164 0.58      
31 A" 1173 1099 9.50      
32 A" 1139 1067 0.18      
33 A" 967 907 0.05      
34 A" 896 840 2.07      
35 A" 643 603 0.76      
36 A" 65 61 0.18      

Unscaled Zero Point Vibrational Energy (zpe) 29336.1 cm-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 27505.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/6-31G**
ABC
0.23008 0.22769 0.13074

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.546 -1.092 0.000
H2 0.417 -2.097 0.000
C3 -0.105 -0.469 1.155
C4 -0.105 -0.469 -1.155
C5 -0.105 1.021 0.775
C6 -0.105 1.021 -0.775
H7 -1.139 -0.816 1.296
H8 -1.139 -0.816 -1.296
H9 0.447 -0.676 2.073
H10 0.447 -0.676 -2.073
H11 0.793 1.506 1.157
H12 0.793 1.506 -1.157
H13 -0.965 1.542 1.195
H14 -0.965 1.542 -1.195

Atom - Atom Distances (Å)
  N1 H2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
N11.01371.46511.46512.34262.34262.14342.14342.11632.11632.85492.85493.26293.2629
H21.01372.06402.06403.25533.25532.39622.39622.51332.51333.80363.80364.07214.0721
C31.46512.06402.31091.53722.43851.10042.68341.09033.28152.16933.17072.18733.2102
C41.46512.06402.31092.43851.53722.68341.10043.28151.09033.17072.16933.21022.1873
C52.34263.25531.53722.43851.55072.17182.95562.20633.36091.08942.18511.08972.2118
C62.34263.25532.43851.53721.55072.95562.17183.36092.20632.18511.08942.21181.0897
H72.14342.39621.10042.68342.17182.95562.59181.77173.72603.02443.89162.36703.4343
H82.14342.39622.68341.10042.95562.17182.59183.72601.77173.89163.02443.43432.3670
H92.11632.51331.09033.28152.20633.36091.77173.72604.14542.39223.91342.77234.1939
H102.11632.51333.28151.09033.36092.20633.72601.77174.14543.91342.39224.19392.7723
H112.85493.80362.16933.17071.08942.18513.02443.89162.39223.91342.31381.75852.9360
H122.85493.80363.17072.16932.18511.08943.89163.02443.91342.39222.31382.93601.7585
H133.26294.07212.18733.21021.08972.21182.36703.43432.77234.19391.75852.93602.3890
H143.26294.07213.21022.18732.21181.08973.43432.36704.19392.77232.93601.75852.3890

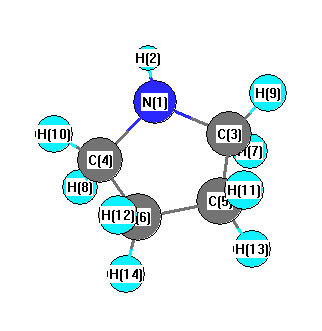
picture of Pyrrolidine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C3 C5 102.544 N1 C3 H7 112.552
N1 C3 H9 110.973 N1 C4 C6 102.544
N1 C4 H8 112.552 N1 C4 H10 110.973
H2 N1 C3 111.442 H2 N1 C4 111.442
C3 N1 C4 104.123 C3 C5 C6 104.316
C3 C5 H11 110.195 C3 C5 H13 111.609
C4 C6 C5 104.316 C4 C6 H12 110.195
C4 C6 H14 111.609 C5 C3 H7 109.741
C5 C3 H9 113.117 C5 C6 H12 110.505
C5 C6 H14 112.623 C6 C4 H8 109.741
C6 C4 H10 113.117 C6 C5 H11 110.505
C6 C5 H13 112.623 H7 C3 H9 107.939
H8 C4 H10 107.939 H11 C5 H13 107.608
H12 C6 H14 107.608
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability