return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3NHCH3 (Dimethylamine)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-134.737667
Energy at 298.15K-134.745927
HF Energy-134.251259
Nuclear repulsion energy84.428595
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3611 3374 0.15      
2 A' 3230 3018 36.21      
3 A' 3174 2966 57.85      
4 A' 3056 2855 110.79      
5 A' 1582 1478 0.60      
6 A' 1561 1459 8.70      
7 A' 1523 1423 0.48      
8 A' 1309 1223 0.69      
9 A' 1232 1151 8.88      
10 A' 981 917 1.24      
11 A' 829 775 122.44      
12 A' 398 372 5.18      
13 A' 287 268 2.70      
14 A" 3229 3017 19.59      
15 A" 3175 2967 18.60      
16 A" 3053 2853 42.58      
17 A" 1576 1472 8.66      
18 A" 1546 1445 6.53      
19 A" 1518 1419 8.32      
20 A" 1490 1392 1.68      
21 A" 1219 1139 29.47      
22 A" 1135 1061 0.48      
23 A" 1059 990 8.43      
24 A" 245 229 0.17      

Unscaled Zero Point Vibrational Energy (zpe) 21009.4 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 19631.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
1.14863 0.31525 0.27749

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.028 0.592 0.000
H2 -0.804 1.172 0.000
C3 0.028 -0.224 1.203
C4 0.028 -0.224 -1.203
H5 -0.788 -0.959 1.246
H6 -0.788 -0.959 -1.246
H7 0.967 -0.771 1.260
H8 0.967 -0.771 -1.260
H9 -0.040 0.417 2.079
H10 -0.040 0.417 -2.079

Atom - Atom Distances (Å)
  N1 H2 C3 C4 H5 H6 H7 H8 H9 H10
N11.01321.45411.45412.15022.15022.08102.08102.08742.0874
H21.01322.02162.02162.46802.46802.91542.91542.34002.3400
C31.45412.02162.40611.09812.68351.08902.69221.08743.3446
C41.45412.02162.40612.68351.09812.69221.08903.34461.0874
H52.15022.46801.09812.68352.49171.76503.06481.77343.6749
H62.15022.46802.68351.09812.49173.06481.76503.67491.7734
H72.08102.91541.08902.69221.76503.06482.51951.75983.6841
H82.08102.91542.69221.08903.06481.76502.51953.68411.7598
H92.08742.34001.08743.34461.77343.67491.75983.68414.1578
H102.08742.34003.34461.08743.67491.77343.68411.75984.1578

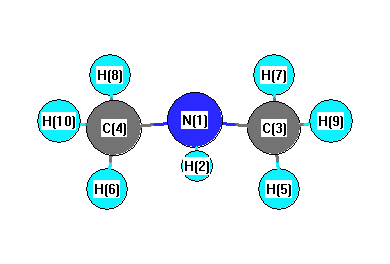
picture of Dimethylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C3 H5 114.079 N1 C3 H7 108.985
N1 C3 H9 109.593 N1 C4 H6 114.079
N1 C4 H8 108.985 N1 C4 H10 109.593
H2 N1 C3 108.727 H2 N1 C4 108.727
C3 N1 C4 111.653 H5 C3 H7 107.607
H5 C3 H9 108.474 H6 C4 H8 107.607
H6 C4 H10 108.474 H7 C3 H9 107.919
H8 C4 H10 107.919
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability