Jump to
S1C2
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -1194.096900 |
| Energy at 298.15K | -1194.099340 |
| HF Energy | -1193.692220 |
| Nuclear repulsion energy | 195.560235 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
2802 |
2618 |
0.48 |
|
|
|
| 2 |
A |
932 |
871 |
1.05 |
|
|
|
| 3 |
A |
508 |
475 |
1.07 |
|
|
|
| 4 |
A |
321 |
300 |
29.20 |
|
|
|
| 5 |
A |
214 |
200 |
0.05 |
|
|
|
| 6 |
B |
2802 |
2618 |
11.11 |
|
|
|
| 7 |
B |
916 |
856 |
16.79 |
|
|
|
| 8 |
B |
513 |
479 |
21.64 |
|
|
|
| 9 |
B |
354 |
330 |
21.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4680.6 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 4373.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.839 |
| S2 |
0.000 |
1.667 |
-0.383 |
| S3 |
0.000 |
-1.667 |
-0.383 |
| H4 |
-1.315 |
1.771 |
-0.580 |
| H5 |
1.315 |
-1.771 |
-0.580 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
H4 |
H5 |
| S1 | | 2.0671 | 2.0671 | 2.6234 | 2.6234 |
S2 | 2.0671 | | 3.3341 | 1.3342 | 3.6867 | S3 | 2.0671 | 3.3341 | | 3.6867 | 1.3342 | H4 | 2.6234 | 1.3342 | 3.6867 | | 4.4127 | H5 | 2.6234 | 3.6867 | 1.3342 | 4.4127 | |
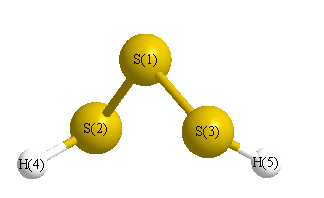 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
H4 |
98.644 |
|
S1 |
S3 |
H5 |
98.644 |
| S2 |
S1 |
S3 |
107.504 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -1194.096525 |
| Energy at 298.15K | -1194.098979 |
| HF Energy | -1193.691744 |
| Nuclear repulsion energy | 195.599814 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
2795 |
2612 |
16.85 |
|
|
|
| 2 |
A' |
933 |
871 |
9.72 |
|
|
|
| 3 |
A' |
508 |
475 |
0.99 |
|
|
|
| 4 |
A' |
353 |
330 |
22.00 |
|
|
|
| 5 |
A' |
215 |
201 |
0.23 |
|
|
|
| 6 |
A" |
2797 |
2614 |
0.62 |
|
|
|
| 7 |
A" |
924 |
864 |
11.12 |
|
|
|
| 8 |
A" |
514 |
481 |
23.54 |
|
|
|
| 9 |
A" |
329 |
308 |
12.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4684.5 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 4377.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
-0.053 |
0.836 |
0.000 |
| S2 |
-0.053 |
-0.385 |
1.667 |
| S3 |
-0.053 |
-0.385 |
-1.667 |
| H4 |
1.267 |
-0.524 |
1.813 |
| H5 |
1.267 |
-0.524 |
-1.813 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
H4 |
H5 |
| S1 | | 2.0664 | 2.0664 | 2.6230 | 2.6230 |
S2 | 2.0664 | | 3.3330 | 1.3348 | 3.7240 | S3 | 2.0664 | 3.3330 | | 3.7240 | 1.3348 | H4 | 2.6230 | 1.3348 | 3.7240 | | 3.6264 | H5 | 2.6230 | 3.7240 | 1.3348 | 3.6264 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
H4 |
98.639 |
|
S1 |
S3 |
H5 |
98.639 |
| S2 |
S1 |
S3 |
107.510 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
