Jump to
S1C2
Energy calculated at M06-2X/6-31G**
| | hartrees |
|---|
| Energy at 0K | -1195.650283 |
| Energy at 298.15K | -1195.652649 |
| HF Energy | -1195.650283 |
| Nuclear repulsion energy | 195.002112 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
2715 |
2580 |
0.42 |
|
|
|
| 2 |
A |
921 |
875 |
1.02 |
|
|
|
| 3 |
A |
495 |
471 |
0.77 |
|
|
|
| 4 |
A |
306 |
291 |
30.41 |
|
|
|
| 5 |
A |
198 |
188 |
0.01 |
|
|
|
| 6 |
B |
2714 |
2580 |
8.05 |
|
|
|
| 7 |
B |
902 |
857 |
13.33 |
|
|
|
| 8 |
B |
486 |
462 |
25.02 |
|
|
|
| 9 |
B |
357 |
340 |
24.19 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4546.8 cm
-1
Scaled (by 0.9504) Zero Point Vibrational Energy (zpe) 4321.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-31G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.848 |
| S2 |
0.000 |
1.667 |
-0.387 |
| S3 |
0.000 |
-1.667 |
-0.387 |
| H4 |
-1.326 |
1.751 |
-0.600 |
| H5 |
1.326 |
-1.751 |
-0.600 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
H4 |
H5 |
| S1 | | 2.0744 | 2.0744 | 2.6305 | 2.6305 |
S2 | 2.0744 | | 3.3336 | 1.3456 | 3.6717 | S3 | 2.0744 | 3.3336 | | 3.6717 | 1.3456 | H4 | 2.6305 | 1.3456 | 3.6717 | | 4.3920 | H5 | 2.6305 | 3.6717 | 1.3456 | 4.3920 | |
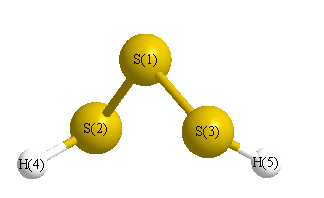 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
H4 |
98.297 |
|
S1 |
S3 |
H5 |
98.297 |
| S2 |
S1 |
S3 |
106.930 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.051 |
|
|
|
| 2 |
S |
-0.076 |
|
|
|
| 3 |
S |
-0.076 |
|
|
|
| 4 |
H |
0.102 |
|
|
|
| 5 |
H |
0.102 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.710 |
0.710 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.433 |
-4.392 |
0.000 |
| y |
-4.392 |
-40.116 |
0.000 |
| z |
0.000 |
0.000 |
-41.933 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.592 |
-4.392 |
0.000 |
| y |
-4.392 |
-0.433 |
0.000 |
| z |
0.000 |
0.000 |
-3.159 |
|
| Polar |
|---|
| 3z2-r2 | -6.317 |
|---|
| x2-y2 | 2.683 |
|---|
| xy | -4.392 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.544 |
-0.690 |
0.000 |
| y |
-0.690 |
10.880 |
0.000 |
| z |
0.000 |
0.000 |
6.012 |
<r2> (average value of r
2) Å
2
| <r2> |
140.438 |
| (<r2>)1/2 |
11.851 |
Jump to
S1C1
Energy calculated at M06-2X/6-31G**
| | hartrees |
|---|
| Energy at 0K | -1195.649793 |
| Energy at 298.15K | -1195.651824 |
| HF Energy | -1195.649793 |
| Nuclear repulsion energy | 194.977593 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
2658 |
2526 |
12.48 |
|
|
|
| 2 |
A' |
906 |
861 |
9.43 |
|
|
|
| 3 |
A' |
494 |
470 |
0.71 |
|
|
|
| 4 |
A' |
233 |
221 |
21.41 |
|
|
|
| 5 |
A' |
196 |
187 |
0.92 |
|
|
|
| 6 |
A" |
2659 |
2527 |
0.33 |
|
|
|
| 7 |
A" |
899 |
855 |
8.05 |
|
|
|
| 8 |
A" |
487 |
462 |
26.70 |
|
|
|
| 9 |
A" |
177 |
168 |
12.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4354.0 cm
-1
Scaled (by 0.9504) Zero Point Vibrational Energy (zpe) 4138.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
-0.053 |
0.843 |
0.000 |
| S2 |
-0.053 |
-0.391 |
1.667 |
| S3 |
-0.053 |
-0.391 |
-1.667 |
| H4 |
1.278 |
-0.495 |
1.829 |
| H5 |
1.278 |
-0.495 |
-1.829 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
H4 |
H5 |
| S1 | | 2.0739 | 2.0739 | 2.6285 | 2.6285 |
S2 | 2.0739 | | 3.3336 | 1.3454 | 3.7423 | S3 | 2.0739 | 3.3336 | | 3.7423 | 1.3454 | H4 | 2.6285 | 1.3454 | 3.7423 | | 3.6580 | H5 | 2.6285 | 3.7423 | 1.3454 | 3.6580 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
H4 |
98.219 |
|
S1 |
S3 |
H5 |
98.219 |
| S2 |
S1 |
S3 |
106.969 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.052 |
|
|
|
| 2 |
S |
-0.067 |
|
|
|
| 3 |
S |
-0.067 |
|
|
|
| 4 |
H |
0.093 |
|
|
|
| 5 |
H |
0.093 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.140 |
-0.536 |
0.000 |
2.206 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.672 |
-1.468 |
0.000 |
| y |
-1.468 |
-42.200 |
0.000 |
| z |
0.000 |
0.000 |
-39.626 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.241 |
-1.468 |
0.000 |
| y |
-1.468 |
-3.551 |
0.000 |
| z |
0.000 |
0.000 |
0.310 |
|
| Polar |
|---|
| 3z2-r2 | 0.620 |
|---|
| x2-y2 | 4.528 |
|---|
| xy | -1.468 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.517 |
-0.316 |
0.000 |
| y |
-0.316 |
5.942 |
0.000 |
| z |
0.000 |
0.000 |
10.965 |
<r2> (average value of r
2) Å
2
| <r2> |
140.631 |
| (<r2>)1/2 |
11.859 |
