Jump to
S1C2
S1C3
Vibrational Frequencies calculated at B2PLYP/6-31G**
Geometric Data calculated at B2PLYP/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at B2PLYP/6-31G**
Geometric Data calculated at B2PLYP/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at B2PLYP/6-31G**
| | hartrees |
|---|
| Energy at 0K | -210.174253 |
| Energy at 298.15K | -210.183465 |
| HF Energy | -209.962473 |
| Nuclear repulsion energy | 134.729694 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3748 |
3748 |
64.91 |
|
|
|
| 2 |
A |
3637 |
3637 |
1.40 |
|
|
|
| 3 |
A |
3542 |
3542 |
0.26 |
|
|
|
| 4 |
A |
3137 |
3137 |
43.41 |
|
|
|
| 5 |
A |
3125 |
3125 |
34.03 |
|
|
|
| 6 |
A |
3035 |
3035 |
74.15 |
|
|
|
| 7 |
A |
2997 |
2997 |
76.08 |
|
|
|
| 8 |
A |
1684 |
1684 |
29.38 |
|
|
|
| 9 |
A |
1553 |
1553 |
0.34 |
|
|
|
| 10 |
A |
1534 |
1534 |
4.15 |
|
|
|
| 11 |
A |
1474 |
1474 |
84.83 |
|
|
|
| 12 |
A |
1434 |
1434 |
17.49 |
|
|
|
| 13 |
A |
1402 |
1402 |
5.38 |
|
|
|
| 14 |
A |
1349 |
1349 |
2.65 |
|
|
|
| 15 |
A |
1278 |
1278 |
25.25 |
|
|
|
| 16 |
A |
1210 |
1210 |
9.13 |
|
|
|
| 17 |
A |
1129 |
1129 |
54.03 |
|
|
|
| 18 |
A |
1083 |
1083 |
32.57 |
|
|
|
| 19 |
A |
1026 |
1026 |
10.89 |
|
|
|
| 20 |
A |
953 |
953 |
65.40 |
|
|
|
| 21 |
A |
897 |
897 |
12.80 |
|
|
|
| 22 |
A |
845 |
845 |
86.92 |
|
|
|
| 23 |
A |
590 |
590 |
138.27 |
|
|
|
| 24 |
A |
544 |
544 |
12.94 |
|
|
|
| 25 |
A |
333 |
333 |
0.50 |
|
|
|
| 26 |
A |
278 |
278 |
14.37 |
|
|
|
| 27 |
A |
194 |
194 |
6.73 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22004.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 22004.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.350 |
-0.565 |
0.120 |
| C2 |
-0.634 |
0.657 |
-0.270 |
| C3 |
0.791 |
0.557 |
0.267 |
| O4 |
1.399 |
-0.641 |
-0.169 |
| H5 |
-1.643 |
-0.505 |
1.090 |
| H6 |
-2.191 |
-0.683 |
-0.433 |
| H7 |
-1.107 |
1.588 |
0.072 |
| H8 |
-0.593 |
0.684 |
-1.361 |
| H9 |
1.395 |
1.393 |
-0.094 |
| H10 |
0.768 |
0.619 |
1.367 |
| H11 |
0.687 |
-1.299 |
-0.103 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4695 | 2.4218 | 2.7652 | 1.0153 | 1.0132 | 2.1674 | 2.0799 | 3.3786 | 2.7287 | 2.1771 |
C2 | 1.4695 | | 1.5255 | 2.4138 | 2.0540 | 2.0613 | 1.0987 | 1.0921 | 2.1650 | 2.1557 | 2.3661 | C3 | 2.4218 | 1.5255 | | 1.4128 | 2.7802 | 3.3047 | 2.1683 | 2.1401 | 1.0924 | 1.1022 | 1.8952 | O4 | 2.7652 | 2.4138 | 1.4128 | | 3.2950 | 3.6000 | 3.3623 | 2.6724 | 2.0351 | 2.0846 | 0.9708 | H5 | 1.0153 | 2.0540 | 2.7802 | 3.2950 | | 1.6281 | 2.3886 | 2.9195 | 3.7727 | 2.6749 | 2.7359 | H6 | 1.0132 | 2.0613 | 3.3047 | 3.6000 | 1.6281 | | 2.5668 | 2.2990 | 4.1575 | 3.7005 | 2.9623 | H7 | 2.1674 | 1.0987 | 2.1683 | 3.3623 | 2.3886 | 2.5668 | | 1.7705 | 2.5149 | 2.4766 | 3.4033 | H8 | 2.0799 | 1.0921 | 2.1401 | 2.6724 | 2.9195 | 2.2990 | 1.7705 | | 2.4614 | 3.0494 | 2.6741 | H9 | 3.3786 | 2.1650 | 1.0924 | 2.0351 | 3.7727 | 4.1575 | 2.5149 | 2.4614 | | 1.7682 | 2.7826 | H10 | 2.7287 | 2.1557 | 1.1022 | 2.0846 | 2.6749 | 3.7005 | 2.4766 | 3.0494 | 1.7682 | | 2.4177 | H11 | 2.1771 | 2.3661 | 1.8952 | 0.9708 | 2.7359 | 2.9623 | 3.4033 | 2.6741 | 2.7826 | 2.4177 | |
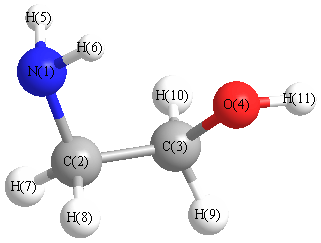 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
107.909 |
|
N1 |
C2 |
H7 |
114.351 |
| N1 |
C2 |
H8 |
107.668 |
|
C2 |
N1 |
H5 |
110.173 |
| C2 |
N1 |
H6 |
110.917 |
|
C2 |
C3 |
O4 |
110.410 |
| C2 |
C3 |
H9 |
110.492 |
|
C2 |
C3 |
H10 |
109.183 |
| C3 |
C2 |
H7 |
110.373 |
|
C3 |
C2 |
H8 |
108.550 |
| C3 |
O4 |
H11 |
103.777 |
|
O4 |
C3 |
H9 |
107.975 |
| O4 |
C3 |
H10 |
111.371 |
|
H5 |
N1 |
H6 |
106.760 |
| H7 |
C2 |
H8 |
107.828 |
|
H9 |
C3 |
H10 |
107.363 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.643 |
|
|
|
| 2 |
C |
-0.092 |
|
|
|
| 3 |
C |
0.088 |
|
|
|
| 4 |
O |
-0.568 |
|
|
|
| 5 |
H |
0.251 |
|
|
|
| 6 |
H |
0.256 |
|
|
|
| 7 |
H |
0.086 |
|
|
|
| 8 |
H |
0.120 |
|
|
|
| 9 |
H |
0.102 |
|
|
|
| 10 |
H |
0.072 |
|
|
|
| 11 |
H |
0.325 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.004 |
1.078 |
0.584 |
3.244 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.011 |
1.987 |
0.315 |
| y |
1.987 |
-25.352 |
-0.298 |
| z |
0.315 |
-0.298 |
-24.447 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.112 |
1.987 |
0.315 |
| y |
1.987 |
-0.123 |
-0.298 |
| z |
0.315 |
-0.298 |
1.235 |
|
| Polar |
|---|
| 3z2-r2 | 2.470 |
|---|
| x2-y2 | -0.659 |
|---|
| xy | 1.987 |
|---|
| xz | 0.315 |
|---|
| yz | -0.298 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.153 |
0.286 |
-0.019 |
| y |
0.286 |
4.924 |
0.049 |
| z |
-0.019 |
0.049 |
4.562 |
<r2> (average value of r
2) Å
2
| <r2> |
86.459 |
| (<r2>)1/2 |
9.298 |
