Jump to
S1C2
S1C3
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
Geometric Data calculated at HSEh1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
Geometric Data calculated at HSEh1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -210.171872 |
| Energy at 298.15K | -210.181104 |
| HF Energy | -210.171872 |
| Nuclear repulsion energy | 135.312856 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3724 |
3554 |
85.55 |
|
|
|
| 2 |
A |
3649 |
3482 |
3.18 |
|
|
|
| 3 |
A |
3551 |
3389 |
0.55 |
|
|
|
| 4 |
A |
3115 |
2973 |
40.06 |
|
|
|
| 5 |
A |
3102 |
2960 |
36.71 |
|
|
|
| 6 |
A |
3015 |
2877 |
73.32 |
|
|
|
| 7 |
A |
2964 |
2829 |
82.86 |
|
|
|
| 8 |
A |
1668 |
1592 |
32.93 |
|
|
|
| 9 |
A |
1528 |
1458 |
0.22 |
|
|
|
| 10 |
A |
1506 |
1438 |
8.40 |
|
|
|
| 11 |
A |
1469 |
1402 |
99.25 |
|
|
|
| 12 |
A |
1416 |
1352 |
12.34 |
|
|
|
| 13 |
A |
1387 |
1324 |
2.21 |
|
|
|
| 14 |
A |
1336 |
1275 |
2.82 |
|
|
|
| 15 |
A |
1266 |
1208 |
24.50 |
|
|
|
| 16 |
A |
1206 |
1151 |
3.37 |
|
|
|
| 17 |
A |
1143 |
1090 |
76.50 |
|
|
|
| 18 |
A |
1098 |
1047 |
26.81 |
|
|
|
| 19 |
A |
1016 |
969 |
7.67 |
|
|
|
| 20 |
A |
942 |
899 |
59.66 |
|
|
|
| 21 |
A |
901 |
859 |
7.52 |
|
|
|
| 22 |
A |
833 |
795 |
99.16 |
|
|
|
| 23 |
A |
617 |
589 |
132.92 |
|
|
|
| 24 |
A |
548 |
523 |
9.59 |
|
|
|
| 25 |
A |
333 |
318 |
0.63 |
|
|
|
| 26 |
A |
284 |
271 |
13.92 |
|
|
|
| 27 |
A |
197 |
188 |
7.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21906.6 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 20905.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.336 |
-0.560 |
0.118 |
| C2 |
-0.631 |
0.663 |
-0.261 |
| C3 |
0.798 |
0.554 |
0.258 |
| O4 |
1.376 |
-0.650 |
-0.167 |
| H5 |
-1.623 |
-0.519 |
1.091 |
| H6 |
-2.175 |
-0.687 |
-0.434 |
| H7 |
-1.101 |
1.593 |
0.095 |
| H8 |
-0.600 |
0.703 |
-1.355 |
| H9 |
1.406 |
1.384 |
-0.119 |
| H10 |
0.789 |
0.634 |
1.361 |
| H11 |
0.642 |
-1.284 |
-0.112 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4612 | 2.4112 | 2.7287 | 1.0149 | 1.0126 | 2.1657 | 2.0752 | 3.3691 | 2.7362 | 2.1188 |
C2 | 1.4612 | | 1.5238 | 2.4001 | 2.0510 | 2.0587 | 1.1017 | 1.0953 | 2.1649 | 2.1559 | 2.3304 | C3 | 2.4112 | 1.5238 | | 1.4018 | 2.7755 | 3.2954 | 2.1711 | 2.1397 | 1.0956 | 1.1063 | 1.8814 | O4 | 2.7287 | 2.4001 | 1.4018 | | 3.2545 | 3.5616 | 3.3525 | 2.6739 | 2.0352 | 2.0808 | 0.9716 | H5 | 1.0149 | 2.0510 | 2.7755 | 3.2545 | | 1.6309 | 2.3924 | 2.9190 | 3.7754 | 2.6865 | 2.6758 | H6 | 1.0126 | 2.0587 | 3.2954 | 3.5616 | 1.6309 | | 2.5757 | 2.2941 | 4.1487 | 3.7090 | 2.8973 | H7 | 2.1657 | 1.1017 | 2.1711 | 3.3525 | 2.3924 | 2.5757 | | 1.7734 | 2.5245 | 2.4692 | 3.3703 | H8 | 2.0752 | 1.0953 | 2.1397 | 2.6739 | 2.9190 | 2.2941 | 1.7734 | | 2.4522 | 3.0517 | 2.6525 | H9 | 3.3691 | 2.1649 | 1.0956 | 2.0352 | 3.7754 | 4.1487 | 2.5245 | 2.4522 | | 1.7704 | 2.7753 | H10 | 2.7362 | 2.1559 | 1.1063 | 2.0808 | 2.6865 | 3.7090 | 2.4692 | 3.0517 | 1.7704 | | 2.4232 | H11 | 2.1188 | 2.3304 | 1.8814 | 0.9716 | 2.6758 | 2.8973 | 3.3703 | 2.6525 | 2.7753 | 2.4232 | |
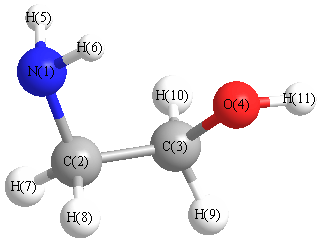 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
107.736 |
|
N1 |
C2 |
H7 |
114.620 |
| N1 |
C2 |
H8 |
107.673 |
|
C2 |
N1 |
H5 |
110.551 |
| C2 |
N1 |
H6 |
111.354 |
|
C2 |
C3 |
O4 |
110.172 |
| C2 |
C3 |
H9 |
110.405 |
|
C2 |
C3 |
H10 |
109.075 |
| C3 |
C2 |
H7 |
110.534 |
|
C3 |
C2 |
H8 |
108.452 |
| C3 |
O4 |
H11 |
103.383 |
|
O4 |
C3 |
H9 |
108.535 |
| O4 |
C3 |
H10 |
111.576 |
|
H5 |
N1 |
H6 |
107.101 |
| H7 |
C2 |
H8 |
107.645 |
|
H9 |
C3 |
H10 |
107.032 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.661 |
|
|
|
| 2 |
C |
-0.152 |
|
|
|
| 3 |
C |
0.035 |
|
|
|
| 4 |
O |
-0.558 |
|
|
|
| 5 |
H |
0.266 |
|
|
|
| 6 |
H |
0.271 |
|
|
|
| 7 |
H |
0.109 |
|
|
|
| 8 |
H |
0.144 |
|
|
|
| 9 |
H |
0.124 |
|
|
|
| 10 |
H |
0.092 |
|
|
|
| 11 |
H |
0.330 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.057 |
1.050 |
0.557 |
3.280 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.622 |
2.192 |
0.263 |
| y |
2.192 |
-25.197 |
-0.333 |
| z |
0.263 |
-0.333 |
-24.208 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.919 |
2.192 |
0.263 |
| y |
2.192 |
-0.282 |
-0.333 |
| z |
0.263 |
-0.333 |
1.201 |
|
| Polar |
|---|
| 3z2-r2 | 2.402 |
|---|
| x2-y2 | -0.425 |
|---|
| xy | 2.192 |
|---|
| xz | 0.263 |
|---|
| yz | -0.333 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.217 |
0.279 |
-0.005 |
| y |
0.279 |
4.982 |
0.052 |
| z |
-0.005 |
0.052 |
4.615 |
<r2> (average value of r
2) Å
2
| <r2> |
85.445 |
| (<r2>)1/2 |
9.244 |
