Jump to
S1C2
S1C3
Vibrational Frequencies calculated at B2PLYP=FULL/6-31G**
Geometric Data calculated at B2PLYP=FULL/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at B2PLYP=FULL/6-31G**
Geometric Data calculated at B2PLYP=FULL/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at B2PLYP=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -210.179171 |
| Energy at 298.15K | -210.188386 |
| HF Energy | -209.962497 |
| Nuclear repulsion energy | 134.771905 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3749 |
3749 |
65.08 |
|
|
|
| 2 |
A |
3638 |
3638 |
1.43 |
|
|
|
| 3 |
A |
3543 |
3543 |
0.26 |
|
|
|
| 4 |
A |
3138 |
3138 |
43.22 |
|
|
|
| 5 |
A |
3126 |
3126 |
34.05 |
|
|
|
| 6 |
A |
3035 |
3035 |
74.02 |
|
|
|
| 7 |
A |
2998 |
2998 |
76.08 |
|
|
|
| 8 |
A |
1685 |
1685 |
29.48 |
|
|
|
| 9 |
A |
1554 |
1554 |
0.35 |
|
|
|
| 10 |
A |
1534 |
1534 |
4.17 |
|
|
|
| 11 |
A |
1475 |
1475 |
84.71 |
|
|
|
| 12 |
A |
1435 |
1435 |
17.38 |
|
|
|
| 13 |
A |
1402 |
1402 |
5.46 |
|
|
|
| 14 |
A |
1350 |
1350 |
2.64 |
|
|
|
| 15 |
A |
1278 |
1278 |
25.42 |
|
|
|
| 16 |
A |
1210 |
1210 |
9.08 |
|
|
|
| 17 |
A |
1130 |
1130 |
54.32 |
|
|
|
| 18 |
A |
1084 |
1084 |
32.49 |
|
|
|
| 19 |
A |
1026 |
1026 |
10.82 |
|
|
|
| 20 |
A |
954 |
954 |
65.21 |
|
|
|
| 21 |
A |
897 |
897 |
12.81 |
|
|
|
| 22 |
A |
845 |
845 |
87.40 |
|
|
|
| 23 |
A |
591 |
591 |
138.38 |
|
|
|
| 24 |
A |
544 |
544 |
12.79 |
|
|
|
| 25 |
A |
333 |
333 |
0.50 |
|
|
|
| 26 |
A |
279 |
279 |
14.37 |
|
|
|
| 27 |
A |
195 |
195 |
6.73 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22012.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 22012.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.350 |
-0.565 |
0.120 |
| C2 |
-0.633 |
0.657 |
-0.270 |
| C3 |
0.791 |
0.557 |
0.267 |
| O4 |
1.398 |
-0.641 |
-0.169 |
| H5 |
-1.642 |
-0.505 |
1.090 |
| H6 |
-2.191 |
-0.683 |
-0.433 |
| H7 |
-1.106 |
1.588 |
0.071 |
| H8 |
-0.592 |
0.683 |
-1.361 |
| H9 |
1.395 |
1.392 |
-0.094 |
| H10 |
0.768 |
0.619 |
1.367 |
| H11 |
0.687 |
-1.298 |
-0.104 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4690 | 2.4210 | 2.7643 | 1.0150 | 1.0129 | 2.1668 | 2.0793 | 3.3775 | 2.7276 | 2.1764 |
C2 | 1.4690 | | 1.5250 | 2.4130 | 2.0535 | 2.0609 | 1.0984 | 1.0918 | 2.1643 | 2.1549 | 2.3651 | C3 | 2.4210 | 1.5250 | | 1.4124 | 2.7792 | 3.3037 | 2.1676 | 2.1393 | 1.0920 | 1.1018 | 1.8947 | O4 | 2.7643 | 2.4130 | 1.4124 | | 3.2939 | 3.5989 | 3.3612 | 2.6712 | 2.0346 | 2.0840 | 0.9706 | H5 | 1.0150 | 2.0535 | 2.7792 | 3.2939 | | 1.6278 | 2.3883 | 2.9189 | 3.7714 | 2.6735 | 2.7349 | H6 | 1.0129 | 2.0609 | 3.3037 | 3.5989 | 1.6278 | | 2.5664 | 2.2987 | 4.1565 | 3.6992 | 2.9611 | H7 | 2.1668 | 1.0984 | 2.1676 | 3.3612 | 2.3883 | 2.5664 | | 1.7699 | 2.5140 | 2.4759 | 3.4022 | H8 | 2.0793 | 1.0918 | 2.1393 | 2.6712 | 2.9189 | 2.2987 | 1.7699 | | 2.4607 | 3.0484 | 2.6727 | H9 | 3.3775 | 2.1643 | 1.0920 | 2.0346 | 3.7714 | 4.1565 | 2.5140 | 2.4607 | | 1.7677 | 2.7819 | H10 | 2.7276 | 2.1549 | 1.1018 | 2.0840 | 2.6735 | 3.6992 | 2.4759 | 3.0484 | 1.7677 | | 2.4171 | H11 | 2.1764 | 2.3651 | 1.8947 | 0.9706 | 2.7349 | 2.9611 | 3.4022 | 2.6727 | 2.7819 | 2.4171 | |
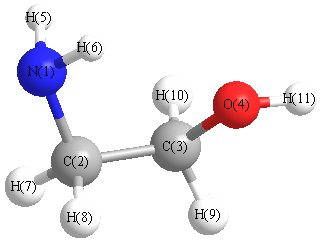 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
107.906 |
|
N1 |
C2 |
H7 |
114.354 |
| N1 |
C2 |
H8 |
107.671 |
|
C2 |
N1 |
H5 |
110.186 |
| C2 |
N1 |
H6 |
110.939 |
|
C2 |
C3 |
O4 |
110.406 |
| C2 |
C3 |
H9 |
110.494 |
|
C2 |
C3 |
H10 |
109.178 |
| C3 |
C2 |
H7 |
110.376 |
|
C3 |
C2 |
H8 |
108.544 |
| C3 |
O4 |
H11 |
103.776 |
|
O4 |
C3 |
H9 |
107.981 |
| O4 |
C3 |
H10 |
111.373 |
|
H5 |
N1 |
H6 |
106.776 |
| H7 |
C2 |
H8 |
107.827 |
|
H9 |
C3 |
H10 |
107.362 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
