Jump to
S1C2
S1C3
Vibrational Frequencies calculated at CCD/6-31G**
Geometric Data calculated at CCD/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at CCD/6-31G**
Geometric Data calculated at CCD/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at CCD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -209.803773 |
| Energy at 298.15K | -209.813010 |
| HF Energy | -209.117744 |
| Nuclear repulsion energy | 134.659413 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3863 |
3622 |
53.80 |
|
|
|
| 2 |
A |
3681 |
3451 |
0.84 |
|
|
|
| 3 |
A |
3586 |
3362 |
0.21 |
|
|
|
| 4 |
A |
3177 |
2978 |
45.98 |
|
|
|
| 5 |
A |
3167 |
2969 |
30.72 |
|
|
|
| 6 |
A |
3079 |
2887 |
66.71 |
|
|
|
| 7 |
A |
3052 |
2862 |
65.17 |
|
|
|
| 8 |
A |
1712 |
1605 |
31.28 |
|
|
|
| 9 |
A |
1581 |
1482 |
0.18 |
|
|
|
| 10 |
A |
1562 |
1465 |
2.99 |
|
|
|
| 11 |
A |
1499 |
1406 |
69.79 |
|
|
|
| 12 |
A |
1460 |
1369 |
26.28 |
|
|
|
| 13 |
A |
1426 |
1337 |
8.07 |
|
|
|
| 14 |
A |
1371 |
1286 |
2.37 |
|
|
|
| 15 |
A |
1298 |
1217 |
24.50 |
|
|
|
| 16 |
A |
1226 |
1150 |
8.08 |
|
|
|
| 17 |
A |
1158 |
1085 |
52.71 |
|
|
|
| 18 |
A |
1106 |
1037 |
29.72 |
|
|
|
| 19 |
A |
1045 |
980 |
17.79 |
|
|
|
| 20 |
A |
980 |
919 |
67.11 |
|
|
|
| 21 |
A |
914 |
857 |
14.25 |
|
|
|
| 22 |
A |
868 |
813 |
76.51 |
|
|
|
| 23 |
A |
568 |
532 |
116.77 |
|
|
|
| 24 |
A |
537 |
503 |
48.30 |
|
|
|
| 25 |
A |
338 |
317 |
0.56 |
|
|
|
| 26 |
A |
277 |
259 |
14.49 |
|
|
|
| 27 |
A |
198 |
186 |
5.75 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22362.8 cm
-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 20967.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.360 |
-0.566 |
0.121 |
| C2 |
-0.634 |
0.647 |
-0.278 |
| C3 |
0.782 |
0.555 |
0.275 |
| O4 |
1.414 |
-0.628 |
-0.171 |
| H5 |
-1.651 |
-0.489 |
1.089 |
| H6 |
-2.205 |
-0.670 |
-0.426 |
| H7 |
-1.108 |
1.581 |
0.047 |
| H8 |
-0.579 |
0.661 |
-1.367 |
| H9 |
1.383 |
1.399 |
-0.068 |
| H10 |
0.747 |
0.598 |
1.373 |
| H11 |
0.730 |
-1.308 |
-0.101 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4692 | 2.4226 | 2.7895 | 1.0141 | 1.0121 | 2.1635 | 2.0807 | 3.3789 | 2.7133 | 2.2288 |
C2 | 1.4692 | | 1.5231 | 2.4150 | 2.0483 | 2.0559 | 1.0968 | 1.0907 | 2.1622 | 2.1531 | 2.3907 | C3 | 2.4226 | 1.5231 | | 1.4140 | 2.7703 | 3.3042 | 2.1633 | 2.1357 | 1.0905 | 1.0993 | 1.9013 | O4 | 2.7895 | 2.4150 | 1.4140 | | 3.3169 | 3.6282 | 3.3604 | 2.6579 | 2.0303 | 2.0815 | 0.9661 | H5 | 1.0141 | 2.0483 | 2.7703 | 3.3169 | | 1.6230 | 2.3808 | 2.9164 | 3.7557 | 2.6485 | 2.7853 | H6 | 1.0121 | 2.0559 | 3.3042 | 3.6282 | 1.6230 | | 2.5486 | 2.3027 | 4.1571 | 3.6824 | 3.0216 | H7 | 2.1635 | 1.0968 | 2.1633 | 3.3604 | 2.3808 | 2.5486 | | 1.7682 | 2.5003 | 2.4840 | 3.4279 | H8 | 2.0807 | 1.0907 | 2.1357 | 2.6579 | 2.9164 | 2.3027 | 1.7682 | | 2.4662 | 3.0449 | 2.6821 | H9 | 3.3789 | 2.1622 | 1.0905 | 2.0303 | 3.7557 | 4.1571 | 2.5003 | 2.4662 | | 1.7663 | 2.7843 | H10 | 2.7133 | 2.1531 | 1.0993 | 2.0815 | 2.6485 | 3.6824 | 2.4840 | 3.0449 | 1.7663 | | 2.4091 | H11 | 2.2288 | 2.3907 | 1.9013 | 0.9661 | 2.7853 | 3.0216 | 3.4279 | 2.6821 | 2.7843 | 2.4091 | |
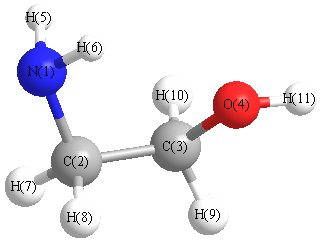 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
108.104 |
|
N1 |
C2 |
H7 |
114.164 |
| N1 |
C2 |
H8 |
107.826 |
|
C2 |
N1 |
H5 |
109.779 |
| C2 |
N1 |
H6 |
110.544 |
|
C2 |
C3 |
O4 |
110.568 |
| C2 |
C3 |
H9 |
110.550 |
|
C2 |
C3 |
H10 |
109.313 |
| C3 |
C2 |
H7 |
110.262 |
|
C3 |
C2 |
H8 |
108.460 |
| C3 |
O4 |
H11 |
104.461 |
|
O4 |
C3 |
H9 |
107.616 |
| O4 |
C3 |
H10 |
111.212 |
|
H5 |
N1 |
H6 |
106.446 |
| H7 |
C2 |
H8 |
107.867 |
|
H9 |
C3 |
H10 |
107.529 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
