Jump to
S1C2
S1C3
Vibrational Frequencies calculated at TPSSh/6-31G**
Geometric Data calculated at TPSSh/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at TPSSh/6-31G**
Geometric Data calculated at TPSSh/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at TPSSh/6-31G**
| | hartrees |
|---|
| Energy at 0K | -210.398637 |
| Energy at 298.15K | -210.407834 |
| HF Energy | -210.398637 |
| Nuclear repulsion energy | 134.696729 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3618 |
3468 |
80.22 |
|
|
|
| 2 |
A |
3578 |
3430 |
0.46 |
|
|
|
| 3 |
A |
3486 |
3342 |
0.50 |
|
|
|
| 4 |
A |
3096 |
2969 |
46.93 |
|
|
|
| 5 |
A |
3082 |
2955 |
42.12 |
|
|
|
| 6 |
A |
2992 |
2869 |
85.36 |
|
|
|
| 7 |
A |
2942 |
2820 |
91.87 |
|
|
|
| 8 |
A |
1672 |
1603 |
26.85 |
|
|
|
| 9 |
A |
1539 |
1475 |
0.56 |
|
|
|
| 10 |
A |
1517 |
1455 |
4.75 |
|
|
|
| 11 |
A |
1463 |
1402 |
100.86 |
|
|
|
| 12 |
A |
1413 |
1355 |
11.22 |
|
|
|
| 13 |
A |
1380 |
1323 |
2.51 |
|
|
|
| 14 |
A |
1330 |
1275 |
2.70 |
|
|
|
| 15 |
A |
1259 |
1207 |
25.04 |
|
|
|
| 16 |
A |
1193 |
1144 |
8.53 |
|
|
|
| 17 |
A |
1107 |
1061 |
57.67 |
|
|
|
| 18 |
A |
1064 |
1020 |
29.49 |
|
|
|
| 19 |
A |
1007 |
965 |
10.46 |
|
|
|
| 20 |
A |
939 |
900 |
67.83 |
|
|
|
| 21 |
A |
883 |
846 |
8.73 |
|
|
|
| 22 |
A |
831 |
797 |
79.82 |
|
|
|
| 23 |
A |
625 |
599 |
129.05 |
|
|
|
| 24 |
A |
536 |
514 |
8.40 |
|
|
|
| 25 |
A |
324 |
311 |
0.79 |
|
|
|
| 26 |
A |
284 |
272 |
13.55 |
|
|
|
| 27 |
A |
195 |
187 |
7.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21677.2 cm
-1
Scaled (by 0.9587) Zero Point Vibrational Energy (zpe) 20781.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.335 |
-0.567 |
0.120 |
| C2 |
-0.633 |
0.672 |
-0.263 |
| C3 |
0.803 |
0.562 |
0.260 |
| O4 |
1.376 |
-0.659 |
-0.169 |
| H5 |
-1.633 |
-0.515 |
1.093 |
| H6 |
-2.177 |
-0.692 |
-0.437 |
| H7 |
-1.112 |
1.596 |
0.098 |
| H8 |
-0.606 |
0.710 |
-1.356 |
| H9 |
1.419 |
1.381 |
-0.125 |
| H10 |
0.797 |
0.640 |
1.363 |
| H11 |
0.626 |
-1.285 |
-0.106 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4744 | 2.4211 | 2.7272 | 1.0190 | 1.0168 | 2.1744 | 2.0827 | 3.3815 | 2.7473 | 2.1002 |
C2 | 1.4744 | | 1.5320 | 2.4117 | 2.0605 | 2.0672 | 1.1015 | 1.0946 | 2.1750 | 2.1653 | 2.3325 | C3 | 2.4211 | 1.5320 | | 1.4148 | 2.7900 | 3.3065 | 2.1824 | 2.1494 | 1.0949 | 1.1059 | 1.8907 | O4 | 2.7272 | 2.4117 | 1.4148 | | 3.2652 | 3.5626 | 3.3684 | 2.6857 | 2.0412 | 2.0901 | 0.9786 | H5 | 1.0190 | 2.0605 | 2.7900 | 3.2652 | | 1.6326 | 2.3908 | 2.9241 | 3.7931 | 2.7040 | 2.6703 | H6 | 1.0168 | 2.0672 | 3.3065 | 3.5626 | 1.6326 | | 2.5793 | 2.2967 | 4.1616 | 3.7223 | 2.8838 | H7 | 2.1744 | 1.1015 | 2.1824 | 3.3684 | 2.3908 | 2.5793 | | 1.7771 | 2.5495 | 2.4813 | 3.3708 | H8 | 2.0827 | 1.0946 | 2.1494 | 2.6857 | 2.9241 | 2.2967 | 1.7771 | | 2.4634 | 3.0608 | 2.6572 | H9 | 3.3815 | 2.1750 | 1.0949 | 2.0412 | 3.7931 | 4.1616 | 2.5495 | 2.4634 | | 1.7743 | 2.7815 | H10 | 2.7473 | 2.1653 | 1.1059 | 2.0901 | 2.7040 | 3.7223 | 2.4813 | 3.0608 | 1.7743 | | 2.4277 | H11 | 2.1002 | 2.3325 | 1.8907 | 0.9786 | 2.6703 | 2.8838 | 3.3708 | 2.6572 | 2.7815 | 2.4277 | |
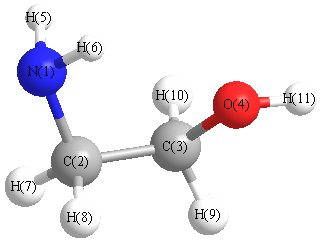 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
107.269 |
|
N1 |
C2 |
H7 |
114.383 |
| N1 |
C2 |
H8 |
107.409 |
|
C2 |
N1 |
H5 |
110.122 |
| C2 |
N1 |
H6 |
110.817 |
|
C2 |
C3 |
O4 |
109.791 |
| C2 |
C3 |
H9 |
110.681 |
|
C2 |
C3 |
H10 |
109.279 |
| C3 |
C2 |
H7 |
110.878 |
|
C3 |
C2 |
H8 |
108.690 |
| C3 |
O4 |
H11 |
102.840 |
|
O4 |
C3 |
H9 |
108.168 |
| O4 |
C3 |
H10 |
111.441 |
|
H5 |
N1 |
H6 |
106.641 |
| H7 |
C2 |
H8 |
108.040 |
|
H9 |
C3 |
H10 |
107.458 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.655 |
|
|
|
| 2 |
C |
-0.134 |
|
|
|
| 3 |
C |
0.047 |
|
|
|
| 4 |
O |
-0.560 |
|
|
|
| 5 |
H |
0.262 |
|
|
|
| 6 |
H |
0.266 |
|
|
|
| 7 |
H |
0.103 |
|
|
|
| 8 |
H |
0.139 |
|
|
|
| 9 |
H |
0.118 |
|
|
|
| 10 |
H |
0.087 |
|
|
|
| 11 |
H |
0.328 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.055 |
1.126 |
0.555 |
3.302 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.357 |
0.277 |
-0.011 |
| y |
0.277 |
5.078 |
0.061 |
| z |
-0.011 |
0.061 |
4.674 |
<r2> (average value of r
2) Å
2
| <r2> |
85.926 |
| (<r2>)1/2 |
9.270 |
