Jump to
S1C2
S1C3
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -254.912005 |
| Energy at 298.15K | -254.911402 |
| HF Energy | -254.912005 |
| Nuclear repulsion energy | 46.675017 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.679 |
| N2 |
0.000 |
0.000 |
-1.849 |
| Na3 |
0.000 |
0.000 |
1.548 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
Na3 |
| C1 | | 1.1700 | 2.2270 |
N2 | 1.1700 | | 3.3969 | Na3 | 2.2270 | 3.3969 | |
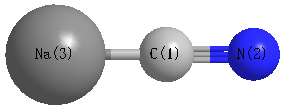 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
Na3 |
0.000 |
|
C1 |
Na3 |
N2 |
0.000 |
| N2 |
C1 |
Na3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.111 |
|
|
|
| 2 |
N |
-0.461 |
|
|
|
| 3 |
Na |
0.571 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
10.323 |
10.323 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.029 |
0.000 |
0.000 |
| y |
0.000 |
-17.029 |
0.000 |
| z |
0.000 |
0.000 |
-12.877 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.076 |
0.000 |
0.000 |
| y |
0.000 |
-2.076 |
0.000 |
| z |
0.000 |
0.000 |
4.152 |
|
| Polar |
|---|
| 3z2-r2 | 8.304 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.699 |
0.000 |
0.000 |
| y |
0.000 |
2.699 |
0.000 |
| z |
0.000 |
0.000 |
5.550 |
<r2> (average value of r
2) Å
2
| <r2> |
62.827 |
| (<r2>)1/2 |
7.926 |
Jump to
S1C1
S1C3
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -254.917433 |
| Energy at 298.15K | -254.917191 |
| HF Energy | -254.917433 |
| Nuclear repulsion energy | 51.884903 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.112 |
0.670 |
0.000 |
| N2 |
0.000 |
1.072 |
0.000 |
| Na3 |
-0.607 |
-1.048 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
Na3 |
| C1 | | 1.1826 | 2.4294 |
N2 | 1.1826 | | 2.2049 | Na3 | 2.4294 | 2.2049 | |
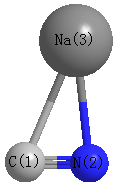 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
Na3 |
86.059 |
|
C1 |
Na3 |
N2 |
29.055 |
| N2 |
C1 |
Na3 |
64.886 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.053 |
|
|
|
| 2 |
N |
-0.447 |
|
|
|
| 3 |
Na |
0.500 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.447 |
-7.153 |
0.000 |
8.423 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.259 |
3.611 |
0.000 |
| y |
3.611 |
-14.070 |
0.000 |
| z |
0.000 |
0.000 |
-17.326 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.562 |
3.611 |
0.000 |
| y |
3.611 |
4.223 |
0.000 |
| z |
0.000 |
0.000 |
-0.661 |
|
| Polar |
|---|
| 3z2-r2 | -1.322 |
|---|
| x2-y2 | -5.190 |
|---|
| xy | 3.611 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.886 |
-0.100 |
0.000 |
| y |
-0.100 |
3.545 |
0.000 |
| z |
0.000 |
0.000 |
2.819 |
<r2> (average value of r
2) Å
2
| <r2> |
44.277 |
| (<r2>)1/2 |
6.654 |
Jump to
S1C1
S1C2
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -254.910392 |
| Energy at 298.15K | |
| HF Energy | -254.910392 |
| Nuclear repulsion energy | 48.892342 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.847 |
| N2 |
0.000 |
0.000 |
-0.667 |
| Na3 |
0.000 |
0.000 |
1.432 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
Na3 |
| C1 | | 1.1808 | 3.2791 |
N2 | 1.1808 | | 2.0983 | Na3 | 3.2791 | 2.0983 | |
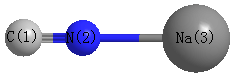 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
Na3 |
180.000 |
|
C1 |
Na3 |
N2 |
0.000 |
| N2 |
C1 |
Na3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.125 |
|
|
|
| 2 |
N |
-0.528 |
|
|
|
| 3 |
Na |
0.654 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
10.342 |
10.342 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.969 |
0.000 |
0.000 |
| y |
0.000 |
-16.969 |
0.000 |
| z |
0.000 |
0.000 |
-14.972 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.999 |
0.000 |
0.000 |
| y |
0.000 |
-0.999 |
0.000 |
| z |
0.000 |
0.000 |
1.997 |
|
| Polar |
|---|
| 3z2-r2 | 3.995 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.583 |
0.000 |
0.000 |
| y |
0.000 |
2.583 |
0.000 |
| z |
0.000 |
0.000 |
5.469 |
<r2> (average value of r
2) Å
2
| <r2> |
56.318 |
| (<r2>)1/2 |
7.505 |
