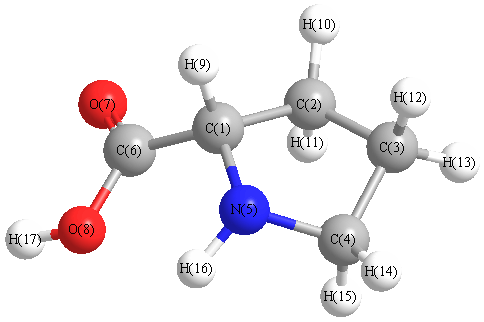
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3748 |
3572 |
64.05 |
|
|
|
| 2 |
A |
3577 |
3410 |
4.70 |
|
|
|
| 3 |
A |
3177 |
3028 |
17.56 |
|
|
|
| 4 |
A |
3154 |
3006 |
11.25 |
|
|
|
| 5 |
A |
3127 |
2980 |
25.28 |
|
|
|
| 6 |
A |
3122 |
2976 |
43.87 |
|
|
|
| 7 |
A |
3105 |
2959 |
15.79 |
|
|
|
| 8 |
A |
3065 |
2921 |
19.03 |
|
|
|
| 9 |
A |
2976 |
2836 |
72.65 |
|
|
|
| 10 |
A |
1878 |
1790 |
241.85 |
|
|
|
| 11 |
A |
1553 |
1480 |
0.91 |
|
|
|
| 12 |
A |
1535 |
1463 |
6.20 |
|
|
|
| 13 |
A |
1514 |
1443 |
0.84 |
|
|
|
| 14 |
A |
1473 |
1404 |
23.22 |
|
|
|
| 15 |
A |
1399 |
1334 |
30.86 |
|
|
|
| 16 |
A |
1389 |
1324 |
7.04 |
|
|
|
| 17 |
A |
1352 |
1289 |
11.28 |
|
|
|
| 18 |
A |
1344 |
1281 |
13.48 |
|
|
|
| 19 |
A |
1332 |
1270 |
2.18 |
|
|
|
| 20 |
A |
1275 |
1215 |
6.44 |
|
|
|
| 21 |
A |
1265 |
1206 |
2.12 |
|
|
|
| 22 |
A |
1236 |
1178 |
18.88 |
|
|
|
| 23 |
A |
1220 |
1163 |
14.17 |
|
|
|
| 24 |
A |
1195 |
1139 |
216.34 |
|
|
|
| 25 |
A |
1174 |
1119 |
25.15 |
|
|
|
| 26 |
A |
1123 |
1070 |
1.83 |
|
|
|
| 27 |
A |
1100 |
1048 |
4.85 |
|
|
|
| 28 |
A |
1021 |
973 |
5.08 |
|
|
|
| 29 |
A |
985 |
938 |
6.52 |
|
|
|
| 30 |
A |
951 |
907 |
4.78 |
|
|
|
| 31 |
A |
935 |
891 |
3.30 |
|
|
|
| 32 |
A |
908 |
866 |
65.72 |
|
|
|
| 33 |
A |
858 |
818 |
28.21 |
|
|
|
| 34 |
A |
796 |
758 |
2.66 |
|
|
|
| 35 |
A |
760 |
724 |
42.15 |
|
|
|
| 36 |
A |
691 |
659 |
111.94 |
|
|
|
| 37 |
A |
630 |
600 |
45.94 |
|
|
|
| 38 |
A |
598 |
570 |
15.94 |
|
|
|
| 39 |
A |
514 |
490 |
33.84 |
|
|
|
| 40 |
A |
500 |
477 |
15.76 |
|
|
|
| 41 |
A |
354 |
337 |
2.25 |
|
|
|
| 42 |
A |
262 |
249 |
3.09 |
|
|
|
| 43 |
A |
186 |
177 |
0.30 |
|
|
|
| 44 |
A |
58 |
56 |
0.29 |
|
|
|
| 45 |
A |
42 |
40 |
1.81 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32227.1 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 30715.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.120 |
|
|
|
| 2 |
C |
-0.353 |
|
|
|
| 3 |
C |
-0.335 |
|
|
|
| 4 |
C |
-0.198 |
|
|
|
| 5 |
N |
-0.571 |
|
|
|
| 6 |
C |
0.578 |
|
|
|
| 7 |
O |
-0.462 |
|
|
|
| 8 |
O |
-0.591 |
|
|
|
| 9 |
H |
0.210 |
|
|
|
| 10 |
H |
0.184 |
|
|
|
| 11 |
H |
0.207 |
|
|
|
| 12 |
H |
0.184 |
|
|
|
| 13 |
H |
0.175 |
|
|
|
| 14 |
H |
0.177 |
|
|
|
| 15 |
H |
0.150 |
|
|
|
| 16 |
H |
0.337 |
|
|
|
| 17 |
H |
0.428 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.041 |
-1.324 |
-0.457 |
1.401 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.310 |
5.885 |
-1.134 |
| y |
5.885 |
-48.806 |
2.688 |
| z |
-1.134 |
2.688 |
-46.830 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.507 |
5.885 |
-1.134 |
| y |
5.885 |
-2.735 |
2.688 |
| z |
-1.134 |
2.688 |
0.228 |
|
| Polar |
|---|
| 3z2-r2 | 0.456 |
|---|
| x2-y2 | 3.495 |
|---|
| xy | 5.885 |
|---|
| xz | -1.134 |
|---|
| yz | 2.688 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.745 |
-0.008 |
0.124 |
| y |
-0.008 |
9.215 |
0.162 |
| z |
0.124 |
0.162 |
7.659 |
<r2> (average value of r
2) Å
2
| <r2> |
256.274 |
| (<r2>)1/2 |
16.009 |