Jump to
S1C2
S1C3
Energy calculated at CCD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -691.750669 |
| Energy at 298.15K | -691.750101 |
| HF Energy | -691.441597 |
| Nuclear repulsion energy | 60.370726 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at CCD/6-31G**
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.324 |
| C2 |
0.000 |
0.000 |
-1.301 |
| N3 |
0.000 |
0.000 |
-2.478 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.6247 | 3.8022 |
C2 | 2.6247 | | 1.1774 | N3 | 3.8022 | 1.1774 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
180.000 |
|
K1 |
N3 |
C2 |
0.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at CCD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -691.750798 |
| Energy at 298.15K | |
| HF Energy | -691.447180 |
| Nuclear repulsion energy | 63.766517 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at CCD/6-31G**
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.225 |
| C2 |
0.000 |
0.000 |
-2.429 |
| N3 |
0.000 |
0.000 |
-1.244 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 3.6544 | 2.4699 |
C2 | 3.6544 | | 1.1845 | N3 | 2.4699 | 1.1845 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
0.000 |
|
K1 |
N3 |
C2 |
180.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at CCD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -691.758197 |
| Energy at 298.15K | -691.758118 |
| HF Energy | -691.450085 |
| Nuclear repulsion energy | 68.136916 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at CCD/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
1.049 |
0.000 |
| C2 |
0.636 |
-1.586 |
0.000 |
| N3 |
-0.545 |
-1.487 |
0.000 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.7109 | 2.5937 |
C2 | 2.7109 | | 1.1854 | N3 | 2.5937 | 1.1854 | |
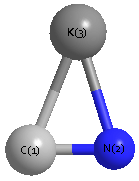 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
71.613 |
|
K1 |
N3 |
C2 |
82.683 |
| C2 |
K1 |
N3 |
25.704 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
