return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H6 (3-Penten-1-yne, (Z)-)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-193.450293
Energy at 298.15K-193.454640
HF Energy-192.755412
Nuclear repulsion energy141.246244
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3547 3314 58.80      
2 A' 3264 3050 24.42      
3 A' 3248 3034 4.28      
4 A' 3238 3025 1.18      
5 A' 3124 2919 17.17      
6 A' 2155 2013 1.34      
7 A' 1718 1605 1.84      
8 A' 1546 1444 4.61      
9 A' 1476 1380 3.35      
10 A' 1437 1342 6.08      
11 A' 1292 1207 0.13      
12 A' 1150 1075 0.62      
13 A' 996 930 15.53      
14 A' 930 869 1.46      
15 A' 663 620 3.50      
16 A' 599 560 42.06      
17 A' 373 349 3.69      
18 A' 160 150 0.12      
19 A" 3203 2993 16.11      
20 A" 1545 1444 5.53      
21 A" 1093 1021 1.13      
22 A" 978 914 0.10      
23 A" 753 703 19.21      
24 A" 544 508 57.72      
25 A" 439 410 3.86      
26 A" 261 244 0.06      
27 A" 141 132 0.45      

Unscaled Zero Point Vibrational Energy (zpe) 19935.5 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 18627.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.39609 0.11194 0.08869

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 3.137 -0.981 0.000
C2 2.239 -0.415 0.000
C3 1.194 0.219 0.000
H4 0.123 2.076 0.000
C5 0.000 1.000 0.000
H6 -2.075 1.168 0.000
C7 -1.240 0.477 0.000
H8 -0.656 -1.579 0.000
H9 -2.154 -1.245 0.877
H10 -2.154 -1.245 -0.877
C11 -1.563 -0.980 0.000

Atom - Atom Distances (Å)
  H1 C2 C3 H4 C5 H6 C7 H8 H9 H10 C11
H11.06152.28404.29343.71045.63804.61423.84025.36965.36964.6997
C21.06151.22273.26872.64904.59573.59243.12104.55604.55603.8437
C32.28401.22272.14401.42673.40382.44772.57953.75683.75683.0055
H44.29343.26872.14401.08262.37782.10083.73744.12014.12013.4900
C53.71042.64901.42671.08262.08191.34622.66183.23213.23212.5227
H65.63804.59573.40382.37782.08191.08353.09242.56812.56812.2087
C74.61423.59242.44772.10081.34621.08352.13812.13712.13711.4927
H83.84023.12102.57953.73742.66183.09242.13811.76741.76741.0865
H95.36964.55603.75684.12013.23212.56812.13711.76741.75321.0899
H105.36964.55603.75684.12013.23212.56812.13711.76741.75321.0899
C114.69973.84373.00553.49002.52272.20871.49271.08651.08991.0899

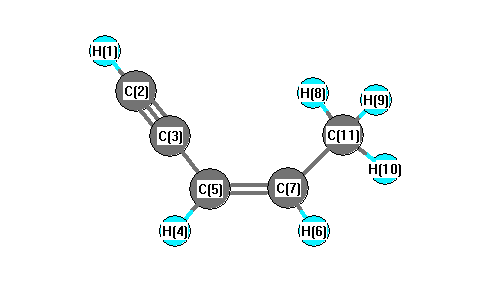
picture of 3-Penten-1-yne, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C3 178.957 C2 C3 C5 177.972
C3 C5 H4 116.726 C3 C5 C7 123.917
H4 C5 C7 119.357 C5 C7 H6 117.526
C5 C7 C11 125.318 H6 C7 C11 117.156
C7 C11 H8 111.008 C7 C11 H9 110.715
C7 C11 H10 110.715 H8 C11 H9 108.599
H8 C11 H10 108.599 H9 C11 H10 107.084
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability