return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (1,3-Pentadiene, (Z)-)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-194.681085
Energy at 298.15K-194.688471
HF Energy-193.968796
Nuclear repulsion energy155.791909
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3342 3122 13.70      
2 A' 3265 3051 22.78      
3 A' 3249 3036 22.57      
4 A' 3240 3028 4.97      
5 A' 3238 3025 8.30      
6 A' 3221 3010 0.56      
7 A' 3126 2921 21.84      
8 A' 1756 1641 3.19      
9 A' 1690 1579 0.78      
10 A' 1553 1451 5.54      
11 A' 1517 1417 7.87      
12 A' 1468 1372 2.96      
13 A' 1429 1335 1.49      
14 A' 1352 1263 1.19      
15 A' 1319 1233 0.69      
16 A' 1221 1141 1.14      
17 A' 1090 1019 0.06      
18 A' 993 928 7.76      
19 A' 919 859 1.21      
20 A' 622 581 4.31      
21 A' 392 366 2.98      
22 A' 227 212 0.25      
23 A" 3201 2991 17.96      
24 A" 1541 1440 5.06      
25 A" 1089 1017 1.28      
26 A" 1038 970 15.92      
27 A" 978 914 3.72      
28 A" 915 855 32.59      
29 A" 797 745 10.50      
30 A" 636 595 20.81      
31 A" 347 324 0.09      
32 A" 117 109 0.01      
33 A" 104 97 0.46      

Unscaled Zero Point Vibrational Energy (zpe) 25495.1 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 23822.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.53235 0.08834 0.07683

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.389 0.160 0.000
C2 1.099 -0.215 0.000
C3 0.000 0.739 0.000
C4 -1.315 0.449 0.000
C5 -1.945 -0.909 0.000
H6 2.670 1.204 0.000
H7 3.189 -0.564 0.000
H8 0.863 -1.271 0.000
H9 0.289 1.784 0.000
H10 -2.009 1.282 0.000
H11 -1.213 -1.711 0.000
H12 -2.581 -1.035 0.876
H13 -2.581 -1.035 -0.876

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.34242.45783.71494.46291.08141.07942.09112.65474.53854.05795.18525.1852
C21.34241.45522.50393.12212.11642.11911.08242.15703.44992.75383.87073.8707
C32.45781.45521.34652.54872.71053.44532.18731.08492.08112.73323.25153.2515
C43.71492.50391.34651.49674.05624.61692.77542.08741.08422.16212.13792.1379
C54.46293.12212.54871.49675.07545.14552.83113.49912.19181.08571.09001.0900
H61.08142.11642.71054.05625.07541.84243.06412.45094.68004.85505.77495.7749
H71.07942.11913.44534.61695.14551.84242.43113.73195.51654.54895.85505.8550
H82.09111.08242.18732.77542.83113.06412.43113.10883.84282.12203.56143.5614
H92.65472.15701.08492.08743.49912.45093.73193.10882.35253.80424.11724.1172
H104.53853.44992.08111.08422.19184.68005.51653.84282.35253.09692.54202.5420
H114.05792.75382.73322.16211.08574.85504.54892.12203.80423.09691.75951.7595
H125.18523.87073.25152.13791.09005.77495.85503.56144.11722.54201.75951.7524
H135.18523.87073.25152.13791.09005.77495.85503.56144.11722.54201.75951.7524

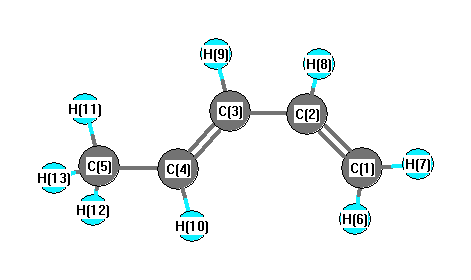
picture of 1,3-Pentadiene, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 122.885 C1 C2 H8 118.782
C2 C1 H6 121.287 C2 C1 H7 121.709
C2 C3 C4 126.639 C2 C3 H9 115.468
C3 C2 H8 118.334 C3 C4 C5 127.306
C3 C4 H10 117.367 C4 C3 H9 117.893
C4 C5 H11 112.733 C4 C5 H12 110.504
C4 C5 H13 110.504 C5 C4 H10 115.327
H6 C1 H7 117.004 H11 C5 H12 107.941
H11 C5 H13 107.941 H12 C5 H13 107.002
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability