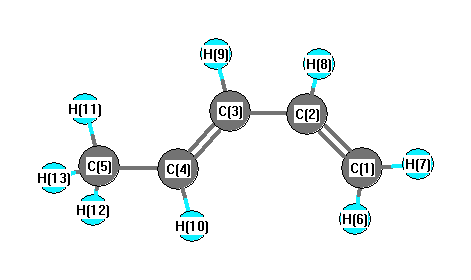
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3271 |
3122 |
13.67 |
|
|
|
| 2 |
A' |
3194 |
3048 |
41.80 |
|
|
|
| 3 |
A' |
3188 |
3042 |
5.55 |
|
|
|
| 4 |
A' |
3174 |
3029 |
8.20 |
|
|
|
| 5 |
A' |
3170 |
3025 |
5.23 |
|
|
|
| 6 |
A' |
3159 |
3015 |
2.94 |
|
|
|
| 7 |
A' |
3053 |
2914 |
26.55 |
|
|
|
| 8 |
A' |
1753 |
1673 |
10.19 |
|
|
|
| 9 |
A' |
1695 |
1617 |
2.04 |
|
|
|
| 10 |
A' |
1500 |
1431 |
6.50 |
|
|
|
| 11 |
A' |
1479 |
1411 |
13.74 |
|
|
|
| 12 |
A' |
1425 |
1360 |
3.04 |
|
|
|
| 13 |
A' |
1398 |
1334 |
1.91 |
|
|
|
| 14 |
A' |
1322 |
1262 |
1.02 |
|
|
|
| 15 |
A' |
1295 |
1235 |
1.48 |
|
|
|
| 16 |
A' |
1200 |
1145 |
1.62 |
|
|
|
| 17 |
A' |
1067 |
1018 |
0.00 |
|
|
|
| 18 |
A' |
977 |
932 |
13.73 |
|
|
|
| 19 |
A' |
909 |
867 |
2.16 |
|
|
|
| 20 |
A' |
618 |
590 |
5.94 |
|
|
|
| 21 |
A' |
388 |
370 |
4.33 |
|
|
|
| 22 |
A' |
217 |
207 |
0.29 |
|
|
|
| 23 |
A" |
3111 |
2969 |
17.62 |
|
|
|
| 24 |
A" |
1488 |
1420 |
7.04 |
|
|
|
| 25 |
A" |
1059 |
1011 |
0.69 |
|
|
|
| 26 |
A" |
1046 |
999 |
11.62 |
|
|
|
| 27 |
A" |
996 |
951 |
1.51 |
|
|
|
| 28 |
A" |
932 |
890 |
46.65 |
|
|
|
| 29 |
A" |
810 |
773 |
6.84 |
|
|
|
| 30 |
A" |
647 |
617 |
25.01 |
|
|
|
| 31 |
A" |
360 |
343 |
0.07 |
|
|
|
| 32 |
A" |
150 |
143 |
0.06 |
|
|
|
| 33 |
A" |
124 |
118 |
0.65 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25086.6 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 23940.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.309 |
|
|
|
| 2 |
C |
-0.063 |
|
|
|
| 3 |
C |
-0.104 |
|
|
|
| 4 |
C |
-0.116 |
|
|
|
| 5 |
C |
-0.426 |
|
|
|
| 6 |
H |
0.123 |
|
|
|
| 7 |
H |
0.126 |
|
|
|
| 8 |
H |
0.111 |
|
|
|
| 9 |
H |
0.111 |
|
|
|
| 10 |
H |
0.118 |
|
|
|
| 11 |
H |
0.135 |
|
|
|
| 12 |
H |
0.146 |
|
|
|
| 13 |
H |
0.146 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.502 |
-0.219 |
0.000 |
0.547 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.538 |
0.200 |
0.000 |
| y |
0.200 |
-29.141 |
0.000 |
| z |
0.000 |
0.000 |
-34.238 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.151 |
0.200 |
0.000 |
| y |
0.200 |
2.247 |
0.000 |
| z |
0.000 |
0.000 |
-5.398 |
|
| Polar |
|---|
| 3z2-r2 | -10.796 |
|---|
| x2-y2 | 0.603 |
|---|
| xy | 0.200 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.274 |
0.864 |
0.000 |
| y |
0.864 |
7.286 |
0.000 |
| z |
0.000 |
0.000 |
3.907 |
<r2> (average value of r
2) Å
2
| <r2> |
155.428 |
| (<r2>)1/2 |
12.467 |