return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10S (2H-Thiopyran, tetrahydro-)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-593.576113
Energy at 298.15K-593.588597
HF Energy-592.696446
Nuclear repulsion energy313.190332
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3202 2992 24.47      
2 A 3186 2977 36.95      
3 A 3173 2965 27.88      
4 A 3126 2921 19.37      
5 A 3120 2915 24.14      
6 A 3111 2907 19.44      
7 A 1552 1450 1.37      
8 A 1539 1438 10.01      
9 A 1522 1422 7.31      
10 A 1430 1336 0.82      
11 A 1393 1301 7.22      
12 A 1313 1227 0.69      
13 A 1288 1204 6.83      
14 A 1126 1053 2.46      
15 A 1068 998 0.61      
16 A 1012 946 8.21      
17 A 862 806 4.40      
18 A 855 799 0.29      
19 A 699 654 2.66      
20 A 514 480 0.27      
21 A 375 350 0.34      
22 A 354 331 0.12      
23 A 190 177 1.80      
24 A 3203 2993 6.24      
25 A 3181 2972 17.24      
26 A 3129 2924 20.74      
27 A 3120 2915 34.37      
28 A 1532 1432 2.15      
29 A 1516 1417 0.57      
30 A 1436 1341 0.18      
31 A 1413 1320 0.63      
32 A 1355 1266 18.25      
33 A 1330 1243 2.06      
34 A 1207 1128 0.01      
35 A 1143 1068 0.41      
36 A 1133 1059 0.25      
37 A 982 918 0.18      
38 A 943 882 3.01      
39 A 827 773 0.10      
40 A 729 681 0.15      
41 A 401 375 0.29      
42 A 248 231 0.43      

Unscaled Zero Point Vibrational Energy (zpe) 32417.3 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 30290.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.13455 0.10051 0.06424

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.000 1.506 0.000
C2 0.069 -1.717 0.000
C3 0.387 -0.936 1.272
C4 0.387 -0.936 -1.272
C5 -0.389 0.371 1.358
C6 -0.389 0.371 -1.358
H7 0.623 -2.657 0.000
H8 -0.993 -1.981 0.000
H9 -1.463 0.172 1.365
H10 -1.463 0.172 -1.365
H11 0.148 -1.546 2.146
H12 0.148 -1.546 -2.146
H13 1.455 -0.718 1.310
H14 1.455 -0.718 -1.310
H15 -0.148 0.905 -2.276
H16 -0.148 0.905 2.276

Atom - Atom Distances (Å)
  S1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
S13.22402.78002.78001.81191.81194.20883.62572.40462.40463.73353.73352.96272.96272.35832.3583
C23.22401.52621.52622.53282.53281.09081.09382.78952.78952.15382.15382.15342.15343.47933.4793
C32.78001.52622.54341.52223.03722.15282.14792.15813.40631.09223.47951.09132.80224.03252.1643
C42.78001.52622.54343.03721.52222.15282.14793.40632.15813.47951.09222.80221.09132.16434.0325
C51.81192.53281.52223.03722.71573.46922.78241.09182.93402.14124.02962.14253.42093.68061.0891
C61.81192.53283.03721.52222.71573.46922.78242.93401.09184.02962.14123.42092.14251.08913.6806
H74.20881.09082.15282.15283.46923.46921.75173.77093.77092.46212.46212.48312.48314.29684.2968
H83.62571.09382.14792.14792.78242.78241.75172.59302.59302.46882.46883.05043.05043.77173.7717
H92.40462.78952.15813.40631.09182.93403.77092.59302.73082.48154.22803.05144.05753.94001.7590
H102.40462.78953.40632.15812.93401.09183.77092.59302.73084.22802.48154.05753.05141.75903.9400
H113.73352.15381.09223.47952.14124.02962.46212.46882.48154.22804.29101.75873.78575.06412.4728
H123.73352.15383.47951.09224.02962.14122.46212.46884.22802.48154.29103.78571.75872.47285.0641
H132.96272.15341.09132.80222.14253.42092.48313.05043.05144.05751.75873.78572.61914.24982.4779
H142.96272.15342.80221.09133.42092.14252.48313.05044.05753.05143.78571.75872.61912.47794.2498
H152.35833.47934.03252.16433.68061.08914.29683.77173.94001.75905.06412.47284.24982.47794.5515
H162.35833.47932.16434.03251.08913.68064.29683.77171.75903.94002.47285.06412.47794.24984.5515

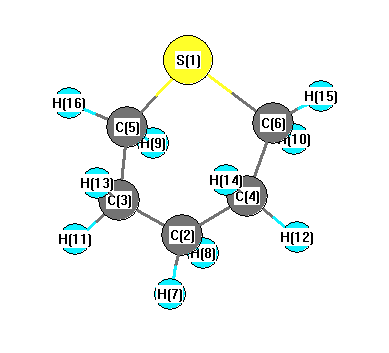
picture of 2H-Thiopyran, tetrahydro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C5 C3 112.691 S1 C5 H9 109.288
S1 C5 H16 106.068 S1 C6 C4 112.691
S1 C6 H10 109.288 S1 C6 H15 106.068
C2 C3 C5 112.371 C2 C3 H11 109.571
C2 C3 H13 109.589 C2 C4 C6 112.371
C2 C4 H12 109.571 C2 C4 H14 109.589
C3 C2 C4 112.863 C3 C2 H7 109.571
C3 C2 H8 109.014 C3 C5 H9 110.208
C3 C5 H16 110.863 C4 C2 H7 109.571
C4 C2 H8 109.014 C4 C6 H10 110.208
C4 C6 H15 110.863 C5 S1 C6 97.078
C5 C3 H11 108.859 C5 C3 H13 109.008
C6 C4 H12 108.859 C6 C4 H14 109.008
H7 C2 H8 106.612 H9 C5 H16 107.517
H10 C6 H15 107.517 H11 C3 H13 107.310
H12 C4 H14 107.310
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability