Jump to
S1C2
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -2610.561891 |
| Energy at 298.15K | -2610.565548 |
| HF Energy | -2610.561891 |
| Nuclear repulsion energy | 80.866717 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3227 |
3075 |
4.71 |
|
|
|
| 2 |
A' |
1410 |
1344 |
20.35 |
|
|
|
| 3 |
A' |
723 |
689 |
19.20 |
|
|
|
| 4 |
A' |
162 |
155 |
80.44 |
|
|
|
| 5 |
A" |
3380 |
3221 |
0.10 |
|
|
|
| 6 |
A" |
938 |
894 |
1.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4919.5 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 4688.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
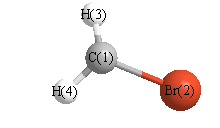
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.004 |
1.481 |
0.000 |
| Br2 |
-0.004 |
-0.367 |
0.000 |
| H3 |
0.074 |
1.983 |
0.953 |
| H4 |
0.074 |
1.983 |
-0.953 |
Atom - Atom Distances (Å)
| |
C1 |
Br2 |
H3 |
H4 |
| C1 | | 1.8486 | 1.0803 | 1.0803 |
Br2 | 1.8486 | | 2.5379 | 2.5379 | H3 | 1.0803 | 2.5379 | | 1.9067 | H4 | 1.0803 | 2.5379 | 1.9067 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
117.697 |
|
Br2 |
C1 |
H4 |
117.697 |
| H3 |
C1 |
H4 |
123.882 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.380 |
|
|
|
| 2 |
Br |
-0.054 |
|
|
|
| 3 |
H |
0.217 |
|
|
|
| 4 |
H |
0.217 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.145 |
1.058 |
0.000 |
1.068 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.953 |
0.312 |
0.000 |
| y |
0.312 |
-21.251 |
0.000 |
| z |
0.000 |
0.000 |
-23.989 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.333 |
0.312 |
0.000 |
| y |
0.312 |
3.720 |
0.000 |
| z |
0.000 |
0.000 |
-0.386 |
|
| Polar |
|---|
| 3z2-r2 | -0.773 |
|---|
| x2-y2 | -4.702 |
|---|
| xy | 0.312 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.381 |
0.036 |
0.000 |
| y |
0.036 |
5.376 |
0.000 |
| z |
0.000 |
0.000 |
3.150 |
<r2> (average value of r
2) Å
2
| <r2> |
42.406 |
| (<r2>)1/2 |
6.512 |
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -2610.561882 |
| Energy at 298.15K | |
| HF Energy | -2610.561882 |
| Nuclear repulsion energy | 80.881984 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3228 |
3077 |
4.25 |
107.08 |
0.12 |
0.21 |
| 2 |
A1 |
1409 |
1343 |
20.55 |
2.65 |
0.65 |
0.79 |
| 3 |
A1 |
718 |
684 |
18.85 |
7.88 |
0.22 |
0.36 |
| 4 |
B1 |
116i |
110i |
83.98 |
0.42 |
0.75 |
0.86 |
| 5 |
B2 |
3382 |
3224 |
0.16 |
57.73 |
0.75 |
0.86 |
| 6 |
B2 |
935 |
892 |
1.76 |
4.11 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 4778.2 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 4554.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.481 |
| Br2 |
0.000 |
0.000 |
0.367 |
| H3 |
0.000 |
0.954 |
-1.986 |
| H4 |
0.000 |
-0.954 |
-1.986 |
Atom - Atom Distances (Å)
| |
C1 |
Br2 |
H3 |
H4 |
| C1 | | 1.8478 | 1.0800 | 1.0800 |
Br2 | 1.8478 | | 2.5395 | 2.5395 | H3 | 1.0800 | 2.5395 | | 1.9090 | H4 | 1.0800 | 2.5395 | 1.9090 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
117.903 |
|
Br2 |
C1 |
H4 |
117.903 |
| H3 |
C1 |
H4 |
124.194 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.381 |
|
|
|
| 2 |
Br |
-0.053 |
|
|
|
| 3 |
H |
0.217 |
|
|
|
| 4 |
H |
0.217 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.061 |
1.061 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.969 |
0.000 |
0.000 |
| y |
0.000 |
-23.976 |
0.000 |
| z |
0.000 |
0.000 |
-21.225 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.369 |
0.000 |
0.000 |
| y |
0.000 |
-0.379 |
0.000 |
| z |
0.000 |
0.000 |
3.747 |
|
| Polar |
|---|
| 3z2-r2 | 7.495 |
|---|
| x2-y2 | -1.993 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.375 |
0.000 |
0.000 |
| y |
0.000 |
3.146 |
0.000 |
| z |
0.000 |
0.000 |
5.372 |
<r2> (average value of r
2) Å
2
| <r2> |
42.401 |
| (<r2>)1/2 |
6.512 |
