Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -2610.544030 |
| Energy at 298.15K | -2610.547534 |
| HF Energy | -2610.544030 |
| Nuclear repulsion energy | 80.857998 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3218 |
3071 |
4.26 |
|
|
|
| 2 |
A' |
1401 |
1337 |
20.13 |
|
|
|
| 3 |
A' |
720 |
687 |
18.99 |
|
|
|
| 4 |
A' |
74 |
70 |
77.64 |
|
|
|
| 5 |
A" |
3379 |
3224 |
0.23 |
|
|
|
| 6 |
A" |
938 |
895 |
1.66 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4864.6 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 4642.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.002 |
1.482 |
0.000 |
| Br2 |
-0.002 |
-0.367 |
0.000 |
| H3 |
0.039 |
1.984 |
0.955 |
| H4 |
0.039 |
1.984 |
-0.955 |
Atom - Atom Distances (Å)
| |
C1 |
Br2 |
H3 |
H4 |
| C1 | | 1.8490 | 1.0793 | 1.0793 |
Br2 | 1.8490 | | 2.5376 | 2.5376 | H3 | 1.0793 | 2.5376 | | 1.9093 | H4 | 1.0793 | 2.5376 | 1.9093 | |
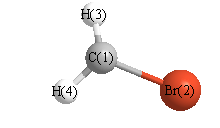 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
117.707 |
|
Br2 |
C1 |
H4 |
117.707 |
| H3 |
C1 |
H4 |
124.390 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.289 |
|
|
|
| 2 |
Br |
-0.061 |
|
|
|
| 3 |
H |
0.175 |
|
|
|
| 4 |
H |
0.175 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.074 |
1.054 |
0.000 |
1.057 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.938 |
0.158 |
0.000 |
| y |
0.158 |
-21.265 |
0.000 |
| z |
0.000 |
0.000 |
-23.996 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.307 |
0.158 |
0.000 |
| y |
0.158 |
3.702 |
0.000 |
| z |
0.000 |
0.000 |
-0.394 |
|
| Polar |
|---|
| 3z2-r2 | -0.788 |
|---|
| x2-y2 | -4.673 |
|---|
| xy | 0.158 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.405 |
0.019 |
0.000 |
| y |
0.019 |
5.411 |
0.000 |
| z |
0.000 |
0.000 |
3.179 |
<r2> (average value of r
2) Å
2
| <r2> |
42.413 |
| (<r2>)1/2 |
6.513 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -2610.544024 |
| Energy at 298.15K | |
| HF Energy | -2610.544024 |
| Nuclear repulsion energy | 80.864767 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3217 |
3070 |
4.13 |
108.97 |
0.12 |
0.21 |
| 2 |
A1 |
1401 |
1337 |
20.14 |
2.57 |
0.65 |
0.79 |
| 3 |
A1 |
715 |
683 |
18.90 |
8.07 |
0.22 |
0.36 |
| 4 |
B1 |
62i |
59i |
78.65 |
0.47 |
0.75 |
0.86 |
| 5 |
B2 |
3378 |
3224 |
0.25 |
57.56 |
0.75 |
0.86 |
| 6 |
B2 |
937 |
894 |
1.67 |
4.07 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 4793.3 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 4574.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.481 |
| Br2 |
0.000 |
0.000 |
0.367 |
| H3 |
0.000 |
0.955 |
-1.984 |
| H4 |
0.000 |
-0.955 |
-1.984 |
Atom - Atom Distances (Å)
| |
C1 |
Br2 |
H3 |
H4 |
| C1 | | 1.8488 | 1.0792 | 1.0792 |
Br2 | 1.8488 | | 2.5381 | 2.5381 | H3 | 1.0792 | 2.5381 | | 1.9098 | H4 | 1.0792 | 2.5381 | 1.9098 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
117.770 |
|
Br2 |
C1 |
H4 |
117.770 |
| H3 |
C1 |
H4 |
124.459 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.289 |
|
|
|
| 2 |
Br |
-0.061 |
|
|
|
| 3 |
H |
0.175 |
|
|
|
| 4 |
H |
0.175 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.055 |
1.055 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.942 |
0.000 |
0.000 |
| y |
0.000 |
-23.993 |
0.000 |
| z |
0.000 |
0.000 |
-21.258 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.316 |
0.000 |
0.000 |
| y |
0.000 |
-0.393 |
0.000 |
| z |
0.000 |
0.000 |
3.709 |
|
| Polar |
|---|
| 3z2-r2 | 7.418 |
|---|
| x2-y2 | -1.949 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.402 |
0.000 |
0.000 |
| y |
0.000 |
3.175 |
0.000 |
| z |
0.000 |
0.000 |
5.410 |
<r2> (average value of r
2) Å
2
| <r2> |
42.411 |
| (<r2>)1/2 |
6.512 |
