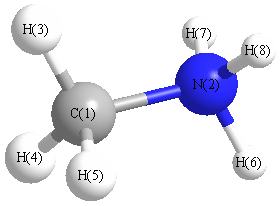
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3096 |
2951 |
53.85 |
|
|
|
| 2 |
A1 |
2511 |
2394 |
86.43 |
|
|
|
| 3 |
A1 |
1337 |
1274 |
130.53 |
|
|
|
| 4 |
A1 |
1140 |
1086 |
9.69 |
|
|
|
| 5 |
A1 |
871 |
830 |
28.28 |
|
|
|
| 6 |
A2 |
281 |
268 |
0.00 |
|
|
|
| 7 |
E |
3196 |
3046 |
14.00 |
|
|
|
| 7 |
E |
3196 |
3046 |
14.01 |
|
|
|
| 8 |
E |
1564 |
1491 |
1819.75 |
|
|
|
| 8 |
E |
1564 |
1491 |
1819.87 |
|
|
|
| 9 |
E |
1485 |
1415 |
351.11 |
|
|
|
| 9 |
E |
1485 |
1415 |
351.26 |
|
|
|
| 10 |
E |
1225 |
1168 |
4.18 |
|
|
|
| 10 |
E |
1225 |
1168 |
4.17 |
|
|
|
| 11 |
E |
861 |
820 |
6.07 |
|
|
|
| 11 |
E |
861 |
820 |
6.07 |
|
|
|
| 12 |
E |
512i |
488i |
4197.54 |
|
|
|
| 12 |
E |
512i |
488i |
4198.35 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12436.9 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 11853.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.419 |
|
|
|
| 2 |
N |
-0.422 |
|
|
|
| 3 |
H |
0.184 |
|
|
|
| 4 |
H |
0.184 |
|
|
|
| 5 |
H |
0.184 |
|
|
|
| 6 |
H |
0.096 |
|
|
|
| 7 |
H |
0.096 |
|
|
|
| 8 |
H |
0.096 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.013 |
1.013 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.845 |
0.000 |
0.000 |
| y |
0.000 |
-16.845 |
0.000 |
| z |
0.000 |
0.000 |
-16.591 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.127 |
0.000 |
0.000 |
| y |
0.000 |
-0.127 |
0.000 |
| z |
0.000 |
0.000 |
0.254 |
|
| Polar |
|---|
| 3z2-r2 | 0.508 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.530 |
0.000 |
0.000 |
| y |
0.000 |
8.531 |
0.000 |
| z |
0.000 |
0.000 |
5.202 |
<r2> (average value of r
2) Å
2
| <r2> |
31.373 |
| (<r2>)1/2 |
5.601 |