Jump to
S2C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -2609.863666 |
| Energy at 298.15K | -2609.866390 |
| HF Energy | -2609.863666 |
| Nuclear repulsion energy | 70.338646 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.026 |
1.550 |
0.000 |
| Br2 |
0.026 |
-0.316 |
0.000 |
| H3 |
-1.073 |
1.746 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
Br2 |
H3 |
| C1 | | 1.8655 | 1.1166 |
Br2 | 1.8655 | | 2.3365 | H3 | 1.1166 | 2.3365 | |
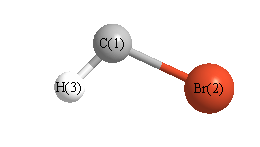 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
100.122 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.120 |
|
|
|
| 2 |
Br |
-0.025 |
|
|
|
| 3 |
H |
0.145 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.373 |
-0.660 |
0.000 |
1.523 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.019 |
-2.971 |
0.000 |
| y |
-2.971 |
-25.292 |
0.000 |
| z |
0.000 |
0.000 |
-23.445 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.650 |
-2.971 |
0.000 |
| y |
-2.971 |
-1.060 |
0.000 |
| z |
0.000 |
0.000 |
1.711 |
|
| Polar |
|---|
| 3z2-r2 | 3.421 |
|---|
| x2-y2 | 0.273 |
|---|
| xy | -2.971 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.177 |
-0.354 |
0.000 |
| y |
-0.354 |
5.924 |
0.000 |
| z |
0.000 |
0.000 |
2.620 |
<r2> (average value of r
2) Å
2
| <r2> |
37.484 |
| (<r2>)1/2 |
6.122 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -2609.867055 |
| Energy at 298.15K | -2609.869767 |
| HF Energy | -2609.867055 |
| Nuclear repulsion energy | 71.261886 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.021 |
1.497 |
0.000 |
| Br2 |
0.021 |
-0.318 |
0.000 |
| H3 |
-0.852 |
2.142 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
Br2 |
H3 |
| C1 | | 1.8146 | 1.0854 |
Br2 | 1.8146 | | 2.6098 | H3 | 1.0854 | 2.6098 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.152 |
|
|
|
| 2 |
Br |
-0.022 |
|
|
|
| 3 |
H |
0.174 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.735 |
0.254 |
0.000 |
0.777 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.935 |
-1.692 |
0.000 |
| y |
-1.692 |
-21.595 |
0.000 |
| z |
0.000 |
0.000 |
-25.060 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.608 |
-1.692 |
0.000 |
| y |
-1.692 |
2.903 |
0.000 |
| z |
0.000 |
0.000 |
-2.295 |
|
| Polar |
|---|
| 3z2-r2 | -4.590 |
|---|
| x2-y2 | -2.341 |
|---|
| xy | -1.692 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.726 |
-0.243 |
0.000 |
| y |
-0.243 |
5.471 |
0.000 |
| z |
0.000 |
0.000 |
2.350 |
<r2> (average value of r
2) Å
2
| <r2> |
37.005 |
| (<r2>)1/2 |
6.083 |
