Jump to
S1C2
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -246.568353 |
| Energy at 298.15K | -246.573924 |
| HF Energy | -245.794537 |
| Nuclear repulsion energy | 156.735117 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3262 |
3048 |
16.33 |
|
|
|
| 2 |
A' |
3090 |
2888 |
46.40 |
|
|
|
| 3 |
A' |
3073 |
2871 |
22.09 |
|
|
|
| 4 |
A' |
2233 |
2086 |
9.85 |
|
|
|
| 5 |
A' |
1577 |
1474 |
2.38 |
|
|
|
| 6 |
A' |
1560 |
1458 |
5.72 |
|
|
|
| 7 |
A' |
1526 |
1426 |
1.70 |
|
|
|
| 8 |
A' |
1448 |
1353 |
45.73 |
|
|
|
| 9 |
A' |
1254 |
1171 |
51.52 |
|
|
|
| 10 |
A' |
1189 |
1111 |
112.65 |
|
|
|
| 11 |
A' |
1016 |
949 |
30.16 |
|
|
|
| 12 |
A' |
973 |
910 |
7.76 |
|
|
|
| 13 |
A' |
544 |
508 |
1.15 |
|
|
|
| 14 |
A' |
383 |
358 |
2.21 |
|
|
|
| 15 |
A' |
179 |
167 |
2.26 |
|
|
|
| 16 |
A" |
3168 |
2960 |
45.45 |
|
|
|
| 17 |
A" |
3120 |
2916 |
30.08 |
|
|
|
| 18 |
A" |
1552 |
1450 |
6.36 |
|
|
|
| 19 |
A" |
1289 |
1204 |
3.22 |
|
|
|
| 20 |
A" |
1210 |
1131 |
4.03 |
|
|
|
| 21 |
A" |
1063 |
994 |
2.25 |
|
|
|
| 22 |
A" |
362 |
338 |
1.87 |
|
|
|
| 23 |
A" |
233 |
217 |
4.43 |
|
|
|
| 24 |
A" |
88 |
82 |
0.10 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17694.0 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 16533.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.132 |
-0.336 |
0.000 |
| O2 |
-0.739 |
-0.605 |
0.000 |
| C3 |
0.000 |
0.596 |
0.000 |
| C4 |
1.427 |
0.259 |
0.000 |
| N5 |
2.581 |
0.022 |
0.000 |
| H6 |
-2.630 |
-1.299 |
0.000 |
| H7 |
-2.429 |
0.227 |
0.889 |
| H8 |
-2.429 |
0.227 |
-0.889 |
| H9 |
-0.220 |
1.205 |
-0.884 |
| H10 |
-0.220 |
1.205 |
0.884 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4196 | 2.3273 | 3.6086 | 4.7273 | 1.0842 | 1.0929 | 1.0929 | 2.6108 | 2.6108 |
O2 | 1.4196 | | 1.4100 | 2.3313 | 3.3785 | 2.0149 | 2.0828 | 2.0828 | 2.0804 | 2.0804 | C3 | 2.3273 | 1.4100 | | 1.4662 | 2.6445 | 3.2419 | 2.6125 | 2.6125 | 1.0960 | 1.0960 | C4 | 3.6086 | 2.3313 | 1.4662 | | 1.1786 | 4.3458 | 3.9568 | 3.9568 | 2.0949 | 2.0949 | N5 | 4.7273 | 3.3785 | 2.6445 | 1.1786 | | 5.3762 | 5.0924 | 5.0924 | 3.1667 | 3.1667 | H6 | 1.0842 | 2.0149 | 3.2419 | 4.3458 | 5.3762 | | 1.7773 | 1.7773 | 3.5867 | 3.5867 | H7 | 1.0929 | 2.0828 | 2.6125 | 3.9568 | 5.0924 | 1.7773 | | 1.7779 | 2.9970 | 2.4161 | H8 | 1.0929 | 2.0828 | 2.6125 | 3.9568 | 5.0924 | 1.7773 | 1.7779 | | 2.4161 | 2.9970 | H9 | 2.6108 | 2.0804 | 1.0960 | 2.0949 | 3.1667 | 3.5867 | 2.9970 | 2.4161 | | 1.7687 | H10 | 2.6108 | 2.0804 | 1.0960 | 2.0949 | 3.1667 | 3.5867 | 2.4161 | 2.9970 | 1.7687 | |
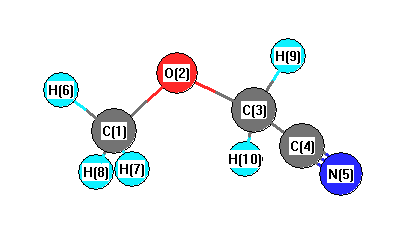 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
110.666 |
|
O2 |
C1 |
H6 |
106.403 |
| O2 |
C1 |
H7 |
111.324 |
|
O2 |
C1 |
H8 |
111.324 |
| O2 |
C3 |
C4 |
108.285 |
|
O2 |
C3 |
H9 |
111.617 |
| O2 |
C3 |
H10 |
111.617 |
|
C3 |
C4 |
N5 |
178.300 |
| C4 |
C3 |
H9 |
108.838 |
|
C4 |
C3 |
H10 |
108.838 |
| H6 |
C1 |
H7 |
109.446 |
|
H6 |
C1 |
H8 |
109.446 |
| H7 |
C1 |
H8 |
108.852 |
|
H9 |
C3 |
H10 |
107.580 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -246.571079 |
| Energy at 298.15K | -246.576771 |
| HF Energy | -245.796397 |
| Nuclear repulsion energy | 159.763081 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3263 |
3049 |
16.72 |
|
|
|
| 2 |
A |
3213 |
3002 |
5.51 |
|
|
|
| 3 |
A |
3184 |
2975 |
34.95 |
|
|
|
| 4 |
A |
3101 |
2897 |
32.78 |
|
|
|
| 5 |
A |
3098 |
2895 |
29.51 |
|
|
|
| 6 |
A |
2217 |
2071 |
9.70 |
|
|
|
| 7 |
A |
1572 |
1469 |
8.10 |
|
|
|
| 8 |
A |
1555 |
1453 |
5.84 |
|
|
|
| 9 |
A |
1539 |
1438 |
2.60 |
|
|
|
| 10 |
A |
1526 |
1426 |
0.09 |
|
|
|
| 11 |
A |
1423 |
1329 |
19.25 |
|
|
|
| 12 |
A |
1338 |
1251 |
5.74 |
|
|
|
| 13 |
A |
1245 |
1163 |
40.97 |
|
|
|
| 14 |
A |
1214 |
1134 |
10.43 |
|
|
|
| 15 |
A |
1186 |
1108 |
95.12 |
|
|
|
| 16 |
A |
1057 |
988 |
6.51 |
|
|
|
| 17 |
A |
968 |
904 |
31.02 |
|
|
|
| 18 |
A |
924 |
863 |
17.29 |
|
|
|
| 19 |
A |
599 |
560 |
2.41 |
|
|
|
| 20 |
A |
401 |
374 |
2.83 |
|
|
|
| 21 |
A |
351 |
328 |
1.17 |
|
|
|
| 22 |
A |
262 |
245 |
10.05 |
|
|
|
| 23 |
A |
180 |
168 |
4.62 |
|
|
|
| 24 |
A |
120 |
112 |
8.56 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17767.2 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 16601.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.537 |
-0.788 |
0.141 |
| O2 |
1.087 |
0.430 |
-0.444 |
| C3 |
-0.049 |
0.942 |
0.212 |
| C4 |
-1.231 |
0.073 |
0.040 |
| N5 |
-2.158 |
-0.642 |
-0.100 |
| H6 |
2.439 |
-1.065 |
-0.393 |
| H7 |
0.793 |
-1.579 |
0.035 |
| H8 |
1.767 |
-0.651 |
1.200 |
| H9 |
-0.256 |
1.915 |
-0.228 |
| H10 |
0.128 |
1.074 |
1.284 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4235 | 2.3476 | 2.9004 | 3.7055 | 1.0842 | 1.0910 | 1.0926 | 3.2638 | 2.5996 |
O2 | 1.4235 | | 1.4081 | 2.3947 | 3.4346 | 2.0160 | 2.0862 | 2.0816 | 2.0138 | 2.0789 | C3 | 2.3476 | 1.4081 | | 1.4771 | 2.6560 | 3.2531 | 2.6641 | 2.6099 | 1.0874 | 1.0950 | C4 | 2.9004 | 2.3947 | 1.4771 | | 1.1791 | 3.8666 | 2.6133 | 3.2952 | 2.1012 | 2.0969 | N5 | 3.7055 | 3.4346 | 2.6560 | 1.1791 | | 4.6254 | 3.0997 | 4.1348 | 3.1895 | 3.1761 | H6 | 1.0842 | 2.0160 | 3.2531 | 3.8666 | 4.6254 | | 1.7762 | 1.7779 | 4.0209 | 3.5677 | H7 | 1.0910 | 2.0862 | 2.6641 | 2.6133 | 3.0997 | 1.7762 | | 1.7792 | 3.6578 | 3.0070 | H8 | 1.0926 | 2.0816 | 2.6099 | 3.2952 | 4.1348 | 1.7779 | 1.7792 | | 3.5656 | 2.3808 | H9 | 3.2638 | 2.0138 | 1.0874 | 2.1012 | 3.1895 | 4.0209 | 3.6578 | 3.5656 | | 1.7723 | H10 | 2.5996 | 2.0789 | 1.0950 | 2.0969 | 3.1761 | 3.5677 | 3.0070 | 2.3808 | 1.7723 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
112.005 |
|
O2 |
C1 |
H6 |
106.227 |
| O2 |
C1 |
H7 |
111.449 |
|
O2 |
C1 |
H8 |
110.964 |
| O2 |
C3 |
C4 |
112.177 |
|
O2 |
C3 |
H9 |
106.892 |
| O2 |
C3 |
H10 |
111.692 |
|
C3 |
C4 |
N5 |
178.662 |
| C4 |
C3 |
H9 |
109.091 |
|
C4 |
C3 |
H10 |
108.313 |
| H6 |
C1 |
H7 |
109.493 |
|
H6 |
C1 |
H8 |
109.522 |
| H7 |
C1 |
H8 |
109.133 |
|
H9 |
C3 |
H10 |
108.591 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
