Jump to
S1C2
Energy calculated at QCISD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -246.581343 |
| Energy at 298.15K | -246.586900 |
| HF Energy | -245.796786 |
| Nuclear repulsion energy | 156.643399 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3227 |
3038 |
21.17 |
|
|
|
| 2 |
A' |
3070 |
2890 |
52.26 |
|
|
|
| 3 |
A' |
3061 |
2882 |
20.51 |
|
|
|
| 4 |
A' |
2357 |
2219 |
2.01 |
|
|
|
| 5 |
A' |
1576 |
1484 |
0.90 |
|
|
|
| 6 |
A' |
1560 |
1468 |
6.96 |
|
|
|
| 7 |
A' |
1526 |
1437 |
1.57 |
|
|
|
| 8 |
A' |
1454 |
1369 |
54.14 |
|
|
|
| 9 |
A' |
1260 |
1186 |
62.30 |
|
|
|
| 10 |
A' |
1192 |
1122 |
108.02 |
|
|
|
| 11 |
A' |
1021 |
961 |
30.95 |
|
|
|
| 12 |
A' |
969 |
912 |
9.01 |
|
|
|
| 13 |
A' |
542 |
510 |
1.10 |
|
|
|
| 14 |
A' |
383 |
361 |
2.24 |
|
|
|
| 15 |
A' |
179 |
168 |
2.45 |
|
|
|
| 16 |
A" |
3132 |
2949 |
51.24 |
|
|
|
| 17 |
A" |
3101 |
2919 |
34.53 |
|
|
|
| 18 |
A" |
1547 |
1456 |
5.42 |
|
|
|
| 19 |
A" |
1289 |
1214 |
3.44 |
|
|
|
| 20 |
A" |
1209 |
1138 |
4.85 |
|
|
|
| 21 |
A" |
1066 |
1003 |
2.73 |
|
|
|
| 22 |
A" |
360 |
339 |
2.31 |
|
|
|
| 23 |
A" |
226 |
213 |
4.55 |
|
|
|
| 24 |
A" |
85 |
80 |
0.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17695.5 cm
-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 16658.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.137 |
-0.341 |
0.000 |
| O2 |
-0.740 |
-0.602 |
0.000 |
| C3 |
0.000 |
0.598 |
0.000 |
| C4 |
1.437 |
0.257 |
0.000 |
| N5 |
2.580 |
0.025 |
0.000 |
| H6 |
-2.631 |
-1.308 |
0.000 |
| H7 |
-2.437 |
0.221 |
0.891 |
| H8 |
-2.437 |
0.221 |
-0.891 |
| H9 |
-0.215 |
1.208 |
-0.886 |
| H10 |
-0.215 |
1.208 |
0.886 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4210 | 2.3344 | 3.6235 | 4.7320 | 1.0865 | 1.0951 | 1.0951 | 2.6229 | 2.6229 |
O2 | 1.4210 | | 1.4095 | 2.3402 | 3.3796 | 2.0188 | 2.0853 | 2.0853 | 2.0823 | 2.0823 | C3 | 2.3344 | 1.4095 | | 1.4764 | 2.6432 | 3.2494 | 2.6218 | 2.6218 | 1.0971 | 1.0971 | C4 | 3.6235 | 2.3402 | 1.4764 | | 1.1671 | 4.3589 | 3.9747 | 3.9747 | 2.1017 | 2.1017 | N5 | 4.7320 | 3.3796 | 2.6432 | 1.1671 | | 5.3799 | 5.0995 | 5.0995 | 3.1620 | 3.1620 | H6 | 1.0865 | 2.0188 | 3.2494 | 4.3589 | 5.3799 | | 1.7808 | 1.7808 | 3.5996 | 3.5996 | H7 | 1.0951 | 2.0853 | 2.6218 | 3.9747 | 5.0995 | 1.7808 | | 1.7818 | 3.0113 | 2.4312 | H8 | 1.0951 | 2.0853 | 2.6218 | 3.9747 | 5.0995 | 1.7808 | 1.7818 | | 2.4312 | 3.0113 | H9 | 2.6229 | 2.0823 | 1.0971 | 2.1017 | 3.1620 | 3.5996 | 3.0113 | 2.4312 | | 1.7719 | H10 | 2.6229 | 2.0823 | 1.0971 | 2.1017 | 3.1620 | 3.5996 | 2.4312 | 3.0113 | 1.7719 | |
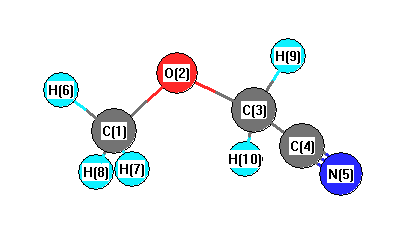 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
111.119 |
|
O2 |
C1 |
H6 |
106.485 |
| O2 |
C1 |
H7 |
111.286 |
|
O2 |
C1 |
H8 |
111.286 |
| O2 |
C3 |
C4 |
108.344 |
|
O2 |
C3 |
H9 |
111.740 |
| O2 |
C3 |
H10 |
111.740 |
|
C3 |
C4 |
N5 |
178.103 |
| C4 |
C3 |
H9 |
108.608 |
|
C4 |
C3 |
H10 |
108.608 |
| H6 |
C1 |
H7 |
109.429 |
|
H6 |
C1 |
H8 |
109.429 |
| H7 |
C1 |
H8 |
108.882 |
|
H9 |
C3 |
H10 |
107.716 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -246.583793 |
| Energy at 298.15K | -246.589475 |
| HF Energy | -245.798755 |
| Nuclear repulsion energy | 159.532898 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3228 |
3039 |
21.90 |
|
|
|
| 2 |
A |
3191 |
3004 |
8.04 |
|
|
|
| 3 |
A |
3151 |
2967 |
40.09 |
|
|
|
| 4 |
A |
3087 |
2906 |
32.76 |
|
|
|
| 5 |
A |
3078 |
2897 |
34.55 |
|
|
|
| 6 |
A |
2343 |
2206 |
2.57 |
|
|
|
| 7 |
A |
1568 |
1476 |
7.65 |
|
|
|
| 8 |
A |
1550 |
1460 |
4.96 |
|
|
|
| 9 |
A |
1542 |
1451 |
1.80 |
|
|
|
| 10 |
A |
1526 |
1437 |
0.38 |
|
|
|
| 11 |
A |
1429 |
1345 |
24.25 |
|
|
|
| 12 |
A |
1342 |
1263 |
6.78 |
|
|
|
| 13 |
A |
1251 |
1178 |
56.77 |
|
|
|
| 14 |
A |
1213 |
1142 |
14.47 |
|
|
|
| 15 |
A |
1191 |
1121 |
83.90 |
|
|
|
| 16 |
A |
1061 |
999 |
6.85 |
|
|
|
| 17 |
A |
971 |
914 |
30.66 |
|
|
|
| 18 |
A |
916 |
862 |
21.91 |
|
|
|
| 19 |
A |
597 |
562 |
2.14 |
|
|
|
| 20 |
A |
402 |
378 |
2.81 |
|
|
|
| 21 |
A |
353 |
332 |
1.31 |
|
|
|
| 22 |
A |
258 |
243 |
10.93 |
|
|
|
| 23 |
A |
177 |
167 |
4.28 |
|
|
|
| 24 |
A |
118 |
111 |
8.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17771.5 cm
-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 16730.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.560 |
-0.778 |
0.140 |
| O2 |
1.082 |
0.431 |
-0.445 |
| C3 |
-0.053 |
0.936 |
0.213 |
| C4 |
-1.244 |
0.061 |
0.038 |
| N5 |
-2.168 |
-0.640 |
-0.098 |
| H6 |
2.463 |
-1.043 |
-0.403 |
| H7 |
0.828 |
-1.584 |
0.044 |
| H8 |
1.798 |
-0.633 |
1.198 |
| H9 |
-0.268 |
1.911 |
-0.222 |
| H10 |
0.120 |
1.063 |
1.288 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4251 | 2.3550 | 2.9288 | 3.7379 | 1.0864 | 1.0931 | 1.0948 | 3.2713 | 2.6041 |
O2 | 1.4251 | | 1.4066 | 2.4048 | 3.4398 | 2.0198 | 2.0888 | 2.0842 | 2.0159 | 2.0806 | C3 | 2.3550 | 1.4066 | | 1.4880 | 2.6555 | 3.2601 | 2.6751 | 2.6194 | 1.0888 | 1.0962 | C4 | 2.9288 | 2.4048 | 1.4880 | | 1.1676 | 3.8934 | 2.6459 | 3.3296 | 2.1076 | 2.1044 | N5 | 3.7379 | 3.4398 | 2.6555 | 1.1676 | | 4.6587 | 3.1442 | 4.1726 | 3.1832 | 3.1711 | H6 | 1.0864 | 2.0198 | 3.2601 | 3.8934 | 4.6587 | | 1.7797 | 1.7814 | 4.0269 | 3.5754 | H7 | 1.0931 | 2.0888 | 2.6751 | 2.6459 | 3.1442 | 1.7797 | | 1.7830 | 3.6723 | 3.0093 | H8 | 1.0948 | 2.0842 | 2.6194 | 3.3296 | 4.1726 | 1.7814 | 1.7830 | | 3.5717 | 2.3876 | H9 | 3.2713 | 2.0159 | 1.0888 | 2.1076 | 3.1832 | 4.0269 | 3.6723 | 3.5717 | | 1.7746 | H10 | 2.6041 | 2.0806 | 1.0962 | 2.1044 | 3.1711 | 3.5754 | 3.0093 | 2.3876 | 1.7746 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
112.541 |
|
O2 |
C1 |
H6 |
106.285 |
| O2 |
C1 |
H7 |
111.417 |
|
O2 |
C1 |
H8 |
110.929 |
| O2 |
C3 |
C4 |
112.325 |
|
O2 |
C3 |
H9 |
107.084 |
| O2 |
C3 |
H10 |
111.873 |
|
C3 |
C4 |
N5 |
179.094 |
| C4 |
C3 |
H9 |
108.761 |
|
C4 |
C3 |
H10 |
108.084 |
| H6 |
C1 |
H7 |
109.478 |
|
H6 |
C1 |
H8 |
109.512 |
| H7 |
C1 |
H8 |
109.168 |
|
H9 |
C3 |
H10 |
108.617 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
