Jump to
S1C2
Energy calculated at B2PLYP=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -247.025105 |
| Energy at 298.15K | -247.030676 |
| HF Energy | -246.769182 |
| Nuclear repulsion energy | 156.811162 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3200 |
3200 |
18.18 |
|
|
|
| 2 |
A' |
3038 |
3038 |
52.44 |
|
|
|
| 3 |
A' |
3022 |
3022 |
23.39 |
|
|
|
| 4 |
A' |
2318 |
2318 |
1.73 |
|
|
|
| 5 |
A' |
1553 |
1553 |
2.76 |
|
|
|
| 6 |
A' |
1533 |
1533 |
7.04 |
|
|
|
| 7 |
A' |
1508 |
1508 |
1.36 |
|
|
|
| 8 |
A' |
1436 |
1436 |
47.10 |
|
|
|
| 9 |
A' |
1242 |
1242 |
47.19 |
|
|
|
| 10 |
A' |
1175 |
1175 |
130.97 |
|
|
|
| 11 |
A' |
1005 |
1005 |
32.06 |
|
|
|
| 12 |
A' |
962 |
962 |
7.39 |
|
|
|
| 13 |
A' |
545 |
545 |
1.42 |
|
|
|
| 14 |
A' |
381 |
381 |
2.25 |
|
|
|
| 15 |
A' |
181 |
181 |
2.31 |
|
|
|
| 16 |
A" |
3098 |
3098 |
50.82 |
|
|
|
| 17 |
A" |
3057 |
3057 |
33.38 |
|
|
|
| 18 |
A" |
1526 |
1526 |
6.79 |
|
|
|
| 19 |
A" |
1276 |
1276 |
3.32 |
|
|
|
| 20 |
A" |
1199 |
1199 |
3.42 |
|
|
|
| 21 |
A" |
1051 |
1051 |
1.84 |
|
|
|
| 22 |
A" |
373 |
373 |
1.93 |
|
|
|
| 23 |
A" |
231 |
231 |
4.43 |
|
|
|
| 24 |
A" |
88 |
88 |
0.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17498.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 17498.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.133 |
-0.357 |
0.000 |
| O2 |
-0.737 |
-0.606 |
0.000 |
| C3 |
0.000 |
0.593 |
0.000 |
| C4 |
1.432 |
0.270 |
0.000 |
| N5 |
2.578 |
0.047 |
0.000 |
| H6 |
-2.622 |
-1.328 |
0.000 |
| H7 |
-2.441 |
0.203 |
0.891 |
| H8 |
-2.441 |
0.203 |
-0.891 |
| H9 |
-0.220 |
1.206 |
-0.886 |
| H10 |
-0.220 |
1.206 |
0.886 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4179 | 2.3348 | 3.6199 | 4.7276 | 1.0874 | 1.0965 | 1.0965 | 2.6238 | 2.6238 |
O2 | 1.4179 | | 1.4077 | 2.3397 | 3.3783 | 2.0184 | 2.0863 | 2.0863 | 2.0820 | 2.0820 | C3 | 2.3348 | 1.4077 | | 1.4685 | 2.6347 | 3.2504 | 2.6277 | 2.6277 | 1.0991 | 1.0991 | C4 | 3.6199 | 2.3397 | 1.4685 | | 1.1665 | 4.3579 | 3.9752 | 3.9752 | 2.0955 | 2.0955 | N5 | 4.7276 | 3.3783 | 2.6347 | 1.1665 | | 5.3782 | 5.0994 | 5.0994 | 3.1550 | 3.1550 | H6 | 1.0874 | 2.0184 | 3.2504 | 4.3579 | 5.3782 | | 1.7805 | 1.7805 | 3.6018 | 3.6018 | H7 | 1.0965 | 2.0863 | 2.6277 | 3.9752 | 5.0994 | 1.7805 | | 1.7820 | 3.0157 | 2.4369 | H8 | 1.0965 | 2.0863 | 2.6277 | 3.9752 | 5.0994 | 1.7805 | 1.7820 | | 2.4369 | 3.0157 | H9 | 2.6238 | 2.0820 | 1.0991 | 2.0955 | 3.1550 | 3.6018 | 3.0157 | 2.4369 | | 1.7710 | H10 | 2.6238 | 2.0820 | 1.0991 | 2.0955 | 3.1550 | 3.6018 | 2.4369 | 3.0157 | 1.7710 | |
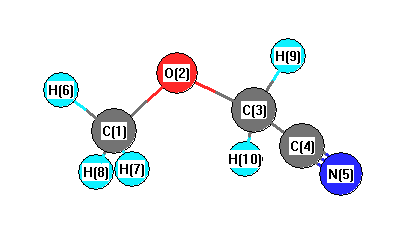 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
111.441 |
|
O2 |
C1 |
H6 |
106.608 |
| O2 |
C1 |
H7 |
111.505 |
|
O2 |
C1 |
H8 |
111.505 |
| O2 |
C3 |
C4 |
108.851 |
|
O2 |
C3 |
H9 |
111.720 |
| O2 |
C3 |
H10 |
111.720 |
|
C3 |
C4 |
N5 |
178.290 |
| C4 |
C3 |
H9 |
108.549 |
|
C4 |
C3 |
H10 |
108.549 |
| H6 |
C1 |
H7 |
109.236 |
|
H6 |
C1 |
H8 |
109.236 |
| H7 |
C1 |
H8 |
108.695 |
|
H9 |
C3 |
H10 |
107.354 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -247.027609 |
| Energy at 298.15K | -247.033281 |
| HF Energy | -246.771347 |
| Nuclear repulsion energy | 159.664174 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3201 |
3201 |
18.79 |
|
|
|
| 2 |
A |
3157 |
3157 |
6.08 |
|
|
|
| 3 |
A |
3119 |
3119 |
37.55 |
|
|
|
| 4 |
A |
3049 |
3049 |
42.29 |
|
|
|
| 5 |
A |
3045 |
3045 |
27.54 |
|
|
|
| 6 |
A |
2300 |
2300 |
1.86 |
|
|
|
| 7 |
A |
1546 |
1546 |
8.85 |
|
|
|
| 8 |
A |
1528 |
1528 |
5.93 |
|
|
|
| 9 |
A |
1517 |
1517 |
2.80 |
|
|
|
| 10 |
A |
1505 |
1505 |
0.49 |
|
|
|
| 11 |
A |
1411 |
1411 |
19.42 |
|
|
|
| 12 |
A |
1328 |
1328 |
6.70 |
|
|
|
| 13 |
A |
1235 |
1235 |
38.41 |
|
|
|
| 14 |
A |
1201 |
1201 |
8.62 |
|
|
|
| 15 |
A |
1172 |
1172 |
108.33 |
|
|
|
| 16 |
A |
1047 |
1047 |
6.96 |
|
|
|
| 17 |
A |
955 |
955 |
31.52 |
|
|
|
| 18 |
A |
908 |
908 |
20.01 |
|
|
|
| 19 |
A |
599 |
599 |
2.05 |
|
|
|
| 20 |
A |
395 |
395 |
2.61 |
|
|
|
| 21 |
A |
361 |
361 |
1.29 |
|
|
|
| 22 |
A |
258 |
258 |
10.35 |
|
|
|
| 23 |
A |
179 |
179 |
4.08 |
|
|
|
| 24 |
A |
119 |
119 |
8.83 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17566.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 17566.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.563 |
-0.775 |
0.138 |
| O2 |
1.081 |
0.432 |
-0.440 |
| C3 |
-0.060 |
0.934 |
0.210 |
| C4 |
-1.245 |
0.064 |
0.038 |
| N5 |
-2.163 |
-0.645 |
-0.097 |
| H6 |
2.463 |
-1.040 |
-0.413 |
| H7 |
0.832 |
-1.584 |
0.048 |
| H8 |
1.813 |
-0.634 |
1.196 |
| H9 |
-0.274 |
1.909 |
-0.229 |
| H10 |
0.108 |
1.070 |
1.286 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4223 | 2.3576 | 2.9323 | 3.7354 | 1.0873 | 1.0942 | 1.0961 | 3.2725 | 2.6150 |
O2 | 1.4223 | | 1.4049 | 2.4026 | 3.4347 | 2.0194 | 2.0892 | 2.0853 | 2.0146 | 2.0811 | C3 | 2.3576 | 1.4049 | | 1.4805 | 2.6477 | 3.2627 | 2.6758 | 2.6341 | 1.0903 | 1.0981 | C4 | 2.9323 | 2.4026 | 1.4805 | | 1.1674 | 3.8945 | 2.6509 | 3.3435 | 2.1021 | 2.0985 | N5 | 3.7354 | 3.4347 | 2.6477 | 1.1674 | | 4.6527 | 3.1415 | 4.1804 | 3.1793 | 3.1645 | H6 | 1.0873 | 2.0194 | 3.2627 | 3.8945 | 4.6527 | | 1.7797 | 1.7814 | 4.0268 | 3.5889 | H7 | 1.0942 | 2.0892 | 2.6758 | 2.6509 | 3.1415 | 1.7797 | | 1.7833 | 3.6738 | 3.0164 | H8 | 1.0961 | 2.0853 | 2.6341 | 3.3435 | 4.1804 | 1.7814 | 1.7833 | | 3.5850 | 2.4121 | H9 | 3.2725 | 2.0146 | 1.0903 | 2.1021 | 3.1793 | 4.0268 | 3.6738 | 3.5850 | | 1.7738 | H10 | 2.6150 | 2.0811 | 1.0981 | 2.0985 | 3.1645 | 3.5889 | 3.0164 | 2.4121 | 1.7738 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
112.999 |
|
O2 |
C1 |
H6 |
106.385 |
| O2 |
C1 |
H7 |
111.569 |
|
O2 |
C1 |
H8 |
111.132 |
| O2 |
C3 |
C4 |
112.725 |
|
O2 |
C3 |
H9 |
107.008 |
| O2 |
C3 |
H10 |
111.906 |
|
C3 |
C4 |
N5 |
178.606 |
| C4 |
C3 |
H9 |
108.761 |
|
C4 |
C3 |
H10 |
108.018 |
| H6 |
C1 |
H7 |
109.337 |
|
H6 |
C1 |
H8 |
109.350 |
| H7 |
C1 |
H8 |
109.010 |
|
H9 |
C3 |
H10 |
108.299 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
