Jump to
S1C2
Energy calculated at CCD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -246.572418 |
| Energy at 298.15K | -246.577989 |
| HF Energy | -245.797581 |
| Nuclear repulsion energy | 156.866534 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3239 |
3037 |
20.29 |
|
|
|
| 2 |
A' |
3084 |
2891 |
49.14 |
|
|
|
| 3 |
A' |
3075 |
2883 |
18.18 |
|
|
|
| 4 |
A' |
2395 |
2245 |
2.45 |
|
|
|
| 5 |
A' |
1580 |
1481 |
1.20 |
|
|
|
| 6 |
A' |
1564 |
1466 |
6.55 |
|
|
|
| 7 |
A' |
1533 |
1437 |
1.24 |
|
|
|
| 8 |
A' |
1462 |
1371 |
53.17 |
|
|
|
| 9 |
A' |
1269 |
1190 |
69.45 |
|
|
|
| 10 |
A' |
1205 |
1129 |
97.35 |
|
|
|
| 11 |
A' |
1032 |
968 |
30.53 |
|
|
|
| 12 |
A' |
977 |
916 |
9.42 |
|
|
|
| 13 |
A' |
547 |
513 |
1.15 |
|
|
|
| 14 |
A' |
386 |
362 |
2.33 |
|
|
|
| 15 |
A' |
180 |
169 |
2.47 |
|
|
|
| 16 |
A" |
3150 |
2953 |
49.21 |
|
|
|
| 17 |
A" |
3117 |
2922 |
30.92 |
|
|
|
| 18 |
A" |
1552 |
1455 |
5.61 |
|
|
|
| 19 |
A" |
1296 |
1215 |
3.40 |
|
|
|
| 20 |
A" |
1213 |
1138 |
4.95 |
|
|
|
| 21 |
A" |
1069 |
1003 |
2.82 |
|
|
|
| 22 |
A" |
362 |
339 |
2.30 |
|
|
|
| 23 |
A" |
224 |
210 |
4.67 |
|
|
|
| 24 |
A" |
85 |
79 |
0.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17796.4 cm
-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 16685.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.134 |
-0.344 |
0.000 |
| O2 |
-0.739 |
-0.601 |
0.000 |
| C3 |
0.000 |
0.597 |
0.000 |
| C4 |
1.436 |
0.259 |
0.000 |
| N5 |
2.576 |
0.030 |
0.000 |
| H6 |
-2.626 |
-1.312 |
0.000 |
| H7 |
-2.435 |
0.216 |
0.890 |
| H8 |
-2.435 |
0.216 |
-0.890 |
| H9 |
-0.216 |
1.206 |
-0.885 |
| H10 |
-0.216 |
1.206 |
0.885 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4186 | 2.3326 | 3.6205 | 4.7252 | 1.0859 | 1.0943 | 1.0943 | 2.6209 | 2.6209 |
O2 | 1.4186 | | 1.4077 | 2.3387 | 3.3746 | 2.0169 | 2.0827 | 2.0827 | 2.0796 | 2.0796 | C3 | 2.3326 | 1.4077 | | 1.4748 | 2.6378 | 3.2470 | 2.6208 | 2.6208 | 1.0963 | 1.0963 | C4 | 3.6205 | 2.3387 | 1.4748 | | 1.1633 | 4.3554 | 3.9722 | 3.9722 | 2.0996 | 2.0996 | N5 | 4.7252 | 3.3746 | 2.6378 | 1.1633 | | 5.3728 | 5.0933 | 5.0933 | 3.1565 | 3.1565 | H6 | 1.0859 | 2.0169 | 3.2470 | 4.3554 | 5.3728 | | 1.7790 | 1.7790 | 3.5970 | 3.5970 | H7 | 1.0943 | 2.0827 | 2.6208 | 3.9722 | 5.0933 | 1.7790 | | 1.7808 | 3.0100 | 2.4304 | H8 | 1.0943 | 2.0827 | 2.6208 | 3.9722 | 5.0933 | 1.7790 | 1.7808 | | 2.4304 | 3.0100 | H9 | 2.6209 | 2.0796 | 1.0963 | 2.0996 | 3.1565 | 3.5970 | 3.0100 | 2.4304 | | 1.7707 | H10 | 2.6209 | 2.0796 | 1.0963 | 2.0996 | 3.1565 | 3.5970 | 2.4304 | 3.0100 | 1.7707 | |
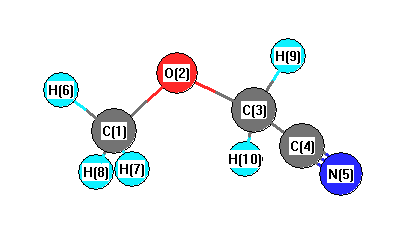 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
111.244 |
|
O2 |
C1 |
H6 |
106.527 |
| O2 |
C1 |
H7 |
111.300 |
|
O2 |
C1 |
H8 |
111.300 |
| O2 |
C3 |
C4 |
108.432 |
|
O2 |
C3 |
H9 |
111.700 |
| O2 |
C3 |
H10 |
111.700 |
|
C3 |
C4 |
N5 |
178.142 |
| C4 |
C3 |
H9 |
108.599 |
|
C4 |
C3 |
H10 |
108.599 |
| H6 |
C1 |
H7 |
109.377 |
|
H6 |
C1 |
H8 |
109.377 |
| H7 |
C1 |
H8 |
108.912 |
|
H9 |
C3 |
H10 |
107.725 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -246.574780 |
| Energy at 298.15K | -246.580480 |
| HF Energy | -245.799540 |
| Nuclear repulsion energy | 159.763554 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3240 |
3038 |
21.02 |
|
|
|
| 2 |
A |
3201 |
3002 |
7.58 |
|
|
|
| 3 |
A |
3167 |
2969 |
38.52 |
|
|
|
| 4 |
A |
3100 |
2907 |
29.81 |
|
|
|
| 5 |
A |
3091 |
2898 |
32.71 |
|
|
|
| 6 |
A |
2381 |
2233 |
3.06 |
|
|
|
| 7 |
A |
1572 |
1474 |
7.76 |
|
|
|
| 8 |
A |
1556 |
1459 |
5.15 |
|
|
|
| 9 |
A |
1547 |
1451 |
1.75 |
|
|
|
| 10 |
A |
1532 |
1437 |
0.29 |
|
|
|
| 11 |
A |
1436 |
1346 |
23.89 |
|
|
|
| 12 |
A |
1348 |
1264 |
7.39 |
|
|
|
| 13 |
A |
1260 |
1181 |
67.84 |
|
|
|
| 14 |
A |
1219 |
1143 |
18.17 |
|
|
|
| 15 |
A |
1202 |
1127 |
66.41 |
|
|
|
| 16 |
A |
1066 |
999 |
7.23 |
|
|
|
| 17 |
A |
982 |
921 |
30.12 |
|
|
|
| 18 |
A |
926 |
868 |
21.36 |
|
|
|
| 19 |
A |
603 |
565 |
2.22 |
|
|
|
| 20 |
A |
404 |
378 |
2.93 |
|
|
|
| 21 |
A |
355 |
333 |
1.28 |
|
|
|
| 22 |
A |
259 |
243 |
11.36 |
|
|
|
| 23 |
A |
177 |
166 |
4.19 |
|
|
|
| 24 |
A |
118 |
111 |
8.79 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17870.8 cm
-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 16755.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.560 |
-0.775 |
0.139 |
| O2 |
1.081 |
0.431 |
-0.443 |
| C3 |
-0.055 |
0.934 |
0.212 |
| C4 |
-1.243 |
0.059 |
0.037 |
| N5 |
-2.165 |
-0.639 |
-0.097 |
| H6 |
2.463 |
-1.039 |
-0.403 |
| H7 |
0.830 |
-1.583 |
0.042 |
| H8 |
1.799 |
-0.632 |
1.197 |
| H9 |
-0.271 |
1.908 |
-0.224 |
| H10 |
0.118 |
1.064 |
1.286 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4223 | 2.3531 | 2.9267 | 3.7345 | 1.0859 | 1.0924 | 1.0940 | 3.2686 | 2.6036 |
O2 | 1.4223 | | 1.4047 | 2.4023 | 3.4349 | 2.0178 | 2.0861 | 2.0816 | 2.0143 | 2.0780 | C3 | 2.3531 | 1.4047 | | 1.4860 | 2.6498 | 3.2578 | 2.6734 | 2.6190 | 1.0884 | 1.0954 | C4 | 2.9267 | 2.4023 | 1.4860 | | 1.1639 | 3.8909 | 2.6449 | 3.3283 | 2.1053 | 2.1027 | N5 | 3.7345 | 3.4349 | 2.6498 | 1.1639 | | 4.6552 | 3.1430 | 4.1692 | 3.1768 | 3.1657 | H6 | 1.0859 | 2.0178 | 3.2578 | 3.8909 | 4.6552 | | 1.7780 | 1.7796 | 4.0237 | 3.5743 | H7 | 1.0924 | 2.0861 | 2.6734 | 2.6449 | 3.1430 | 1.7780 | | 1.7818 | 3.6698 | 3.0096 | H8 | 1.0940 | 2.0816 | 2.6190 | 3.3283 | 4.1692 | 1.7796 | 1.7818 | | 3.5708 | 2.3893 | H9 | 3.2686 | 2.0143 | 1.0884 | 2.1053 | 3.1768 | 4.0237 | 3.6698 | 3.5708 | | 1.7726 | H10 | 2.6036 | 2.0780 | 1.0954 | 2.1027 | 3.1657 | 3.5743 | 3.0096 | 2.3893 | 1.7726 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
112.679 |
|
O2 |
C1 |
H6 |
106.350 |
| O2 |
C1 |
H7 |
111.437 |
|
O2 |
C1 |
H8 |
110.960 |
| O2 |
C3 |
C4 |
112.380 |
|
O2 |
C3 |
H9 |
107.110 |
| O2 |
C3 |
H10 |
111.838 |
|
C3 |
C4 |
N5 |
179.185 |
| C4 |
C3 |
H9 |
108.749 |
|
C4 |
C3 |
H10 |
108.134 |
| H6 |
C1 |
H7 |
109.420 |
|
H6 |
C1 |
H8 |
109.447 |
| H7 |
C1 |
H8 |
109.171 |
|
H9 |
C3 |
H10 |
108.527 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
