Jump to
S1C2
Energy calculated at TPSSh/6-31G**
| | hartrees |
|---|
| Energy at 0K | -247.271740 |
| Energy at 298.15K | -247.277266 |
| HF Energy | -247.271740 |
| Nuclear repulsion energy | 156.575498 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3157 |
3026 |
20.74 |
|
|
|
| 2 |
A' |
2994 |
2870 |
59.10 |
|
|
|
| 3 |
A' |
2977 |
2854 |
26.76 |
|
|
|
| 4 |
A' |
2356 |
2258 |
0.26 |
|
|
|
| 5 |
A' |
1538 |
1475 |
3.27 |
|
|
|
| 6 |
A' |
1517 |
1454 |
6.08 |
|
|
|
| 7 |
A' |
1492 |
1430 |
2.92 |
|
|
|
| 8 |
A' |
1412 |
1354 |
42.88 |
|
|
|
| 9 |
A' |
1222 |
1171 |
32.05 |
|
|
|
| 10 |
A' |
1148 |
1101 |
151.27 |
|
|
|
| 11 |
A' |
988 |
947 |
32.37 |
|
|
|
| 12 |
A' |
948 |
908 |
6.95 |
|
|
|
| 13 |
A' |
542 |
520 |
1.36 |
|
|
|
| 14 |
A' |
368 |
353 |
1.91 |
|
|
|
| 15 |
A' |
177 |
170 |
2.17 |
|
|
|
| 16 |
A" |
3047 |
2922 |
58.28 |
|
|
|
| 17 |
A" |
3006 |
2882 |
41.02 |
|
|
|
| 18 |
A" |
1513 |
1451 |
6.24 |
|
|
|
| 19 |
A" |
1256 |
1204 |
3.06 |
|
|
|
| 20 |
A" |
1182 |
1133 |
3.32 |
|
|
|
| 21 |
A" |
1033 |
990 |
1.70 |
|
|
|
| 22 |
A" |
375 |
360 |
1.47 |
|
|
|
| 23 |
A" |
228 |
219 |
4.27 |
|
|
|
| 24 |
A" |
86 |
83 |
0.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17281.0 cm
-1
Scaled (by 0.9587) Zero Point Vibrational Energy (zpe) 16567.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.136 |
-0.361 |
0.000 |
| O2 |
-0.736 |
-0.614 |
0.000 |
| C3 |
0.000 |
0.591 |
0.000 |
| C4 |
1.435 |
0.275 |
0.000 |
| N5 |
2.578 |
0.058 |
0.000 |
| H6 |
-2.624 |
-1.336 |
0.000 |
| H7 |
-2.443 |
0.201 |
0.894 |
| H8 |
-2.443 |
0.201 |
-0.894 |
| H9 |
-0.222 |
1.205 |
-0.889 |
| H10 |
-0.222 |
1.205 |
0.889 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4223 | 2.3384 | 3.6269 | 4.7325 | 1.0903 | 1.1001 | 1.1001 | 2.6271 | 2.6271 |
O2 | 1.4223 | | 1.4125 | 2.3461 | 3.3817 | 2.0212 | 2.0926 | 2.0926 | 2.0885 | 2.0885 | C3 | 2.3384 | 1.4125 | | 1.4696 | 2.6330 | 3.2557 | 2.6311 | 2.6311 | 1.1026 | 1.1026 | C4 | 3.6269 | 2.3461 | 1.4696 | | 1.1637 | 4.3670 | 3.9810 | 3.9810 | 2.0981 | 2.0981 | N5 | 4.7325 | 3.3817 | 2.6330 | 1.1637 | | 5.3857 | 5.1027 | 5.1027 | 3.1544 | 3.1544 | H6 | 1.0903 | 2.0212 | 3.2557 | 4.3670 | 5.3857 | | 1.7870 | 1.7870 | 3.6072 | 3.6072 | H7 | 1.1001 | 2.0926 | 2.6311 | 3.9810 | 5.1027 | 1.7870 | | 1.7889 | 3.0201 | 2.4373 | H8 | 1.1001 | 2.0926 | 2.6311 | 3.9810 | 5.1027 | 1.7870 | 1.7889 | | 2.4373 | 3.0201 | H9 | 2.6271 | 2.0885 | 1.1026 | 2.0981 | 3.1544 | 3.6072 | 3.0201 | 2.4373 | | 1.7777 | H10 | 2.6271 | 2.0885 | 1.1026 | 2.0981 | 3.1544 | 3.6072 | 2.4373 | 3.0201 | 1.7777 | |
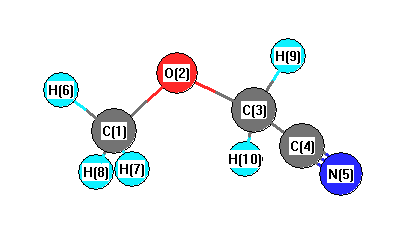 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
111.151 |
|
O2 |
C1 |
H6 |
106.359 |
| O2 |
C1 |
H7 |
111.478 |
|
O2 |
C1 |
H8 |
111.478 |
| O2 |
C3 |
C4 |
108.968 |
|
O2 |
C3 |
H9 |
111.693 |
| O2 |
C3 |
H10 |
111.693 |
|
C3 |
C4 |
N5 |
178.324 |
| C4 |
C3 |
H9 |
108.470 |
|
C4 |
C3 |
H10 |
108.470 |
| H6 |
C1 |
H7 |
109.337 |
|
H6 |
C1 |
H8 |
109.337 |
| H7 |
C1 |
H8 |
108.799 |
|
H9 |
C3 |
H10 |
107.445 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.114 |
|
|
|
| 2 |
O |
-0.472 |
|
|
|
| 3 |
C |
0.018 |
|
|
|
| 4 |
C |
0.354 |
|
|
|
| 5 |
N |
-0.482 |
|
|
|
| 6 |
H |
0.153 |
|
|
|
| 7 |
H |
0.117 |
|
|
|
| 8 |
H |
0.117 |
|
|
|
| 9 |
H |
0.154 |
|
|
|
| 10 |
H |
0.154 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.308 |
1.754 |
0.000 |
4.652 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.207 |
0.014 |
0.000 |
| y |
0.014 |
4.700 |
0.000 |
| z |
0.000 |
0.000 |
4.410 |
<r2> (average value of r
2) Å
2
| <r2> |
143.236 |
| (<r2>)1/2 |
11.968 |
Jump to
S1C1
Energy calculated at TPSSh/6-31G**
| | hartrees |
|---|
| Energy at 0K | -247.274558 |
| Energy at 298.15K | -247.280164 |
| HF Energy | -247.274558 |
| Nuclear repulsion energy | 159.449045 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3161 |
3031 |
21.52 |
|
|
|
| 2 |
A |
3116 |
2987 |
6.72 |
|
|
|
| 3 |
A |
3070 |
2944 |
42.08 |
|
|
|
| 4 |
A |
3005 |
2881 |
49.74 |
|
|
|
| 5 |
A |
2999 |
2875 |
30.91 |
|
|
|
| 6 |
A |
2336 |
2240 |
0.03 |
|
|
|
| 7 |
A |
1532 |
1469 |
8.46 |
|
|
|
| 8 |
A |
1514 |
1451 |
5.91 |
|
|
|
| 9 |
A |
1499 |
1437 |
2.27 |
|
|
|
| 10 |
A |
1487 |
1426 |
0.97 |
|
|
|
| 11 |
A |
1389 |
1331 |
16.35 |
|
|
|
| 12 |
A |
1311 |
1256 |
5.99 |
|
|
|
| 13 |
A |
1214 |
1164 |
25.28 |
|
|
|
| 14 |
A |
1182 |
1133 |
5.92 |
|
|
|
| 15 |
A |
1144 |
1097 |
125.01 |
|
|
|
| 16 |
A |
1030 |
987 |
7.21 |
|
|
|
| 17 |
A |
937 |
899 |
31.74 |
|
|
|
| 18 |
A |
895 |
858 |
19.91 |
|
|
|
| 19 |
A |
593 |
568 |
1.89 |
|
|
|
| 20 |
A |
380 |
364 |
1.91 |
|
|
|
| 21 |
A |
358 |
343 |
1.61 |
|
|
|
| 22 |
A |
249 |
238 |
9.94 |
|
|
|
| 23 |
A |
174 |
167 |
3.73 |
|
|
|
| 24 |
A |
117 |
112 |
8.89 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17344.8 cm
-1
Scaled (by 0.9587) Zero Point Vibrational Energy (zpe) 16628.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.562 |
-0.779 |
0.139 |
| O2 |
1.083 |
0.436 |
-0.439 |
| C3 |
-0.063 |
0.938 |
0.210 |
| C4 |
-1.247 |
0.064 |
0.039 |
| N5 |
-2.158 |
-0.649 |
-0.099 |
| H6 |
2.458 |
-1.047 |
-0.421 |
| H7 |
0.821 |
-1.584 |
0.051 |
| H8 |
1.819 |
-0.638 |
1.198 |
| H9 |
-0.277 |
1.912 |
-0.236 |
| H10 |
0.104 |
1.078 |
1.290 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4272 | 2.3642 | 2.9343 | 3.7298 | 1.0902 | 1.0976 | 1.0995 | 3.2804 | 2.6259 |
O2 | 1.4272 | | 1.4096 | 2.4078 | 3.4349 | 2.0219 | 2.0943 | 2.0917 | 2.0177 | 2.0881 | C3 | 2.3642 | 1.4096 | | 1.4819 | 2.6463 | 3.2697 | 2.6764 | 2.6456 | 1.0931 | 1.1015 | C4 | 2.9343 | 2.4078 | 1.4819 | | 1.1647 | 3.8956 | 2.6442 | 3.3523 | 2.1057 | 2.1023 | N5 | 3.7298 | 3.4349 | 2.6463 | 1.1647 | | 4.6448 | 3.1260 | 4.1831 | 3.1808 | 3.1666 | H6 | 1.0902 | 2.0219 | 3.2697 | 3.8956 | 4.6448 | | 1.7866 | 1.7882 | 4.0335 | 3.6028 | H7 | 1.0976 | 2.0943 | 2.6764 | 2.6442 | 3.1260 | 1.7866 | | 1.7905 | 3.6754 | 3.0217 | H8 | 1.0995 | 2.0917 | 2.6456 | 3.3523 | 4.1831 | 1.7882 | 1.7905 | | 3.5990 | 2.4272 | H9 | 3.2804 | 2.0177 | 1.0931 | 2.1057 | 3.1808 | 4.0335 | 3.6754 | 3.5990 | | 1.7806 | H10 | 2.6259 | 2.0881 | 1.1015 | 2.1023 | 3.1666 | 3.6028 | 3.0217 | 2.4272 | 1.7806 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
112.897 |
|
O2 |
C1 |
H6 |
106.088 |
| O2 |
C1 |
H7 |
111.428 |
|
O2 |
C1 |
H8 |
111.095 |
| O2 |
C3 |
C4 |
112.737 |
|
O2 |
C3 |
H9 |
106.775 |
| O2 |
C3 |
H10 |
111.936 |
|
C3 |
C4 |
N5 |
178.389 |
| C4 |
C3 |
H9 |
108.785 |
|
C4 |
C3 |
H10 |
108.032 |
| H6 |
C1 |
H7 |
109.501 |
|
H6 |
C1 |
H8 |
109.499 |
| H7 |
C1 |
H8 |
109.172 |
|
H9 |
C3 |
H10 |
108.457 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.130 |
|
|
|
| 2 |
O |
-0.461 |
|
|
|
| 3 |
C |
-0.018 |
|
|
|
| 4 |
C |
0.345 |
|
|
|
| 5 |
N |
-0.480 |
|
|
|
| 6 |
H |
0.150 |
|
|
|
| 7 |
H |
0.140 |
|
|
|
| 8 |
H |
0.116 |
|
|
|
| 9 |
H |
0.183 |
|
|
|
| 10 |
H |
0.156 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.479 |
1.221 |
1.403 |
3.099 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.918 |
0.479 |
0.186 |
| y |
0.479 |
5.560 |
0.211 |
| z |
0.186 |
0.211 |
4.410 |
<r2> (average value of r
2) Å
2
| <r2> |
122.945 |
| (<r2>)1/2 |
11.088 |
