Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Geometric Data calculated at MP2=FULL/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -235.087323 |
| Energy at 298.15K | -235.100982 |
| Nuclear repulsion energy | 256.388224 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3228 |
3016 |
50.08 |
|
|
|
| 2 |
A' |
3214 |
3003 |
38.40 |
|
|
|
| 3 |
A' |
3210 |
3000 |
20.39 |
|
|
|
| 4 |
A' |
3198 |
2989 |
15.31 |
|
|
|
| 5 |
A' |
3160 |
2952 |
29.16 |
|
|
|
| 6 |
A' |
3136 |
2930 |
6.78 |
|
|
|
| 7 |
A' |
3113 |
2909 |
24.88 |
|
|
|
| 8 |
A' |
3108 |
2904 |
23.65 |
|
|
|
| 9 |
A' |
1570 |
1467 |
1.69 |
|
|
|
| 10 |
A' |
1563 |
1461 |
1.33 |
|
|
|
| 11 |
A' |
1554 |
1452 |
5.58 |
|
|
|
| 12 |
A' |
1538 |
1437 |
0.28 |
|
|
|
| 13 |
A' |
1466 |
1370 |
1.06 |
|
|
|
| 14 |
A' |
1451 |
1356 |
7.42 |
|
|
|
| 15 |
A' |
1364 |
1275 |
5.08 |
|
|
|
| 16 |
A' |
1352 |
1263 |
1.79 |
|
|
|
| 17 |
A' |
1277 |
1193 |
0.82 |
|
|
|
| 18 |
A' |
1245 |
1163 |
0.14 |
|
|
|
| 19 |
A' |
1050 |
981 |
0.76 |
|
|
|
| 20 |
A' |
1026 |
959 |
0.35 |
|
|
|
| 21 |
A' |
986 |
921 |
0.28 |
|
|
|
| 22 |
A' |
959 |
896 |
0.31 |
|
|
|
| 23 |
A' |
907 |
848 |
0.51 |
|
|
|
| 24 |
A' |
729 |
681 |
0.80 |
|
|
|
| 25 |
A' |
614 |
574 |
1.34 |
|
|
|
| 26 |
A' |
404 |
378 |
0.47 |
|
|
|
| 27 |
A' |
360 |
336 |
0.02 |
|
|
|
| 28 |
A' |
193 |
180 |
0.01 |
|
|
|
| 29 |
A" |
3207 |
2997 |
49.26 |
|
|
|
| 30 |
A" |
3203 |
2993 |
0.42 |
|
|
|
| 31 |
A" |
3200 |
2991 |
10.52 |
|
|
|
| 32 |
A" |
3132 |
2927 |
57.78 |
|
|
|
| 33 |
A" |
1566 |
1463 |
5.78 |
|
|
|
| 34 |
A" |
1544 |
1443 |
0.08 |
|
|
|
| 35 |
A" |
1532 |
1432 |
1.60 |
|
|
|
| 36 |
A" |
1319 |
1233 |
0.17 |
|
|
|
| 37 |
A" |
1294 |
1209 |
0.26 |
|
|
|
| 38 |
A" |
1263 |
1180 |
0.02 |
|
|
|
| 39 |
A" |
1225 |
1144 |
1.97 |
|
|
|
| 40 |
A" |
1118 |
1045 |
0.04 |
|
|
|
| 41 |
A" |
1017 |
950 |
0.00 |
|
|
|
| 42 |
A" |
976 |
912 |
1.55 |
|
|
|
| 43 |
A" |
917 |
856 |
0.00 |
|
|
|
| 44 |
A" |
814 |
760 |
0.27 |
|
|
|
| 45 |
A" |
389 |
364 |
0.03 |
|
|
|
| 46 |
A" |
306 |
286 |
0.02 |
|
|
|
| 47 |
A" |
278 |
259 |
0.00 |
|
|
|
| 48 |
A" |
239 |
224 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37756.4 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 35279.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.108 |
0.400 |
0.000 |
| C2 |
0.403 |
-1.703 |
0.000 |
| C3 |
-0.108 |
-0.720 |
1.072 |
| C4 |
-0.108 |
-0.720 |
-1.072 |
| C5 |
1.203 |
1.171 |
0.000 |
| C6 |
-1.292 |
1.347 |
0.000 |
| H7 |
1.489 |
-1.775 |
0.000 |
| H8 |
-0.009 |
-2.709 |
0.000 |
| H9 |
0.498 |
-0.563 |
1.964 |
| H10 |
0.498 |
-0.563 |
-1.964 |
| H11 |
-1.126 |
-0.965 |
1.377 |
| H12 |
-1.126 |
-0.965 |
-1.377 |
| H13 |
2.062 |
0.501 |
0.000 |
| H14 |
-2.230 |
0.794 |
0.000 |
| H15 |
1.274 |
1.807 |
-0.883 |
| H16 |
1.274 |
1.807 |
0.883 |
| H17 |
-1.277 |
1.988 |
0.882 |
| H18 |
-1.277 |
1.988 |
-0.882 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.1640 | 1.5499 | 1.5499 | 1.5215 | 1.5159 | 2.6985 | 3.1108 | 2.2698 | 2.2698 | 2.1897 | 2.1897 | 2.1727 | 2.1584 | 2.1610 | 2.1610 | 2.1605 | 2.1605 |
C2 | 2.1640 | | 1.5418 | 1.5418 | 2.9834 | 3.4891 | 1.0880 | 1.0879 | 2.2729 | 2.2729 | 2.1859 | 2.1859 | 2.7587 | 3.6289 | 3.7227 | 3.7227 | 4.1506 | 4.1506 | C3 | 1.5499 | 1.5418 | | 2.1438 | 2.5386 | 2.6116 | 2.1941 | 2.2622 | 1.0900 | 3.0999 | 1.0904 | 2.6632 | 2.7111 | 2.8183 | 3.4809 | 2.8864 | 2.9558 | 3.5381 | C4 | 1.5499 | 1.5418 | 2.1438 | | 2.5386 | 2.6116 | 2.1941 | 2.2622 | 3.0999 | 1.0900 | 2.6632 | 1.0904 | 2.7111 | 2.8183 | 2.8864 | 3.4809 | 3.5381 | 2.9558 | C5 | 1.5215 | 2.9834 | 2.5386 | 2.5386 | | 2.5012 | 2.9604 | 4.0657 | 2.7133 | 2.7133 | 3.4475 | 3.4475 | 1.0892 | 3.4543 | 1.0904 | 1.0904 | 2.7565 | 2.7565 | C6 | 1.5159 | 3.4891 | 2.6116 | 2.6116 | 2.5012 | | 4.1809 | 4.2541 | 3.2724 | 3.2724 | 2.6961 | 2.6961 | 3.4589 | 1.0895 | 2.7519 | 2.7519 | 1.0907 | 1.0907 | H7 | 2.6985 | 1.0880 | 2.1941 | 2.1941 | 2.9604 | 4.1809 | | 1.7657 | 2.5118 | 2.5118 | 3.0641 | 3.0641 | 2.3478 | 4.5202 | 3.6962 | 3.6962 | 4.7534 | 4.7534 | H8 | 3.1108 | 1.0879 | 2.2622 | 2.2622 | 4.0657 | 4.2541 | 1.7657 | | 2.9534 | 2.9534 | 2.4868 | 2.4868 | 3.8211 | 4.1477 | 4.7778 | 4.7778 | 4.9451 | 4.9451 | H9 | 2.2698 | 2.2729 | 1.0900 | 3.0999 | 2.7133 | 3.2724 | 2.5118 | 2.9534 | | 3.9281 | 1.7732 | 3.7364 | 2.7269 | 3.6252 | 3.7847 | 2.7182 | 3.2912 | 4.2143 | H10 | 2.2698 | 2.2729 | 3.0999 | 1.0900 | 2.7133 | 3.2724 | 2.5118 | 2.9534 | 3.9281 | | 3.7364 | 1.7732 | 2.7269 | 3.6252 | 2.7182 | 3.7847 | 4.2143 | 3.2912 | H11 | 2.1897 | 2.1859 | 1.0904 | 2.6632 | 3.4475 | 2.6961 | 3.0641 | 2.4868 | 1.7732 | 3.7364 | | 2.7538 | 3.7696 | 2.4918 | 4.3071 | 3.6999 | 2.9986 | 3.7214 | H12 | 2.1897 | 2.1859 | 2.6632 | 1.0904 | 3.4475 | 2.6961 | 3.0641 | 2.4868 | 3.7364 | 1.7732 | 2.7538 | | 3.7696 | 2.4918 | 3.6999 | 4.3071 | 3.7214 | 2.9986 | H13 | 2.1727 | 2.7587 | 2.7111 | 2.7111 | 1.0892 | 3.4589 | 2.3478 | 3.8211 | 2.7269 | 2.7269 | 3.7696 | 3.7696 | | 4.3024 | 1.7626 | 1.7626 | 3.7603 | 3.7603 | H14 | 2.1584 | 3.6289 | 2.8183 | 2.8183 | 3.4543 | 1.0895 | 4.5202 | 4.1477 | 3.6252 | 3.6252 | 2.4918 | 2.4918 | 4.3024 | | 3.7530 | 3.7530 | 1.7646 | 1.7646 | H15 | 2.1610 | 3.7227 | 3.4809 | 2.8864 | 1.0904 | 2.7519 | 3.6962 | 4.7778 | 3.7847 | 2.7182 | 4.3071 | 3.6999 | 1.7626 | 3.7530 | | 1.7655 | 3.1071 | 2.5573 | H16 | 2.1610 | 3.7227 | 2.8864 | 3.4809 | 1.0904 | 2.7519 | 3.6962 | 4.7778 | 2.7182 | 3.7847 | 3.6999 | 4.3071 | 1.7626 | 3.7530 | 1.7655 | | 2.5573 | 3.1071 | H17 | 2.1605 | 4.1506 | 2.9558 | 3.5381 | 2.7565 | 1.0907 | 4.7534 | 4.9451 | 3.2912 | 4.2143 | 2.9986 | 3.7214 | 3.7603 | 1.7646 | 3.1071 | 2.5573 | | 1.7639 | H18 | 2.1605 | 4.1506 | 3.5381 | 2.9558 | 2.7565 | 1.0907 | 4.7534 | 4.9451 | 4.2143 | 3.2912 | 3.7214 | 2.9986 | 3.7603 | 1.7646 | 2.5573 | 3.1071 | 1.7639 | |
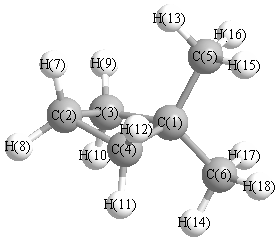 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
88.841 |
|
C1 |
C3 |
H9 |
117.524 |
| C1 |
C3 |
H11 |
110.857 |
|
C1 |
C4 |
C2 |
88.841 |
| C1 |
C4 |
H10 |
117.524 |
|
C1 |
C4 |
H12 |
110.857 |
| C1 |
C5 |
H13 |
111.581 |
|
C1 |
C5 |
H15 |
110.578 |
| C1 |
C5 |
H16 |
110.578 |
|
C1 |
C6 |
H14 |
110.821 |
| C1 |
C6 |
H17 |
110.917 |
|
C1 |
C6 |
H18 |
110.917 |
| C2 |
C3 |
H9 |
118.439 |
|
C2 |
C3 |
H11 |
111.130 |
| C2 |
C4 |
H10 |
118.439 |
|
C2 |
C4 |
H12 |
111.130 |
| C3 |
C1 |
C4 |
87.511 |
|
C3 |
C1 |
C5 |
111.482 |
| C3 |
C1 |
C6 |
116.824 |
|
C3 |
C2 |
C4 |
88.091 |
| C3 |
C2 |
H7 |
111.930 |
|
C3 |
C2 |
H8 |
117.653 |
| C4 |
C1 |
C5 |
111.482 |
|
C4 |
C1 |
C6 |
116.824 |
| C4 |
C2 |
H7 |
111.930 |
|
C4 |
C2 |
H8 |
117.653 |
| C5 |
C1 |
C6 |
110.871 |
|
H7 |
C2 |
H8 |
108.478 |
| H9 |
C3 |
H11 |
108.824 |
|
H10 |
C4 |
H12 |
108.824 |
| H13 |
C5 |
H15 |
107.932 |
|
H13 |
C5 |
H16 |
107.932 |
| H14 |
C6 |
H17 |
108.073 |
|
H14 |
C6 |
H18 |
108.073 |
| H15 |
C5 |
H16 |
108.113 |
|
H17 |
C6 |
H18 |
107.918 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
