Jump to
S1C2
S1C3
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -1590.496679 |
| Energy at 298.15K | |
| HF Energy | -1589.976869 |
| Nuclear repulsion energy | 349.285951 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
510 |
476 |
0.00 |
44.26 |
0.06 |
0.12 |
| 2 |
A1 |
209 |
196 |
0.00 |
2.85 |
0.74 |
0.85 |
| 3 |
B1 |
458 |
428 |
0.00 |
28.31 |
0.75 |
0.86 |
| 4 |
B2 |
326 |
305 |
0.34 |
9.77 |
0.75 |
0.86 |
| 5 |
E |
431 |
402 |
0.00 |
7.15 |
0.75 |
0.86 |
| 5 |
E |
431 |
402 |
0.00 |
7.15 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1182.1 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 1104.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.435 |
0.315 |
| S2 |
0.000 |
-1.435 |
0.315 |
| S3 |
-1.435 |
0.000 |
-0.315 |
| S4 |
1.435 |
0.000 |
-0.315 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.8709 | 2.1258 | 2.1258 |
S2 | 2.8709 | | 2.1258 | 2.1258 | S3 | 2.1258 | 2.1258 | | 2.8709 | S4 | 2.1258 | 2.1258 | 2.8709 | |
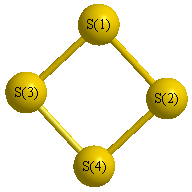 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
47.526 |
|
S1 |
S3 |
S4 |
47.526 |
| S2 |
S1 |
S3 |
47.526 |
|
S2 |
S4 |
S3 |
47.526 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -1590.573737 |
| Energy at 298.15K | |
| HF Energy | -1589.902737 |
| Nuclear repulsion energy | 313.355393 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
638 |
596 |
0.00 |
29.46 |
0.31 |
0.47 |
| 2 |
A1 |
296 |
277 |
0.18 |
0.00 |
0.33 |
0.49 |
| 3 |
A1 |
118 |
110 |
0.00 |
2052.29 |
0.31 |
0.48 |
| 4 |
A2 |
185 |
172 |
0.00 |
0.00 |
0.75 |
0.86 |
| 5 |
B2 |
696 |
650 |
35.38 |
0.00 |
0.75 |
0.86 |
| 6 |
B2 |
256 |
239 |
0.00 |
19.63 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1094.3 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 1022.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.422 |
0.972 |
| S2 |
0.000 |
-1.422 |
0.972 |
| S3 |
0.000 |
1.422 |
-0.972 |
| S4 |
0.000 |
-1.422 |
-0.972 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.8444 | 1.9430 | 3.4446 |
S2 | 2.8444 | | 3.4446 | 1.9430 | S3 | 1.9430 | 3.4446 | | 2.8442 | S4 | 3.4446 | 1.9430 | 2.8442 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
89.996 |
|
S1 |
S3 |
S4 |
90.004 |
| S2 |
S1 |
S3 |
89.996 |
|
S2 |
S4 |
S3 |
90.004 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -1590.573737 |
| Energy at 298.15K | |
| HF Energy | -1589.902686 |
| Nuclear repulsion energy | 313.336897 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
637 |
595 |
0.00 |
29.36 |
0.31 |
0.48 |
| 2 |
Ag |
119 |
111 |
0.00 |
2086.74 |
0.31 |
0.48 |
| 3 |
Au |
185 |
173 |
0.00 |
0.00 |
0.00 |
0.00 |
| 4 |
B1u |
695 |
649 |
35.99 |
0.00 |
0.00 |
0.00 |
| 5 |
B2u |
297 |
278 |
0.19 |
0.00 |
0.33 |
0.49 |
| 6 |
B3g |
256 |
239 |
0.00 |
19.69 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1094.3 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 1022.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is D2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.972 |
1.422 |
| S2 |
0.000 |
0.972 |
-1.422 |
| S3 |
0.000 |
-0.972 |
1.422 |
| S4 |
0.000 |
-0.972 |
-1.422 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.8437 | 1.9436 | 3.4444 |
S2 | 2.8437 | | 3.4444 | 1.9436 | S3 | 1.9436 | 3.4444 | | 2.8437 | S4 | 3.4444 | 1.9436 | 2.8437 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
90.000 |
|
S1 |
S3 |
S4 |
90.000 |
| S2 |
S1 |
S3 |
90.000 |
|
S2 |
S4 |
S3 |
90.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
