Jump to
S1C2
Energy calculated at QCISD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -114.112737 |
| Energy at 298.15K | -114.114153 |
| HF Energy | -113.790009 |
| Nuclear repulsion energy | 30.495840 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3809 |
3586 |
56.65 |
|
|
|
| 2 |
A' |
2903 |
2733 |
174.50 |
|
|
|
| 3 |
A' |
1544 |
1453 |
31.58 |
|
|
|
| 4 |
A' |
1333 |
1255 |
61.17 |
|
|
|
| 5 |
A' |
1231 |
1158 |
139.73 |
|
|
|
| 6 |
A" |
1105 |
1041 |
123.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5962.1 cm
-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 5612.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.011 |
0.751 |
0.000 |
| O2 |
0.011 |
-0.576 |
0.000 |
| H3 |
-1.084 |
0.967 |
0.000 |
| H4 |
0.934 |
-0.867 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
H3 |
H4 |
| C1 | | 1.3270 | 1.1154 | 1.8628 |
O2 | 1.3270 | | 1.8917 | 0.9683 | H3 | 1.1154 | 1.8917 | | 2.7267 | H4 | 1.8628 | 0.9683 | 2.7267 | |
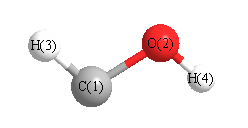 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H4 |
107.473 |
|
O2 |
C1 |
H3 |
101.168 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -114.104597 |
| Energy at 298.15K | -114.106002 |
| HF Energy | -113.781450 |
| Nuclear repulsion energy | 30.443283 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3695 |
3478 |
10.80 |
|
|
|
| 2 |
A' |
2801 |
2637 |
241.39 |
|
|
|
| 3 |
A' |
1509 |
1420 |
44.28 |
|
|
|
| 4 |
A' |
1339 |
1260 |
76.58 |
|
|
|
| 5 |
A' |
1258 |
1185 |
46.93 |
|
|
|
| 6 |
A" |
1020 |
960 |
43.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5810.9 cm
-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 5470.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.123 |
0.750 |
0.000 |
| O2 |
0.123 |
-0.572 |
0.000 |
| H3 |
-0.957 |
1.060 |
0.000 |
| H4 |
-0.758 |
-0.985 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
H3 |
H4 |
| C1 | | 1.3222 | 1.1228 | 1.9457 |
O2 | 1.3222 | | 1.9567 | 0.9727 | H3 | 1.1228 | 1.9567 | | 2.0543 | H4 | 1.9457 | 0.9727 | 2.0543 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H4 |
115.097 |
|
O2 |
C1 |
H3 |
106.021 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
