Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -114.293013 |
| Energy at 298.15K | -114.294430 |
| HF Energy | -114.293013 |
| Nuclear repulsion energy | 30.700789 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3760 |
3589 |
56.22 |
|
|
|
| 2 |
A' |
2833 |
2704 |
168.99 |
|
|
|
| 3 |
A' |
1524 |
1454 |
29.74 |
|
|
|
| 4 |
A' |
1371 |
1308 |
75.70 |
|
|
|
| 5 |
A' |
1218 |
1162 |
126.22 |
|
|
|
| 6 |
A" |
1106 |
1055 |
121.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5905.5 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 5635.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.011 |
0.743 |
0.000 |
| O2 |
0.011 |
-0.569 |
0.000 |
| H3 |
-1.090 |
0.965 |
0.000 |
| H4 |
0.933 |
-0.868 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
H3 |
H4 |
| C1 | | 1.3121 | 1.1232 | 1.8557 |
O2 | 1.3121 | | 1.8884 | 0.9691 | H3 | 1.1232 | 1.8884 | | 2.7297 | H4 | 1.8557 | 0.9691 | 2.7297 | |
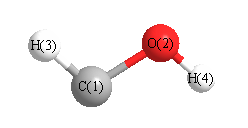 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H4 |
107.928 |
|
O2 |
C1 |
H3 |
101.408 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.064 |
|
|
|
| 2 |
O |
-0.388 |
|
|
|
| 3 |
H |
0.091 |
|
|
|
| 4 |
H |
0.360 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.384 |
-1.627 |
0.000 |
1.672 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-10.782 |
-3.496 |
0.000 |
| y |
-3.496 |
-13.457 |
0.000 |
| z |
0.000 |
0.000 |
-11.632 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.762 |
-3.496 |
0.000 |
| y |
-3.496 |
-2.250 |
0.000 |
| z |
0.000 |
0.000 |
0.487 |
|
| Polar |
|---|
| 3z2-r2 | 0.974 |
|---|
| x2-y2 | 2.675 |
|---|
| xy | -3.496 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.434 |
-0.612 |
0.000 |
| y |
-0.612 |
2.376 |
0.000 |
| z |
0.000 |
0.000 |
1.371 |
<r2> (average value of r
2) Å
2
| <r2> |
17.116 |
| (<r2>)1/2 |
4.137 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -114.284898 |
| Energy at 298.15K | -114.286302 |
| HF Energy | -114.284898 |
| Nuclear repulsion energy | 30.653321 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3585 |
3421 |
22.17 |
|
|
|
| 2 |
A' |
2739 |
2614 |
243.45 |
|
|
|
| 3 |
A' |
1483 |
1416 |
53.73 |
|
|
|
| 4 |
A' |
1378 |
1315 |
68.48 |
|
|
|
| 5 |
A' |
1229 |
1173 |
27.77 |
|
|
|
| 6 |
A" |
1026 |
979 |
40.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5720.1 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 5458.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.122 |
0.741 |
0.000 |
| O2 |
0.122 |
-0.564 |
0.000 |
| H3 |
-0.960 |
1.065 |
0.000 |
| H4 |
-0.752 |
-0.999 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
H3 |
H4 |
| C1 | | 1.3055 | 1.1295 | 1.9474 |
O2 | 1.3055 | | 1.9557 | 0.9763 | H3 | 1.1295 | 1.9557 | | 2.0741 | H4 | 1.9474 | 0.9763 | 2.0741 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H4 |
116.436 |
|
O2 |
C1 |
H3 |
106.641 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.009 |
|
|
|
| 2 |
O |
-0.381 |
|
|
|
| 3 |
H |
0.057 |
|
|
|
| 4 |
H |
0.334 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.915 |
-1.708 |
0.000 |
3.379 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-12.068 |
0.049 |
0.000 |
| y |
0.049 |
-12.888 |
0.000 |
| z |
0.000 |
0.000 |
-11.610 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.181 |
0.049 |
0.000 |
| y |
0.049 |
-1.049 |
0.000 |
| z |
0.000 |
0.000 |
0.868 |
|
| Polar |
|---|
| 3z2-r2 | 1.736 |
|---|
| x2-y2 | 0.820 |
|---|
| xy | 0.049 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.305 |
-0.086 |
0.000 |
| y |
-0.086 |
2.825 |
0.000 |
| z |
0.000 |
0.000 |
1.392 |
<r2> (average value of r
2) Å
2
| <r2> |
17.285 |
| (<r2>)1/2 |
4.157 |
