Jump to
S1C2
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -117.114126 |
| Energy at 298.15K | -117.118374 |
| HF Energy | -117.114126 |
| Nuclear repulsion energy | 64.934160 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3289 |
3134 |
15.66 |
|
|
|
| 2 |
A1 |
3188 |
3038 |
1.64 |
|
|
|
| 3 |
A1 |
3177 |
3028 |
15.49 |
|
|
|
| 4 |
A1 |
1532 |
1460 |
2.36 |
|
|
|
| 5 |
A1 |
1283 |
1223 |
0.90 |
|
|
|
| 6 |
A1 |
1047 |
998 |
0.12 |
|
|
|
| 7 |
A1 |
423 |
403 |
0.02 |
|
|
|
| 8 |
A2 |
780 |
743 |
0.00 |
|
|
|
| 9 |
A2 |
555 |
529 |
0.00 |
|
|
|
| 10 |
B1 |
1013 |
966 |
17.49 |
|
|
|
| 11 |
B1 |
805 |
767 |
77.13 |
|
|
|
| 12 |
B1 |
533 |
508 |
13.48 |
|
|
|
| 13 |
B2 |
3286 |
3132 |
4.38 |
|
|
|
| 14 |
B2 |
3181 |
3032 |
7.44 |
|
|
|
| 15 |
B2 |
1530 |
1459 |
0.10 |
|
|
|
| 16 |
B2 |
1425 |
1358 |
6.52 |
|
|
|
| 17 |
B2 |
1222 |
1164 |
0.00 |
|
|
|
| 18 |
B2 |
934 |
890 |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14601.2 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 13916.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.444 |
| H2 |
0.000 |
0.000 |
1.534 |
| C3 |
0.000 |
1.226 |
-0.196 |
| C4 |
0.000 |
-1.226 |
-0.196 |
| H5 |
0.000 |
2.157 |
0.360 |
| H6 |
0.000 |
-2.157 |
0.360 |
| H7 |
0.000 |
1.291 |
-1.281 |
| H8 |
0.000 |
-1.291 |
-1.281 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.0898 | 1.3831 | 1.3831 | 2.1586 | 2.1586 | 2.1540 | 2.1540 |
H2 | 1.0898 | | 2.1203 | 2.1203 | 2.4557 | 2.4557 | 3.0961 | 3.0961 | C3 | 1.3831 | 2.1203 | | 2.4522 | 1.0842 | 3.4285 | 1.0863 | 2.7406 | C4 | 1.3831 | 2.1203 | 2.4522 | | 3.4285 | 1.0842 | 2.7406 | 1.0863 | H5 | 2.1586 | 2.4557 | 1.0842 | 3.4285 | | 4.3140 | 1.8549 | 3.8181 | H6 | 2.1586 | 2.4557 | 3.4285 | 1.0842 | 4.3140 | | 3.8181 | 1.8549 | H7 | 2.1540 | 3.0961 | 1.0863 | 2.7406 | 1.8549 | 3.8181 | | 2.5816 | H8 | 2.1540 | 3.0961 | 2.7406 | 1.0863 | 3.8181 | 1.8549 | 2.5816 | |
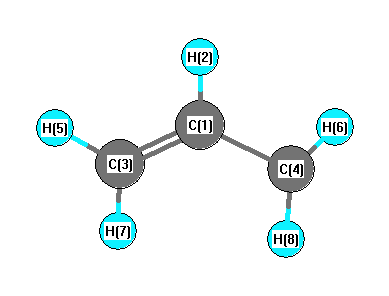 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
H5 |
121.592 |
|
C1 |
C3 |
H7 |
120.978 |
| C1 |
C4 |
H6 |
121.592 |
|
C1 |
C4 |
H8 |
120.978 |
| H2 |
C1 |
C3 |
117.564 |
|
H2 |
C1 |
C4 |
117.564 |
| C3 |
C1 |
C4 |
124.872 |
|
H5 |
C3 |
H7 |
117.430 |
| H6 |
C4 |
H8 |
117.430 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.066 |
|
|
|
| 2 |
H |
0.124 |
|
|
|
| 3 |
C |
-0.290 |
|
|
|
| 4 |
C |
-0.290 |
|
|
|
| 5 |
H |
0.132 |
|
|
|
| 6 |
H |
0.132 |
|
|
|
| 7 |
H |
0.129 |
|
|
|
| 8 |
H |
0.129 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.058 |
0.058 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.582 |
0.000 |
0.000 |
| y |
0.000 |
-17.361 |
0.000 |
| z |
0.000 |
0.000 |
-17.229 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.287 |
0.000 |
0.000 |
| y |
0.000 |
2.045 |
0.000 |
| z |
0.000 |
0.000 |
2.243 |
|
| Polar |
|---|
| 3z2-r2 | 4.485 |
|---|
| x2-y2 | -4.222 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.882 |
0.000 |
0.000 |
| y |
0.000 |
7.368 |
0.000 |
| z |
0.000 |
0.000 |
4.587 |
<r2> (average value of r
2) Å
2
| <r2> |
49.907 |
| (<r2>)1/2 |
7.064 |
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -117.106519 |
| Energy at 298.15K | -117.110769 |
| HF Energy | -117.106519 |
| Nuclear repulsion energy | 64.909517 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3292 |
3137 |
17.16 |
|
|
|
| 2 |
A' |
3290 |
3135 |
5.31 |
|
|
|
| 3 |
A' |
3196 |
3046 |
2.81 |
|
|
|
| 4 |
A' |
3189 |
3040 |
8.72 |
|
|
|
| 5 |
A' |
3181 |
3032 |
15.72 |
|
|
|
| 6 |
A' |
1543 |
1470 |
2.31 |
|
|
|
| 7 |
A' |
1538 |
1466 |
0.11 |
|
|
|
| 8 |
A' |
1435 |
1368 |
6.27 |
|
|
|
| 9 |
A' |
1290 |
1229 |
0.82 |
|
|
|
| 10 |
A' |
1228 |
1170 |
0.00 |
|
|
|
| 11 |
A' |
1051 |
1002 |
0.13 |
|
|
|
| 12 |
A' |
940 |
896 |
0.12 |
|
|
|
| 13 |
A' |
423 |
403 |
0.01 |
|
|
|
| 14 |
A" |
1014 |
967 |
18.02 |
|
|
|
| 15 |
A" |
801 |
764 |
82.00 |
|
|
|
| 16 |
A" |
776 |
740 |
0.00 |
|
|
|
| 17 |
A" |
556 |
530 |
0.00 |
|
|
|
| 18 |
A" |
534 |
509 |
14.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14638.0 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 13951.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.444 |
0.000 |
| H2 |
0.000 |
1.534 |
0.000 |
| C3 |
-1.226 |
-0.196 |
0.000 |
| C4 |
1.226 |
-0.196 |
0.000 |
| H5 |
-2.159 |
0.359 |
0.000 |
| H6 |
2.158 |
0.359 |
0.000 |
| H7 |
-1.291 |
-1.281 |
0.000 |
| H8 |
1.291 |
-1.281 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.0902 | 1.3835 | 1.3835 | 2.1602 | 2.1601 | 2.1550 | 2.1552 |
H2 | 1.0902 | | 2.1213 | 2.1210 | 2.4582 | 2.4576 | 3.0977 | 3.0976 | C3 | 1.3835 | 2.1213 | | 2.4527 | 1.0849 | 3.4299 | 1.0868 | 2.7416 | C4 | 1.3835 | 2.1210 | 2.4527 | | 3.4301 | 1.0848 | 2.7413 | 1.0869 | H5 | 2.1602 | 2.4582 | 1.0849 | 3.4301 | | 4.3169 | 1.8552 | 3.8199 | H6 | 2.1601 | 2.4576 | 3.4299 | 1.0848 | 4.3169 | | 3.8195 | 1.8549 | H7 | 2.1550 | 3.0977 | 1.0868 | 2.7413 | 1.8552 | 3.8195 | | 2.5826 | H8 | 2.1552 | 3.0976 | 2.7416 | 1.0869 | 3.8199 | 1.8549 | 2.5826 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
H5 |
121.654 |
|
C1 |
C3 |
H7 |
120.999 |
| C1 |
C4 |
H6 |
121.651 |
|
C1 |
C4 |
H8 |
121.016 |
| H2 |
C1 |
C3 |
117.593 |
|
H2 |
C1 |
C4 |
117.565 |
| C3 |
C1 |
C4 |
124.843 |
|
H5 |
C3 |
H7 |
117.348 |
| H6 |
C4 |
H8 |
117.333 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.086 |
|
|
|
| 2 |
H |
0.166 |
|
|
|
| 3 |
C |
-0.385 |
|
|
|
| 4 |
C |
-0.385 |
|
|
|
| 5 |
H |
0.174 |
|
|
|
| 6 |
H |
0.174 |
|
|
|
| 7 |
H |
0.171 |
|
|
|
| 8 |
H |
0.171 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.062 |
0.000 |
0.062 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.332 |
0.000 |
0.000 |
| y |
0.000 |
-17.199 |
0.000 |
| z |
0.000 |
0.000 |
-21.648 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.091 |
0.000 |
0.000 |
| y |
0.000 |
2.292 |
0.000 |
| z |
0.000 |
0.000 |
-4.383 |
|
| Polar |
|---|
| 3z2-r2 | -8.766 |
|---|
| x2-y2 | -0.134 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.300 |
-0.000 |
0.000 |
| y |
-0.000 |
4.526 |
0.000 |
| z |
0.000 |
0.000 |
1.815 |
<r2> (average value of r
2) Å
2
| <r2> |
49.937 |
| (<r2>)1/2 |
7.067 |
