Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -117.124400 |
| Energy at 298.15K | -117.128648 |
| HF Energy | -117.124400 |
| Nuclear repulsion energy | 64.941624 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3287 |
3137 |
16.82 |
|
|
|
| 2 |
A1 |
3187 |
3041 |
1.72 |
|
|
|
| 3 |
A1 |
3177 |
3031 |
16.40 |
|
|
|
| 4 |
A1 |
1532 |
1462 |
2.30 |
|
|
|
| 5 |
A1 |
1282 |
1224 |
0.90 |
|
|
|
| 6 |
A1 |
1047 |
999 |
0.11 |
|
|
|
| 7 |
A1 |
423 |
404 |
0.02 |
|
|
|
| 8 |
A2 |
779 |
744 |
0.00 |
|
|
|
| 9 |
A2 |
556 |
531 |
0.00 |
|
|
|
| 10 |
B1 |
1014 |
967 |
17.13 |
|
|
|
| 11 |
B1 |
804 |
767 |
75.80 |
|
|
|
| 12 |
B1 |
533 |
509 |
13.29 |
|
|
|
| 13 |
B2 |
3285 |
3135 |
4.81 |
|
|
|
| 14 |
B2 |
3180 |
3035 |
8.06 |
|
|
|
| 15 |
B2 |
1530 |
1460 |
0.08 |
|
|
|
| 16 |
B2 |
1425 |
1360 |
6.32 |
|
|
|
| 17 |
B2 |
1222 |
1166 |
0.00 |
|
|
|
| 18 |
B2 |
934 |
892 |
0.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14598.9 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 13931.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
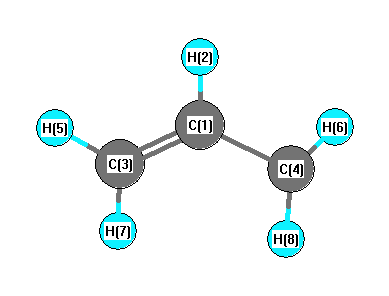
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.444 |
| H2 |
0.000 |
0.000 |
1.533 |
| C3 |
0.000 |
1.226 |
-0.196 |
| C4 |
0.000 |
-1.226 |
-0.196 |
| H5 |
0.000 |
2.157 |
0.359 |
| H6 |
0.000 |
-2.157 |
0.359 |
| H7 |
0.000 |
1.290 |
-1.280 |
| H8 |
0.000 |
-1.290 |
-1.280 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.0897 | 1.3830 | 1.3830 | 2.1586 | 2.1586 | 2.1537 | 2.1537 |
H2 | 1.0897 | | 2.1201 | 2.1201 | 2.4558 | 2.4558 | 3.0957 | 3.0957 | C3 | 1.3830 | 2.1201 | | 2.4519 | 1.0840 | 3.4282 | 1.0862 | 2.7401 | C4 | 1.3830 | 2.1201 | 2.4519 | | 3.4282 | 1.0840 | 2.7401 | 1.0862 | H5 | 2.1586 | 2.4558 | 1.0840 | 3.4282 | | 4.3138 | 1.8545 | 3.8174 | H6 | 2.1586 | 2.4558 | 3.4282 | 1.0840 | 4.3138 | | 3.8174 | 1.8545 | H7 | 2.1537 | 3.0957 | 1.0862 | 2.7401 | 1.8545 | 3.8174 | | 2.5809 | H8 | 2.1537 | 3.0957 | 2.7401 | 1.0862 | 3.8174 | 1.8545 | 2.5809 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
H5 |
121.613 |
|
C1 |
C3 |
H7 |
120.970 |
| C1 |
C4 |
H6 |
121.613 |
|
C1 |
C4 |
H8 |
120.970 |
| H2 |
C1 |
C3 |
117.566 |
|
H2 |
C1 |
C4 |
117.566 |
| C3 |
C1 |
C4 |
124.867 |
|
H5 |
C3 |
H7 |
117.416 |
| H6 |
C4 |
H8 |
117.416 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.060 |
|
|
|
| 2 |
H |
0.120 |
|
|
|
| 3 |
C |
-0.286 |
|
|
|
| 4 |
C |
-0.286 |
|
|
|
| 5 |
H |
0.130 |
|
|
|
| 6 |
H |
0.130 |
|
|
|
| 7 |
H |
0.126 |
|
|
|
| 8 |
H |
0.126 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.059 |
0.059 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.562 |
0.000 |
0.000 |
| y |
0.000 |
-17.377 |
0.000 |
| z |
0.000 |
0.000 |
-17.251 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.248 |
0.000 |
0.000 |
| y |
0.000 |
2.030 |
0.000 |
| z |
0.000 |
0.000 |
2.218 |
|
| Polar |
|---|
| 3z2-r2 | 4.437 |
|---|
| x2-y2 | -4.185 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.880 |
0.000 |
0.000 |
| y |
0.000 |
7.366 |
0.000 |
| z |
0.000 |
0.000 |
4.595 |
<r2> (average value of r
2) Å
2
| <r2> |
49.902 |
| (<r2>)1/2 |
7.064 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -117.124400 |
| Energy at 298.15K | -117.128649 |
| HF Energy | -117.124400 |
| Nuclear repulsion energy | 64.943767 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3287 |
3137 |
16.58 |
|
|
|
| 2 |
A' |
3285 |
3135 |
5.05 |
|
|
|
| 3 |
A' |
3186 |
3041 |
1.60 |
|
|
|
| 4 |
A' |
3180 |
3034 |
8.05 |
|
|
|
| 5 |
A' |
3177 |
3031 |
16.51 |
|
|
|
| 6 |
A' |
1532 |
1462 |
2.32 |
|
|
|
| 7 |
A' |
1530 |
1460 |
0.08 |
|
|
|
| 8 |
A' |
1425 |
1360 |
6.33 |
|
|
|
| 9 |
A' |
1283 |
1224 |
0.91 |
|
|
|
| 10 |
A' |
1221 |
1166 |
0.00 |
|
|
|
| 11 |
A' |
1047 |
999 |
0.11 |
|
|
|
| 12 |
A' |
935 |
892 |
0.11 |
|
|
|
| 13 |
A' |
424 |
404 |
0.02 |
|
|
|
| 14 |
A" |
1014 |
967 |
17.14 |
|
|
|
| 15 |
A" |
804 |
768 |
75.79 |
|
|
|
| 16 |
A" |
779 |
744 |
0.00 |
|
|
|
| 17 |
A" |
557 |
531 |
0.00 |
|
|
|
| 18 |
A" |
534 |
509 |
13.31 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14599.0 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 13931.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.444 |
0.000 |
| H2 |
0.000 |
1.534 |
0.000 |
| C3 |
-1.226 |
-0.196 |
0.000 |
| C4 |
1.226 |
-0.196 |
0.000 |
| H5 |
-2.157 |
0.359 |
0.000 |
| H6 |
2.157 |
0.358 |
0.000 |
| H7 |
-1.289 |
-1.281 |
0.000 |
| H8 |
1.289 |
-1.281 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.0897 | 1.3830 | 1.3830 | 2.1589 | 2.1587 | 2.1532 | 2.1534 |
H2 | 1.0897 | | 2.1205 | 2.1201 | 2.4568 | 2.4562 | 3.0957 | 3.0956 | C3 | 1.3830 | 2.1205 | | 2.4515 | 1.0841 | 3.4280 | 1.0862 | 2.7388 | C4 | 1.3830 | 2.1201 | 2.4515 | | 3.4281 | 1.0840 | 2.7386 | 1.0862 | H5 | 2.1589 | 2.4568 | 1.0841 | 3.4281 | | 4.3142 | 1.8548 | 3.8164 | H6 | 2.1587 | 2.4562 | 3.4280 | 1.0840 | 4.3142 | | 3.8160 | 1.8546 | H7 | 2.1532 | 3.0957 | 1.0862 | 2.7386 | 1.8548 | 3.8160 | | 2.5783 | H8 | 2.1534 | 3.0956 | 2.7388 | 1.0862 | 3.8164 | 1.8546 | 2.5783 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
H5 |
121.633 |
|
C1 |
C3 |
H7 |
120.927 |
| C1 |
C4 |
H6 |
121.629 |
|
C1 |
C4 |
H8 |
120.941 |
| H2 |
C1 |
C3 |
117.601 |
|
H2 |
C1 |
C4 |
117.573 |
| C3 |
C1 |
C4 |
124.826 |
|
H5 |
C3 |
H7 |
117.440 |
| H6 |
C4 |
H8 |
117.430 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.060 |
|
|
|
| 2 |
H |
0.120 |
|
|
|
| 3 |
C |
-0.286 |
|
|
|
| 4 |
C |
-0.286 |
|
|
|
| 5 |
H |
0.130 |
|
|
|
| 6 |
H |
0.130 |
|
|
|
| 7 |
H |
0.126 |
|
|
|
| 8 |
H |
0.126 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.059 |
0.000 |
0.059 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.378 |
0.000 |
0.000 |
| y |
0.000 |
-17.251 |
0.000 |
| z |
0.000 |
0.000 |
-21.562 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.029 |
0.000 |
0.000 |
| y |
0.000 |
2.219 |
0.000 |
| z |
0.000 |
0.000 |
-4.248 |
|
| Polar |
|---|
| 3z2-r2 | -8.496 |
|---|
| x2-y2 | -0.127 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.365 |
-0.000 |
0.000 |
| y |
-0.000 |
4.595 |
0.000 |
| z |
0.000 |
0.000 |
1.880 |
<r2> (average value of r
2) Å
2
| <r2> |
49.894 |
| (<r2>)1/2 |
7.064 |
